return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHClCHCH2CH3 ((E)-1-Chloro-1-butene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-612.560430
Energy at 298.15K-612.567559
HF Energy-612.064639
Nuclear repulsion energy199.459896
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3245 3087 6.45      
2 A 3192 3037 5.21      
3 A 3169 3015 25.38      
4 A 3160 3006 25.46      
5 A 3121 2969 7.52      
6 A 3087 2936 19.95      
7 A 3079 2929 16.72      
8 A 1672 1590 25.60      
9 A 1604 1526 4.48      
10 A 1601 1523 7.09      
11 A 1579 1502 4.81      
12 A 1492 1419 2.86      
13 A 1406 1338 1.23      
14 A 1376 1309 11.49      
15 A 1364 1297 4.15      
16 A 1324 1259 5.21      
17 A 1177 1120 5.58      
18 A 1143 1088 2.61      
19 A 1023 973 5.08      
20 A 996 947 55.02      
21 A 922 877 0.32      
22 A 852 811 21.10      
23 A 820 780 13.90      
24 A 773 736 19.90      
25 A 457 435 1.67      
26 A 383 365 3.95      
27 A 301 286 0.28      
28 A 209 199 0.23      
29 A 169 161 0.23      
30 A 92 87 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 22393.2 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 21302.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.56072 0.04649 0.04543

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.280 -0.683 1.241
C2 0.405 0.055 0.450
H3 -0.600 1.162 -1.082
Cl4 -2.269 -0.198 -0.016
C5 -0.648 0.436 -0.279
H6 1.784 1.373 -0.548
H7 2.201 1.014 1.136
C8 1.812 0.577 0.208
H9 3.772 -0.181 -0.422
H10 2.795 -1.363 0.479
H11 2.390 -0.983 -1.208
C12 2.757 -0.563 -0.267

Atom - Atom Distances (Å)
  H1 C2 H3 Cl4 C5 H6 H7 C8 H9 H10 H11 C12
H11.08873.09392.88312.10323.11262.56602.23653.90032.71423.24672.9027
C21.08872.14072.72651.33662.15272.14881.51983.48612.77882.78702.5354
H33.09392.14072.40251.08342.45193.57592.79644.62124.50973.68213.8612
Cl42.88312.72652.40251.76034.37934.77314.15986.05515.21994.87315.0460
C52.10321.33661.08341.76032.62013.23352.51154.46583.95843.48023.5492
H63.11262.15272.45194.37932.62011.77141.09762.52703.09252.52102.1849
H72.56602.14883.57594.77313.23351.77141.09742.51472.53683.08572.1828
C82.23651.51982.79644.15982.51151.09761.09742.19462.19232.18531.5556
H93.90033.48614.62126.05514.46582.52702.51472.19461.77881.78101.0952
H102.71422.77884.50975.21993.95843.09252.53682.19231.77881.77681.0951
H113.24672.78703.68214.87313.48022.52103.08572.18531.78101.77681.0943
C122.90272.53543.86125.04603.54922.18492.18281.55561.09521.09511.0943

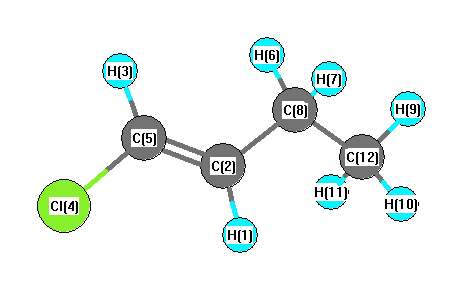
picture of (E)-1-Chloro-1-butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C5 119.914 H1 C2 C8 117.088
C2 C5 H3 124.064 C2 C5 Cl4 122.786
C2 C8 H6 109.610 C2 C8 H7 109.316
C2 C8 C12 111.054 H3 C5 Cl4 113.150
C5 C2 C8 122.977 H6 C8 H7 107.611
H6 C8 C12 109.665 H7 C8 C12 109.516
C8 C12 H9 110.564 C8 C12 H10 110.385
C8 C12 H11 109.885 H9 C12 H10 108.606
H9 C12 H11 108.862 H10 C12 H11 108.490
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability