Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -634.358868 |
| Energy at 298.15K | -634.363757 |
| HF Energy | -633.927762 |
| Nuclear repulsion energy | 155.025211 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3163 |
3009 |
9.16 |
|
|
|
| 2 |
A' |
3124 |
2972 |
15.72 |
|
|
|
| 3 |
A' |
1628 |
1549 |
0.54 |
|
|
|
| 4 |
A' |
1586 |
1508 |
5.76 |
|
|
|
| 5 |
A' |
1485 |
1412 |
3.75 |
|
|
|
| 6 |
A' |
1349 |
1284 |
13.69 |
|
|
|
| 7 |
A' |
1075 |
1023 |
46.52 |
|
|
|
| 8 |
A' |
1047 |
996 |
5.48 |
|
|
|
| 9 |
A' |
762 |
725 |
39.11 |
|
|
|
| 10 |
A' |
369 |
351 |
1.14 |
|
|
|
| 11 |
A' |
238 |
226 |
14.55 |
|
|
|
| 12 |
A" |
3229 |
3072 |
6.50 |
|
|
|
| 13 |
A" |
3177 |
3022 |
20.71 |
|
|
|
| 14 |
A" |
1375 |
1308 |
0.41 |
|
|
|
| 15 |
A" |
1261 |
1200 |
0.56 |
|
|
|
| 16 |
A" |
1115 |
1061 |
2.53 |
|
|
|
| 17 |
A" |
843 |
802 |
1.17 |
|
|
|
| 18 |
A" |
137 |
130 |
11.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13480.9 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12824.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.617 |
0.000 |
| C2 |
0.993 |
-0.550 |
0.000 |
| Cl3 |
-1.687 |
-0.041 |
0.000 |
| F4 |
2.306 |
0.020 |
0.000 |
| H5 |
0.126 |
1.228 |
0.892 |
| H6 |
0.126 |
1.228 |
-0.892 |
| H7 |
0.857 |
-1.165 |
0.892 |
| H8 |
0.857 |
-1.165 |
-0.892 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5323 | 1.8107 | 2.3818 | 1.0889 | 1.0889 | 2.1689 | 2.1689 |
C2 | 1.5323 | | 2.7281 | 1.4308 | 2.1704 | 2.1704 | 1.0919 | 1.0919 | Cl3 | 1.8107 | 2.7281 | | 3.9932 | 2.3861 | 2.3861 | 2.9207 | 2.9207 | F4 | 2.3818 | 1.4308 | 3.9932 | | 2.6475 | 2.6475 | 2.0729 | 2.0729 | H5 | 1.0889 | 2.1704 | 2.3861 | 2.6475 | | 1.7846 | 2.5020 | 3.0732 | H6 | 1.0889 | 2.1704 | 2.3861 | 2.6475 | 1.7846 | | 3.0732 | 2.5020 | H7 | 2.1689 | 1.0919 | 2.9207 | 2.0729 | 2.5020 | 3.0732 | | 1.7845 | H8 | 2.1689 | 1.0919 | 2.9207 | 2.0729 | 3.0732 | 2.5020 | 1.7845 | |
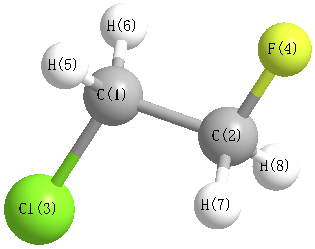 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
106.945 |
|
C1 |
C2 |
H7 |
110.348 |
| C1 |
C2 |
H8 |
110.348 |
|
C2 |
C1 |
Cl3 |
109.099 |
| C2 |
C1 |
H5 |
110.650 |
|
C2 |
C1 |
H6 |
110.650 |
| Cl3 |
C1 |
H5 |
108.158 |
|
Cl3 |
C1 |
H6 |
108.158 |
| F4 |
C2 |
H7 |
109.784 |
|
F4 |
C2 |
H8 |
109.784 |
| H5 |
C1 |
H6 |
110.051 |
|
H7 |
C2 |
H8 |
109.597 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -634.357744 |
| Energy at 298.15K | -634.362726 |
| HF Energy | -633.926252 |
| Nuclear repulsion energy | 159.661346 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3218 |
3061 |
3.47 |
|
|
|
| 2 |
A |
3162 |
3008 |
26.48 |
|
|
|
| 3 |
A |
3150 |
2996 |
4.42 |
|
|
|
| 4 |
A |
3097 |
2947 |
23.81 |
|
|
|
| 5 |
A |
1611 |
1533 |
1.19 |
|
|
|
| 6 |
A |
1564 |
1488 |
8.25 |
|
|
|
| 7 |
A |
1492 |
1420 |
12.54 |
|
|
|
| 8 |
A |
1409 |
1340 |
19.42 |
|
|
|
| 9 |
A |
1316 |
1252 |
1.80 |
|
|
|
| 10 |
A |
1274 |
1212 |
0.37 |
|
|
|
| 11 |
A |
1122 |
1068 |
5.70 |
|
|
|
| 12 |
A |
1098 |
1044 |
37.15 |
|
|
|
| 13 |
A |
991 |
943 |
3.46 |
|
|
|
| 14 |
A |
879 |
836 |
3.15 |
|
|
|
| 15 |
A |
674 |
641 |
25.90 |
|
|
|
| 16 |
A |
462 |
439 |
13.26 |
|
|
|
| 17 |
A |
283 |
270 |
1.17 |
|
|
|
| 18 |
A |
124 |
118 |
2.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13463.9 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12808.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.089 |
0.877 |
-0.303 |
| C2 |
1.214 |
0.440 |
0.360 |
| Cl3 |
-1.422 |
-0.292 |
0.069 |
| F4 |
1.636 |
-0.802 |
-0.199 |
| H5 |
-0.394 |
1.857 |
0.066 |
| H6 |
0.027 |
0.898 |
-1.386 |
| H7 |
1.980 |
1.201 |
0.171 |
| H8 |
1.072 |
0.325 |
1.436 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5260 | 1.8116 | 2.4098 | 1.0902 | 1.0895 | 2.1464 | 2.1632 |
C2 | 1.5260 | | 2.7511 | 1.4255 | 2.1638 | 2.1602 | 1.0956 | 1.0922 | Cl3 | 1.8116 | 2.7511 | | 3.1118 | 2.3815 | 2.3732 | 3.7158 | 2.9106 | F4 | 2.4098 | 1.4255 | 3.1118 | | 3.3560 | 2.6244 | 2.0652 | 2.0647 | H5 | 1.0902 | 2.1638 | 2.3815 | 3.3560 | | 1.7904 | 2.4653 | 2.5251 | H6 | 1.0895 | 2.1602 | 2.3732 | 2.6244 | 1.7904 | | 2.5152 | 3.0639 | H7 | 2.1464 | 1.0956 | 3.7158 | 2.0652 | 2.4653 | 2.5152 | | 1.7868 | H8 | 2.1632 | 1.0922 | 2.9106 | 2.0647 | 2.5251 | 3.0639 | 1.7868 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.415 |
|
C1 |
C2 |
H7 |
108.809 |
| C1 |
C2 |
H8 |
110.321 |
|
C2 |
C1 |
Cl3 |
110.738 |
| C2 |
C1 |
H5 |
110.492 |
|
C2 |
C1 |
H6 |
110.248 |
| Cl3 |
C1 |
H5 |
107.698 |
|
Cl3 |
C1 |
H6 |
107.135 |
| F4 |
C2 |
H7 |
109.298 |
|
F4 |
C2 |
H8 |
109.471 |
| H5 |
C1 |
H6 |
110.447 |
|
H7 |
C2 |
H8 |
109.507 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
