return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (2-Hexyne)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-232.234578
Energy at 298.15K-232.243210
HF Energy-231.688467
Nuclear repulsion energy204.336525
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3155 3001 27.66      
2 A 3142 2989 12.12      
3 A 3100 2949 19.06      
4 A 3079 2929 15.01      
5 A 3078 2929 22.98      
6 A 3067 2918 13.34      
7 A 2276 2165 0.01      
8 A 1606 1528 4.80      
9 A 1592 1514 1.01      
10 A 1590 1513 6.69      
11 A 1575 1498 1.09      
12 A 1497 1425 0.65      
13 A 1492 1419 4.12      
14 A 1424 1355 1.44      
15 A 1366 1299 5.50      
16 A 1151 1095 8.82      
17 A 1127 1072 2.70      
18 A 1114 1059 1.45      
19 A 1024 975 1.08      
20 A 897 854 3.49      
21 A 744 707 0.32      
22 A 445 424 2.62      
23 A 323 308 0.50      
24 A 213 203 4.68      
25 A 92 87 1.64      
26 A 3161 3007 50.41      
27 A 3143 2990 11.87      
28 A 3137 2984 0.15      
29 A 3103 2952 6.68      
30 A 1603 1525 7.23      
31 A 1590 1513 5.89      
32 A 1390 1322 0.11      
33 A 1332 1267 0.17      
34 A 1185 1127 0.75      
35 A 1114 1059 2.09      
36 A 924 879 0.52      
37 A 768 731 3.63      
38 A 280 267 0.49      
39 A 245 233 0.02      
40 A 198 188 8.09      
41 A 82 78 0.49      
42 A 7 6 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 31713.6 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30169.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.58319 0.03912 0.03770

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.255 -2.847 0.000
C2 -1.781 2.873 0.000
C3 0.958 -0.296 0.000
C4 0.000 0.835 0.000
C5 0.238 -1.678 0.000
C6 -0.805 1.757 0.000
H7 0.735 -3.812 0.000
H8 -2.808 2.491 0.000
H9 1.894 -2.798 0.889
H10 1.894 -2.798 -0.889
H11 -0.406 -1.740 -0.885
H12 -0.406 -1.740 0.885
H13 1.606 -0.235 -0.885
H14 1.606 -0.235 0.885
H15 -1.650 3.503 0.888
H16 -1.650 3.503 -0.888

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C16.47572.56813.89011.54955.04421.09536.70821.09601.09602.18312.18312.78052.78057.03907.0390
C26.47574.18862.70644.97861.48207.14261.09636.81596.81594.89414.89414.68134.68131.09631.0963
C32.56814.18861.48231.55792.70663.52234.68512.81532.81532.17372.17371.09911.09914.69294.6929
C43.89012.70641.48232.52421.22444.70453.25994.19254.19252.75252.75252.12332.12333.26013.2601
C51.54954.97861.55792.52423.59002.19105.16292.18802.18801.09581.09582.17682.17685.58505.5850
C65.04421.48202.70661.22443.59005.77802.13275.36915.36913.62913.62913.25093.25092.13292.1329
H71.09537.14263.52234.70452.19105.77807.23031.77731.77732.52542.52543.78623.78627.74477.7447
H86.70821.09634.68513.25995.16292.13277.23037.13247.13244.94484.94485.26305.26301.77581.7758
H91.09606.81592.81534.19252.18805.36911.77737.13241.77803.09092.53133.13082.57947.22937.4444
H101.09606.81592.81534.19252.18805.36911.77737.13241.77802.53133.09092.57943.13087.44447.2293
H112.18314.89412.17372.75251.09583.62912.52544.94483.09092.53131.76952.51233.07335.67235.3882
H122.18314.89412.17372.75251.09583.62912.52544.94482.53133.09091.76953.07332.51235.38825.6723
H132.78054.68131.09912.12332.17683.25093.78625.26303.13082.57942.51233.07331.77075.26494.9572
H142.78054.68131.09912.12332.17683.25093.78625.26302.57943.13083.07332.51231.77074.95725.2649
H157.03901.09634.69293.26015.58502.13297.74471.77587.22937.44445.67235.38825.26494.95721.7759
H167.03901.09634.69293.26015.58502.13297.74471.77587.44447.22935.38825.67234.95725.26491.7759

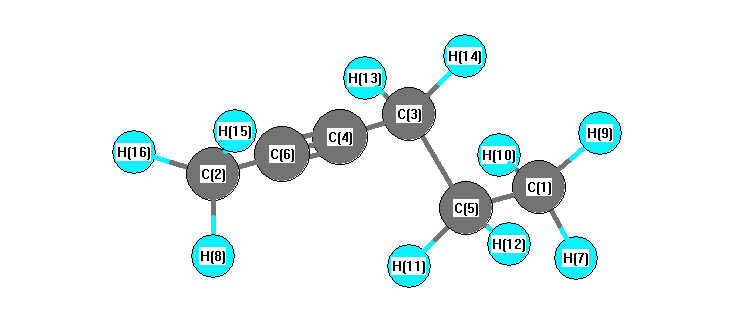
picture of 2-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 C3 111.465 C1 C5 H11 110.048
C1 C5 H12 110.048 C2 C6 C4 179.997
C3 C4 C6 179.113 C3 C5 H11 108.745
C3 C5 H12 108.745 C4 C3 C5 112.224
C4 C3 H13 109.789 C4 C3 H14 109.789
C5 C1 H7 110.694 C5 C1 H9 110.425
C5 C1 H10 110.425 C5 C3 H13 108.793
C5 C3 H14 108.793 C6 C2 H8 110.728
C6 C2 H15 110.744 C6 C2 H16 110.744
H7 C1 H9 108.404 H7 C1 H10 108.404
H8 C2 H15 108.171 H8 C2 H16 108.171
H9 C1 H10 108.417 H11 C5 H12 107.695
H13 C3 H14 107.321 H15 C2 H16 108.182
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability