Jump to
S1C2
Vibrational Frequencies calculated at MP2/3-21G*
Geometric Data calculated at MP2/3-21G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -233.427972 |
| Energy at 298.15K | -233.440270 |
| HF Energy | -232.879950 |
| Nuclear repulsion energy | 229.233293 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3181 |
3026 |
52.75 |
|
|
|
| 2 |
A |
3166 |
3012 |
5.18 |
|
|
|
| 3 |
A |
3156 |
3003 |
56.86 |
|
|
|
| 4 |
A |
3152 |
2999 |
31.37 |
|
|
|
| 5 |
A |
3147 |
2994 |
1.82 |
|
|
|
| 6 |
A |
3132 |
2979 |
7.28 |
|
|
|
| 7 |
A |
3131 |
2978 |
17.13 |
|
|
|
| 8 |
A |
3115 |
2963 |
1.72 |
|
|
|
| 9 |
A |
3085 |
2935 |
29.92 |
|
|
|
| 10 |
A |
3078 |
2928 |
12.74 |
|
|
|
| 11 |
A |
3078 |
2928 |
20.04 |
|
|
|
| 12 |
A |
3071 |
2921 |
11.59 |
|
|
|
| 13 |
A |
1701 |
1618 |
0.24 |
|
|
|
| 14 |
A |
1606 |
1528 |
5.85 |
|
|
|
| 15 |
A |
1603 |
1525 |
7.02 |
|
|
|
| 16 |
A |
1599 |
1521 |
5.72 |
|
|
|
| 17 |
A |
1592 |
1515 |
3.15 |
|
|
|
| 18 |
A |
1590 |
1512 |
2.66 |
|
|
|
| 19 |
A |
1577 |
1500 |
0.81 |
|
|
|
| 20 |
A |
1491 |
1419 |
1.23 |
|
|
|
| 21 |
A |
1490 |
1418 |
7.11 |
|
|
|
| 22 |
A |
1486 |
1413 |
5.88 |
|
|
|
| 23 |
A |
1421 |
1352 |
0.97 |
|
|
|
| 24 |
A |
1389 |
1322 |
0.01 |
|
|
|
| 25 |
A |
1361 |
1295 |
0.41 |
|
|
|
| 26 |
A |
1352 |
1286 |
0.96 |
|
|
|
| 27 |
A |
1322 |
1258 |
0.21 |
|
|
|
| 28 |
A |
1213 |
1154 |
1.66 |
|
|
|
| 29 |
A |
1150 |
1094 |
3.53 |
|
|
|
| 30 |
A |
1129 |
1074 |
0.45 |
|
|
|
| 31 |
A |
1109 |
1055 |
0.76 |
|
|
|
| 32 |
A |
1040 |
989 |
2.30 |
|
|
|
| 33 |
A |
1019 |
969 |
0.42 |
|
|
|
| 34 |
A |
996 |
948 |
4.51 |
|
|
|
| 35 |
A |
935 |
889 |
0.83 |
|
|
|
| 36 |
A |
911 |
867 |
10.35 |
|
|
|
| 37 |
A |
879 |
837 |
3.48 |
|
|
|
| 38 |
A |
771 |
733 |
2.61 |
|
|
|
| 39 |
A |
736 |
700 |
36.04 |
|
|
|
| 40 |
A |
589 |
561 |
7.25 |
|
|
|
| 41 |
A |
483 |
460 |
1.17 |
|
|
|
| 42 |
A |
372 |
354 |
0.02 |
|
|
|
| 43 |
A |
304 |
290 |
0.05 |
|
|
|
| 44 |
A |
249 |
237 |
0.00 |
|
|
|
| 45 |
A |
203 |
193 |
0.01 |
|
|
|
| 46 |
A |
108 |
103 |
0.26 |
|
|
|
| 47 |
A |
95 |
91 |
0.00 |
|
|
|
| 48 |
A |
35 |
33 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37198.9 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 35387.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.387 |
1.017 |
-0.017 |
| C2 |
1.993 |
-0.442 |
-0.187 |
| C3 |
0.799 |
-0.999 |
0.095 |
| C4 |
-0.445 |
-0.293 |
0.608 |
| C5 |
-1.520 |
-0.157 |
-0.511 |
| C6 |
-2.817 |
0.512 |
0.010 |
| H7 |
1.567 |
1.612 |
0.394 |
| H8 |
3.248 |
1.106 |
0.656 |
| H9 |
2.674 |
1.451 |
-0.983 |
| H10 |
2.781 |
-1.085 |
-0.578 |
| H11 |
0.672 |
-2.065 |
-0.095 |
| H12 |
-0.201 |
0.706 |
0.989 |
| H13 |
-0.874 |
-0.866 |
1.442 |
| H14 |
-1.095 |
0.434 |
-1.331 |
| H15 |
-1.749 |
-1.154 |
-0.909 |
| H16 |
-3.560 |
0.603 |
-0.790 |
| H17 |
-3.252 |
-0.083 |
0.821 |
| H18 |
-2.599 |
1.514 |
0.396 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5216 | 2.5687 | 3.1828 | 4.1094 | 5.2286 | 1.0935 | 1.0970 | 1.0971 | 2.2118 | 3.5290 | 2.7943 | 4.0385 | 3.7669 | 4.7558 | 6.0113 | 5.8064 | 5.0275 |
C2 | 1.5216 | | 1.3466 | 2.5688 | 3.5391 | 4.9076 | 2.1770 | 2.1646 | 2.1637 | 1.0900 | 2.0950 | 2.7412 | 3.3241 | 3.4075 | 3.8769 | 5.6825 | 5.3531 | 5.0250 | C3 | 2.5687 | 1.3466 | | 1.5201 | 2.5403 | 3.9200 | 2.7374 | 3.2778 | 3.2672 | 2.0948 | 1.0909 | 2.1693 | 2.1521 | 2.7700 | 2.7434 | 4.7277 | 4.2168 | 4.2369 | C4 | 3.1828 | 2.5688 | 1.5201 | | 1.5566 | 2.5747 | 2.7788 | 3.9502 | 3.9115 | 3.5278 | 2.2097 | 1.0964 | 1.0991 | 2.1696 | 2.1774 | 3.5293 | 2.8226 | 2.8188 | C5 | 4.1094 | 3.5391 | 2.5403 | 1.5566 | | 1.5495 | 3.6707 | 5.0689 | 4.5159 | 4.4006 | 2.9352 | 2.1751 | 2.1752 | 1.0967 | 1.0975 | 2.1950 | 2.1863 | 2.1860 | C6 | 5.2286 | 4.9076 | 3.9200 | 2.5747 | 1.5495 | | 4.5359 | 6.1287 | 5.6582 | 5.8512 | 4.3384 | 2.7996 | 2.7793 | 2.1837 | 2.1815 | 1.0956 | 1.0959 | 1.0960 | H7 | 1.0935 | 2.1770 | 2.7374 | 2.7788 | 3.6707 | 4.5359 | | 1.7756 | 1.7746 | 3.1138 | 3.8162 | 2.0736 | 3.6324 | 3.3831 | 4.5101 | 5.3574 | 5.1259 | 4.1665 | H8 | 1.0970 | 2.1646 | 3.2778 | 3.9502 | 5.0689 | 6.1287 | 1.7756 | | 1.7714 | 2.5584 | 4.1551 | 3.4887 | 4.6369 | 4.8233 | 5.7038 | 6.9784 | 6.6105 | 5.8672 | H9 | 1.0971 | 2.1637 | 3.2672 | 3.9115 | 4.5159 | 5.6582 | 1.7746 | 1.7714 | | 2.5704 | 4.1426 | 3.5650 | 4.8820 | 3.9186 | 5.1331 | 6.2939 | 6.3815 | 5.4503 | H10 | 2.2118 | 1.0900 | 2.0948 | 3.5278 | 4.4006 | 5.8512 | 3.1138 | 2.5584 | 2.5704 | | 2.3761 | 3.8156 | 4.1819 | 4.2308 | 4.5429 | 6.5655 | 6.2742 | 6.0539 | H11 | 3.5290 | 2.0950 | 1.0909 | 2.2097 | 2.9352 | 4.3384 | 3.8162 | 4.1551 | 4.1426 | 2.3761 | | 3.1008 | 2.4873 | 3.3008 | 2.7116 | 5.0505 | 4.4904 | 4.8730 | H12 | 2.7943 | 2.7412 | 2.1693 | 1.0964 | 2.1751 | 2.7996 | 2.0736 | 3.4887 | 3.5650 | 3.8156 | 3.1008 | | 1.7688 | 2.5003 | 3.0750 | 3.8020 | 3.1556 | 2.5985 | H13 | 4.0385 | 3.3241 | 2.1521 | 1.0991 | 2.1752 | 2.7793 | 3.6324 | 4.6369 | 4.8820 | 4.1819 | 2.4873 | 1.7688 | | 3.0701 | 2.5250 | 3.7887 | 2.5797 | 3.1199 | H14 | 3.7669 | 3.4075 | 2.7700 | 2.1696 | 1.0967 | 2.1837 | 3.3831 | 4.8233 | 3.9186 | 4.2308 | 3.3008 | 2.5003 | 3.0701 | | 1.7683 | 2.5292 | 3.0906 | 2.5316 | H15 | 4.7558 | 3.8769 | 2.7434 | 2.1774 | 1.0975 | 2.1815 | 4.5101 | 5.7038 | 5.1331 | 4.5429 | 2.7116 | 3.0750 | 2.5250 | 1.7683 | | 2.5258 | 2.5299 | 3.0890 | H16 | 6.0113 | 5.6825 | 4.7277 | 3.5293 | 2.1950 | 1.0956 | 5.3574 | 6.9784 | 6.2939 | 6.5655 | 5.0505 | 3.8020 | 3.7887 | 2.5292 | 2.5258 | | 1.7781 | 1.7778 | H17 | 5.8064 | 5.3531 | 4.2168 | 2.8226 | 2.1863 | 1.0959 | 5.1259 | 6.6105 | 6.3815 | 6.2742 | 4.4904 | 3.1556 | 2.5797 | 3.0906 | 2.5299 | 1.7781 | | 1.7768 | H18 | 5.0275 | 5.0250 | 4.2369 | 2.8188 | 2.1860 | 1.0960 | 4.1665 | 5.8672 | 5.4503 | 6.0539 | 4.8730 | 2.5985 | 3.1199 | 2.5316 | 3.0890 | 1.7778 | 1.7768 | |
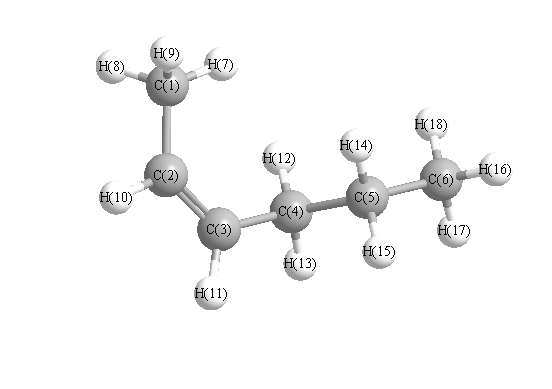 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.062 |
|
C1 |
C2 |
H10 |
114.746 |
| C2 |
C1 |
H7 |
111.662 |
|
C2 |
C1 |
H8 |
110.457 |
| C2 |
C1 |
H9 |
110.385 |
|
C2 |
C3 |
C4 |
127.191 |
| C2 |
C3 |
H11 |
118.140 |
|
C3 |
C2 |
H10 |
118.191 |
| C3 |
C4 |
C5 |
111.302 |
|
C3 |
C4 |
H12 |
110.970 |
| C3 |
C4 |
H13 |
109.451 |
|
C4 |
C3 |
H11 |
114.620 |
| C4 |
C5 |
C6 |
111.971 |
|
C4 |
C5 |
H14 |
108.461 |
| C4 |
C5 |
H15 |
109.022 |
|
C5 |
C4 |
H12 |
108.907 |
| C5 |
C4 |
H13 |
108.758 |
|
C5 |
C6 |
H16 |
111.000 |
| C5 |
C6 |
H17 |
110.295 |
|
C5 |
C6 |
H18 |
110.268 |
| C6 |
C5 |
H14 |
110.046 |
|
C6 |
C5 |
H15 |
109.823 |
| H7 |
C1 |
H8 |
108.316 |
|
H7 |
C1 |
H9 |
108.217 |
| H8 |
C1 |
H9 |
107.678 |
|
H12 |
C4 |
H13 |
107.346 |
| H14 |
C5 |
H16 |
94.483 |
|
H16 |
C6 |
H17 |
108.456 |
| H16 |
C6 |
H18 |
108.428 |
|
H17 |
C6 |
H18 |
108.317 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
