Jump to
S2C1
Energy calculated at mPW1PW91/3-21G*
| | hartrees |
|---|
| Energy at 0K | -149.440743 |
| Energy at 298.15K | -149.440733 |
| HF Energy | -149.440743 |
| Nuclear repulsion energy | 26.264385 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/3-21G*
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.645 |
| O2 |
0.000 |
0.000 |
-0.645 |
Atom - Atom Distances (Å)
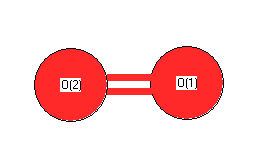 More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
0.000 |
|
|
|
| 2 |
O |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-9.655 |
0.000 |
0.000 |
| y |
0.000 |
-9.655 |
0.000 |
| z |
0.000 |
0.000 |
-10.355 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.350 |
0.000 |
0.000 |
| y |
0.000 |
0.350 |
0.000 |
| z |
0.000 |
0.000 |
-0.701 |
|
| Polar |
|---|
| 3z2-r2 | -1.401 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.297 |
0.000 |
0.000 |
| y |
0.000 |
0.297 |
0.000 |
| z |
0.000 |
0.000 |
1.800 |
<r2> (average value of r
2) Å
2
| <r2> |
12.827 |
| (<r2>)1/2 |
3.581 |
Jump to
S1C1
Energy calculated at mPW1PW91/3-21G*
| | hartrees |
|---|
| Energy at 0K | -149.374985 |
| Energy at 298.15K | -149.374974 |
| HF Energy | -149.374985 |
| Nuclear repulsion energy | 26.174854 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/3-21G*
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.647 |
| O2 |
0.000 |
0.000 |
-0.647 |
Atom - Atom Distances (Å)
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
0.000 |
|
|
|
| 2 |
O |
0.000 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-10.534 |
0.000 |
0.000 |
| y |
0.000 |
-8.829 |
0.000 |
| z |
0.000 |
0.000 |
-10.359 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.940 |
0.000 |
0.000 |
| y |
0.000 |
1.618 |
0.000 |
| z |
0.000 |
0.000 |
-0.678 |
|
| Polar |
|---|
| 3z2-r2 | -1.356 |
|---|
| x2-y2 | -1.705 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.128 |
0.000 |
0.000 |
| y |
0.000 |
0.393 |
0.000 |
| z |
0.000 |
0.000 |
1.637 |
<r2> (average value of r
2) Å
2
| <r2> |
12.885 |
| (<r2>)1/2 |
3.590 |
