Jump to
S1C2
Vibrational Frequencies calculated at TPSSh/3-21G*
Geometric Data calculated at TPSSh/3-21G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at TPSSh/3-21G*
| | hartrees |
|---|
| Energy at 0K | -621.912278 |
| Energy at 298.15K | -621.916037 |
| HF Energy | -621.912278 |
| Nuclear repulsion energy | 189.683333 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3358 |
3243 |
13.51 |
|
|
|
| 2 |
A' |
1225 |
1183 |
122.95 |
|
|
|
| 3 |
A' |
1100 |
1062 |
4.80 |
|
|
|
| 4 |
A' |
745 |
719 |
131.73 |
|
|
|
| 5 |
A' |
500 |
483 |
97.96 |
|
|
|
| 6 |
A' |
419 |
404 |
104.27 |
|
|
|
| 7 |
A' |
281 |
271 |
19.53 |
|
|
|
| 8 |
A" |
3356 |
3241 |
5.77 |
|
|
|
| 9 |
A" |
1064 |
1028 |
43.44 |
|
|
|
| 10 |
A" |
760 |
734 |
326.05 |
|
|
|
| 11 |
A" |
454 |
438 |
90.32 |
|
|
|
| 12 |
A" |
191 |
185 |
10.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6725.9 cm
-1
Scaled (by 0.9657) Zero Point Vibrational Energy (zpe) 6495.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.301 |
0.344 |
0.000 |
| O2 |
-1.043 |
0.974 |
0.000 |
| O3 |
0.301 |
-0.707 |
1.288 |
| O4 |
0.301 |
-0.707 |
-1.288 |
| H5 |
-0.642 |
-0.992 |
1.506 |
| H6 |
-0.642 |
-0.992 |
-1.506 |
Atom - Atom Distances (Å)
| |
S1 |
O2 |
O3 |
O4 |
H5 |
H6 |
| S1 | | 1.4841 | 1.6626 | 1.6626 | 2.2232 | 2.2232 |
O2 | 1.4841 | | 2.5083 | 2.5083 | 2.5090 | 2.5090 | O3 | 1.6626 | 2.5083 | | 2.5768 | 1.0089 | 2.9632 | O4 | 1.6626 | 2.5083 | 2.5768 | | 2.9632 | 1.0089 | H5 | 2.2232 | 2.5090 | 1.0089 | 2.9632 | | 3.0126 | H6 | 2.2232 | 2.5090 | 2.9632 | 1.0089 | 3.0126 | |
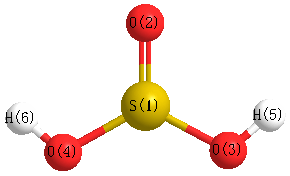 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
O3 |
H5 |
110.236 |
|
S1 |
O4 |
H6 |
110.236 |
| O2 |
S1 |
O3 |
105.571 |
|
O2 |
S1 |
O4 |
105.571 |
| O3 |
S1 |
O4 |
101.597 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.984 |
|
|
|
| 2 |
O |
-0.524 |
|
|
|
| 3 |
O |
-0.583 |
|
|
|
| 4 |
O |
-0.583 |
|
|
|
| 5 |
H |
0.353 |
|
|
|
| 6 |
H |
0.353 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.402 |
-1.523 |
0.000 |
2.070 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.342 |
-0.139 |
0.000 |
| y |
-0.139 |
3.307 |
0.000 |
| z |
0.000 |
0.000 |
3.845 |
<r2> (average value of r
2) Å
2
| <r2> |
81.138 |
| (<r2>)1/2 |
9.008 |
