return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(NH2)H2C(CH3)HCH3 (1-Propanamine, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-211.619396
Energy at 298.15K-211.632159
HF Energy-211.141807
Nuclear repulsion energy192.127633
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3534 3362 0.69      
2 A 3436 3268 2.11      
3 A 3161 3007 24.20      
4 A 3149 2996 55.37      
5 A 3144 2991 35.77      
6 A 3138 2985 9.05      
7 A 3113 2961 18.08      
8 A 3103 2951 8.64      
9 A 3070 2921 16.09      
10 A 3068 2919 20.69      
11 A 2986 2840 59.31      
12 A 1758 1673 19.73      
13 A 1617 1538 13.35      
14 A 1605 1526 3.93      
15 A 1598 1520 1.28      
16 A 1596 1518 1.62      
17 A 1589 1511 1.28      
18 A 1495 1422 10.14      
19 A 1473 1401 6.84      
20 A 1456 1385 3.29      
21 A 1439 1369 3.02      
22 A 1398 1330 0.79      
23 A 1370 1303 5.80      
24 A 1287 1224 0.42      
25 A 1240 1179 3.87      
26 A 1204 1146 4.45      
27 A 1111 1057 2.55      
28 A 1077 1024 13.22      
29 A 1010 961 1.32      
30 A 978 931 1.71      
31 A 949 903 4.72      
32 A 930 885 4.26      
33 A 812 772 77.75      
34 A 784 746 87.91      
35 A 489 466 8.51      
36 A 423 403 0.77      
37 A 372 354 1.05      
38 A 301 286 8.76      
39 A 270 257 2.44      
40 A 246 234 0.84      
41 A 213 203 51.74      
42 A 119 113 11.15      

Unscaled Zero Point Vibrational Energy (zpe) 33553.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 31919.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.24749 0.11389 0.08602

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -2.018 -0.026 -0.176
H2 -2.824 -0.650 -0.091
H3 -2.187 0.809 0.389
C4 -0.757 -0.712 0.246
H5 -0.622 -0.775 1.342
H6 -0.795 -1.738 -0.141
C7 1.767 -0.766 0.028
H8 1.893 -0.763 1.118
H9 1.723 -1.807 -0.315
H10 2.646 -0.289 -0.422
C11 0.551 1.470 0.107
H12 1.434 1.962 -0.316
H13 -0.339 2.015 -0.222
H14 0.620 1.515 1.202
C15 0.477 -0.004 -0.358
H16 0.349 -0.021 -1.447

Atom - Atom Distances (Å)
  N1 H2 H3 C4 H5 H6 C7 H8 H9 H10 C11 H12 H13 H14 C15 H16
N11.02321.02271.49692.19432.10423.86234.18454.14584.67772.98603.98622.64353.35162.50122.6867
H21.02321.66342.09592.63032.30314.59454.87064.69775.49193.99055.00093.64674.26893.37403.5078
H31.02271.66342.09362.42292.95104.27204.43274.75735.02222.83063.86512.28973.00672.88373.2393
C41.49692.09592.09361.10561.09682.53392.78932.76823.49312.54763.50282.79912.78751.54532.1373
H52.19432.63032.42291.10561.77642.72682.52463.05123.74512.81813.80413.21172.60942.16633.0481
H62.10422.30312.95101.09681.77642.74553.12372.52513.74373.48714.32313.78163.79292.16082.4414
C73.86234.59454.27202.53392.72682.74551.09721.09671.09602.54612.76983.49772.81001.54772.1775
H84.18454.87064.43272.78932.52463.12371.09721.78091.77832.79403.11333.80722.61052.18183.0841
H94.14584.69774.75732.76823.05122.52511.09671.78091.77893.50523.78004.34393.81462.19212.5214
H104.67775.49195.02223.49313.74513.74371.09601.77831.77892.78602.55913.77633.16142.18892.5295
C112.98603.99052.83062.54762.81813.48712.54612.79403.50522.78601.09611.09461.09801.54722.1630
H123.98625.00093.86513.50283.80414.32312.76983.11333.78002.55911.09611.77641.77922.18752.5282
H132.64353.64672.28972.79913.21173.78163.49773.80724.34393.77631.09461.77641.78832.18222.4743
H143.35164.26893.00672.78752.60943.79292.81002.61053.81463.16141.09801.77921.78832.18223.0741
C152.50123.37402.88371.54532.16632.16081.54772.18182.19212.18891.54722.18752.18222.18221.0964
H162.68673.50783.23932.13733.04812.44142.17753.08412.52142.52952.16302.52822.47433.07411.0964

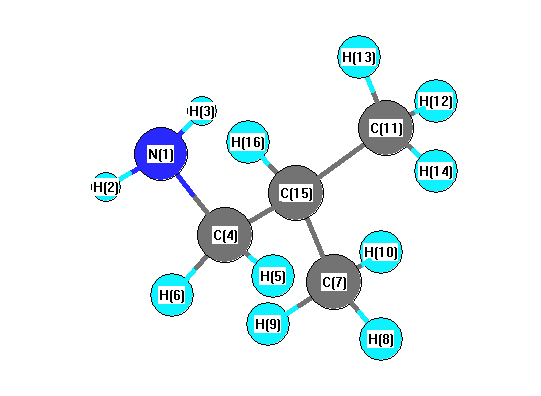
picture of 1-Propanamine, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C4 H5 114.105 N1 C4 H6 107.436
N1 C4 C15 110.594 H2 N1 H3 108.783
H2 N1 C4 111.143 H3 N1 C4 110.985
C4 C15 C7 110.015 C4 C15 C11 110.938
C4 C15 H16 106.775 H5 C4 H6 107.521
H5 C4 C15 108.464 H6 C4 C15 108.547
C7 C15 C11 110.712 C7 C15 H16 109.697
H8 C7 H9 108.531 H8 C7 H10 108.355
H8 C7 C15 109.987 H9 C7 H10 108.448
H9 C7 C15 110.831 H10 C7 C15 110.618
C11 C15 H16 108.608 H12 C11 H13 108.368
H12 C11 H14 108.369 H12 C11 C15 110.545
H13 C11 H14 109.288 H13 C11 C15 110.212
H14 C11 C15 110.013
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability