return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(CH3)CN (Propanenitrile, 2-methyl-)

using model chemistry: MP2=FULL/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/3-21G*
 hartrees
Energy at 0K-209.321022
Energy at 298.15K-209.328468
HF Energy-208.825943
Nuclear repulsion energy157.431341
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3175 3010 23.34      
2 A' 3169 3004 26.45      
3 A' 3101 2939 3.21      
4 A' 3090 2929 9.99      
5 A' 2136 2025 3.36      
6 A' 1607 1523 12.52      
7 A' 1601 1518 7.15      
8 A' 1502 1424 4.70      
9 A' 1416 1342 2.29      
10 A' 1244 1180 3.30      
11 A' 1191 1129 7.26      
12 A' 929 880 0.49      
13 A' 765 725 0.26      
14 A' 573 544 1.45      
15 A' 371 352 0.19      
16 A' 305 289 0.56      
17 A' 240 228 0.98      
18 A" 3172 3007 10.17      
19 A" 3165 3001 2.29      
20 A" 3089 2928 12.39      
21 A" 1591 1508 3.67      
22 A" 1589 1507 0.42      
23 A" 1484 1407 5.84      
24 A" 1385 1313 0.16      
25 A" 1155 1095 3.43      
26 A" 1008 956 1.38      
27 A" 983 932 1.67      
28 A" 596 565 0.57      
29 A" 250 237 0.02      
30 A" 210 199 2.61      

Unscaled Zero Point Vibrational Energy (zpe) 23046.0 cm-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 21847.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/3-21G*
ABC
0.26229 0.12852 0.09467

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.153 -2.245 0.000
C2 -0.107 -1.085 0.000
C3 -0.417 0.359 0.000
C4 0.153 1.033 1.280
C5 0.153 1.033 -1.280
H6 -1.509 0.468 0.000
H7 -0.112 2.095 1.277
H8 -0.256 0.567 2.181
H9 1.243 0.939 1.294
H10 -0.112 2.095 -1.277
H11 -0.256 0.567 -2.181
H12 1.243 0.939 -1.294

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.18872.66553.51873.51873.18194.53193.58183.60514.53193.58183.6051
C21.18871.47692.48822.48822.09263.42722.74012.75553.42722.74012.7555
C32.66551.47691.55501.55501.09762.17712.19712.18362.17712.19712.1836
C43.51872.48821.55502.56032.17281.09501.09411.09442.78223.51672.7973
C53.51872.48821.55502.56032.17282.78223.51672.79731.09501.09411.0944
H63.18192.09261.09762.17282.17282.49642.51753.07792.49642.51753.0779
H74.53193.42722.17711.09502.78222.49641.78181.78182.55493.78433.1286
H83.58182.74012.19711.09413.51672.51751.78181.78173.78434.36273.8036
H93.60512.75552.18361.09442.79733.07791.78181.78173.12863.80362.5884
H104.53193.42722.17712.78221.09502.49642.55493.78433.12861.78181.7818
H113.58182.74012.19713.51671.09412.51753.78434.36273.80361.78181.7817
H123.60512.75552.18362.79731.09443.07793.12863.80362.58841.78181.7817

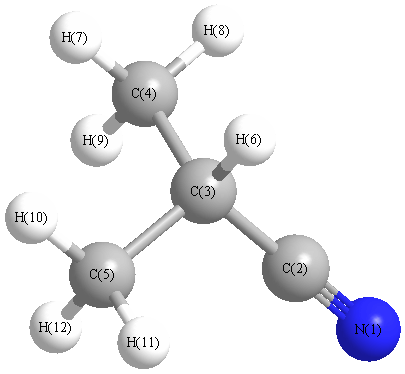
picture of Propanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 179.480 C2 C3 C4 110.278
C2 C3 C5 110.278 C2 C3 H6 107.833
C3 C4 H7 109.247 C3 C4 H8 110.870
C3 C4 H9 109.793 C3 C5 H10 109.247
C3 C5 H11 110.870 C3 C5 H12 109.793
C4 C3 C5 110.829 C4 C3 H6 108.773
C5 C3 H6 108.773 H7 C4 H8 108.963
H7 C4 H9 108.937 H8 C4 H9 108.999
H10 C5 H11 108.963 H10 C5 H12 108.937
H11 C5 H12 108.999
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability