return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

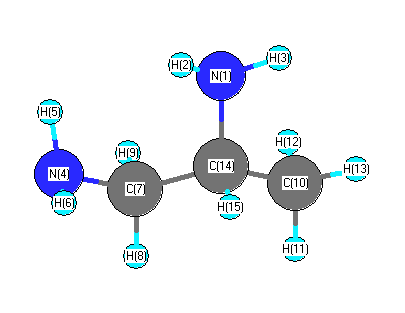
All results from a given calculation for C3H10N2 (1,2-Diaminopropane)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-227.526177
Energy at 298.15K-227.538699
HF Energy-227.035383
Nuclear repulsion energy193.335049
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3549 3376 0.29      
2 A 3514 3343 1.45      
3 A 3435 3268 1.59      
4 A 3415 3249 3.76      
5 A 3172 3017 25.28      
6 A 3151 2997 18.47      
7 A 3148 2995 25.66      
8 A 3088 2937 31.99      
9 A 3079 2929 14.82      
10 A 2972 2827 69.37      
11 A 1751 1666 2.24      
12 A 1744 1659 42.86      
13 A 1605 1527 4.71      
14 A 1592 1515 4.19      
15 A 1564 1488 1.17      
16 A 1484 1412 9.10      
17 A 1453 1382 3.95      
18 A 1448 1377 7.47      
19 A 1421 1351 5.64      
20 A 1397 1329 6.84      
21 A 1312 1248 0.98      
22 A 1252 1191 3.66      
23 A 1183 1125 0.33      
24 A 1100 1046 7.04      
25 A 1082 1029 5.61      
26 A 1042 991 3.33      
27 A 966 919 13.09      
28 A 916 871 11.36      
29 A 868 825 91.55      
30 A 798 759 102.84      
31 A 743 707 207.20      
32 A 497 473 2.88      
33 A 467 444 30.97      
34 A 385 366 62.65      
35 A 371 353 2.68      
36 A 262 250 4.64      
37 A 251 239 5.12      
38 A 225 214 78.92      
39 A 112 107 11.60      

Unscaled Zero Point Vibrational Energy (zpe) 30905.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 29400.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.26254 0.11462 0.08776

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.456 1.397 -0.208
H2 -0.364 1.909 0.129
H3 1.299 1.918 0.055
N4 -2.066 -0.138 0.024
H5 -2.168 0.736 -0.495
H6 -2.244 0.014 1.020
C7 -0.728 -0.753 -0.217
H8 -0.738 -1.770 0.191
H9 -0.585 -0.810 -1.301
C10 1.800 -0.658 -0.042
H11 1.822 -1.701 0.293
H12 1.884 -0.622 -1.133
H13 2.660 -0.135 0.394
C14 0.479 0.018 0.370
H15 0.394 0.002 1.473

Atom - Atom Distances (Å)
  N1 H2 H3 N4 H5 H6 C7 H8 H9 C10 H11 H12 H13 C14 H15
N11.02441.02512.96172.72183.27292.45483.40852.67362.46153.42272.64022.75071.49612.1857
H21.02441.66492.66362.24032.81352.70863.69843.07983.36234.22333.61303.65972.08472.4528
H31.02511.66493.94333.70434.13603.36434.21553.58182.62673.66452.86482.48672.09372.5495
N42.96172.66363.94331.02161.02311.49162.11112.09773.90164.19884.14414.73992.57302.8588
H52.72182.24033.70431.02161.67992.08992.96652.35544.23084.74164.32144.98602.87643.3136
H63.27292.81354.13601.02311.67992.10112.47742.96934.23474.47174.69854.94522.79922.6766
C72.45482.70863.36431.49162.08992.10111.09651.09452.53652.76832.77123.49781.54812.1655
H83.40853.69844.21552.11112.96652.47741.09651.78122.78062.56243.15343.77562.16992.4631
H92.67363.07983.58182.09772.35542.96931.09451.78122.70113.02122.48163.72222.14683.0520
C102.46153.36232.62673.90164.23084.23472.53652.78062.70111.09551.09431.09661.54032.1702
H113.42274.22333.66454.19884.74164.47172.76832.56243.02121.09551.78921.77852.18222.5161
H122.64023.61302.86484.14414.32144.69852.77123.15342.48161.09431.78921.78062.15443.0660
H132.75073.65972.48674.73994.98604.94523.49783.77563.72221.09661.77851.78062.18602.5129
C141.49612.08472.09372.57302.87642.79921.54812.16992.14681.54032.18222.15442.18601.1066
H152.18572.45282.54952.85883.31362.67662.16552.46313.05202.17022.51613.06602.51291.1066

picture of 1,2-Diaminopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C14 C7 107.472 N1 C14 C10 108.312
N1 C14 H15 113.392 H2 N1 H3 108.658
H2 N1 C14 110.199 H3 N1 C14 110.903
N4 C7 H8 108.342 N4 C7 H9 107.418
N4 C7 C14 115.645 H5 N4 H6 110.496
H5 N4 C7 111.135 H6 N4 C7 111.989
C7 C14 C10 110.433 C7 C14 H15 108.164
H8 C7 H9 108.770 H8 C7 C14 109.076
H9 C7 C14 107.411 C10 C14 H15 109.058
H11 C10 H12 109.580 H11 C10 H13 108.452
H11 C10 C14 110.639 H12 C10 H13 108.728
H12 C10 C14 108.537 H13 C10 C14 110.877
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability