Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -1066.254901 |
| Energy at 298.15K | -1066.257364 |
| HF Energy | -1065.677918 |
| Nuclear repulsion energy | 259.975894 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3213 |
3056 |
0.34 |
|
|
|
| 2 |
A' |
3040 |
2892 |
43.96 |
|
|
|
| 3 |
A' |
1628 |
1549 |
46.32 |
|
|
|
| 4 |
A' |
1454 |
1383 |
0.92 |
|
|
|
| 5 |
A' |
1256 |
1195 |
6.31 |
|
|
|
| 6 |
A' |
1003 |
954 |
34.24 |
|
|
|
| 7 |
A' |
777 |
739 |
16.38 |
|
|
|
| 8 |
A' |
425 |
404 |
3.51 |
|
|
|
| 9 |
A' |
319 |
304 |
22.34 |
|
|
|
| 10 |
A' |
262 |
249 |
1.51 |
|
|
|
| 11 |
A" |
1302 |
1238 |
20.29 |
|
|
|
| 12 |
A" |
1044 |
993 |
22.20 |
|
|
|
| 13 |
A" |
711 |
676 |
101.48 |
|
|
|
| 14 |
A" |
287 |
273 |
1.98 |
|
|
|
| 15 |
A" |
80 |
76 |
7.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8400.2 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 7991.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.340 |
0.210 |
0.000 |
| C2 |
-0.938 |
1.068 |
0.000 |
| H3 |
1.225 |
0.841 |
0.000 |
| Cl4 |
0.340 |
-0.807 |
1.483 |
| Cl5 |
0.340 |
-0.807 |
-1.483 |
| O6 |
-0.921 |
2.311 |
0.000 |
| H7 |
-1.852 |
0.458 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5395 | 1.0871 | 1.7982 | 1.7982 | 2.4502 | 2.2068 |
C2 | 1.5395 | | 2.1750 | 2.7108 | 2.7108 | 1.2430 | 1.0990 | H3 | 1.0871 | 2.1750 | | 2.3874 | 2.3874 | 2.6007 | 3.1014 | Cl4 | 1.7982 | 2.7108 | 2.3874 | | 2.9657 | 3.6756 | 2.9341 | Cl5 | 1.7982 | 2.7108 | 2.3874 | 2.9657 | | 3.6756 | 2.9341 | O6 | 2.4502 | 1.2430 | 2.6007 | 3.6756 | 3.6756 | | 2.0735 | H7 | 2.2068 | 1.0990 | 3.1014 | 2.9341 | 2.9341 | 2.0735 | |
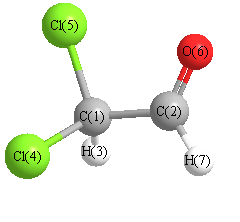 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
123.068 |
|
C1 |
C2 |
H7 |
112.455 |
| C2 |
C1 |
H3 |
110.621 |
|
C2 |
C1 |
Cl4 |
108.373 |
| C2 |
C1 |
Cl5 |
108.373 |
|
H3 |
C1 |
Cl4 |
109.183 |
| H3 |
C1 |
Cl5 |
109.183 |
|
Cl4 |
C1 |
Cl5 |
111.104 |
| O6 |
C2 |
H7 |
124.477 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -1066.255053 |
| Energy at 298.15K | |
| HF Energy | -1065.677113 |
| Nuclear repulsion energy | 263.265328 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3172 |
3018 |
2.38 |
70.93 |
0.27 |
0.42 |
| 2 |
A |
3027 |
2880 |
49.33 |
116.60 |
0.30 |
0.46 |
| 3 |
A |
1630 |
1550 |
37.10 |
1.78 |
0.73 |
0.84 |
| 4 |
A |
1447 |
1376 |
6.65 |
7.28 |
0.71 |
0.83 |
| 5 |
A |
1298 |
1235 |
12.21 |
6.22 |
0.72 |
0.84 |
| 6 |
A |
1289 |
1226 |
12.22 |
11.55 |
0.74 |
0.85 |
| 7 |
A |
1037 |
987 |
11.75 |
5.41 |
0.51 |
0.68 |
| 8 |
A |
932 |
887 |
19.16 |
3.71 |
0.75 |
0.86 |
| 9 |
A |
800 |
761 |
58.92 |
6.39 |
0.74 |
0.85 |
| 10 |
A |
641 |
610 |
32.67 |
9.67 |
0.10 |
0.19 |
| 11 |
A |
612 |
582 |
34.70 |
7.97 |
0.49 |
0.66 |
| 12 |
A |
352 |
335 |
1.78 |
3.62 |
0.25 |
0.40 |
| 13 |
A |
277 |
263 |
2.87 |
5.71 |
0.64 |
0.78 |
| 14 |
A |
228 |
217 |
2.81 |
1.27 |
0.69 |
0.82 |
| 15 |
A |
106 |
101 |
8.85 |
2.14 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 8423.0 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 8012.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.114 |
-0.032 |
0.540 |
| C2 |
-0.725 |
-1.283 |
0.195 |
| H3 |
0.245 |
0.055 |
1.619 |
| Cl4 |
1.758 |
-0.268 |
-0.174 |
| Cl5 |
-0.673 |
1.448 |
-0.058 |
| O6 |
-1.854 |
-1.251 |
-0.318 |
| H7 |
-0.200 |
-2.211 |
0.464 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5454 | 1.0904 | 1.8080 | 1.7796 | 2.4691 | 2.2028 |
C2 | 1.5454 | | 2.1818 | 2.7080 | 2.7432 | 1.2405 | 1.1000 | H3 | 1.0904 | 2.1818 | | 2.3682 | 2.3653 | 3.1410 | 2.5820 | Cl4 | 1.8080 | 2.7080 | 2.3682 | | 2.9780 | 3.7467 | 2.8312 | Cl5 | 1.7796 | 2.7432 | 2.3653 | 2.9780 | | 2.9579 | 3.7263 | O6 | 2.4691 | 1.2405 | 3.1410 | 3.7467 | 2.9579 | | 2.0667 | H7 | 2.2028 | 1.1000 | 2.5820 | 2.8312 | 3.7263 | 2.0667 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.458 |
|
C1 |
C2 |
H7 |
111.646 |
| C2 |
C1 |
H3 |
110.549 |
|
C2 |
C1 |
Cl4 |
107.450 |
| C2 |
C1 |
Cl5 |
110.986 |
|
H3 |
C1 |
Cl4 |
106.959 |
| H3 |
C1 |
Cl5 |
108.616 |
|
Cl4 |
C1 |
Cl5 |
112.212 |
| O6 |
C2 |
H7 |
123.895 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
