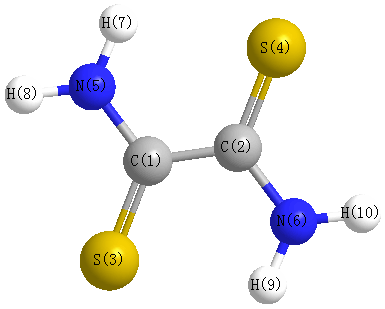
Vibrational Frequencies calculated at B3PW91/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3625 |
3477 |
0.00 |
|
|
|
| 2 |
Ag |
3387 |
3249 |
0.00 |
|
|
|
| 3 |
Ag |
1651 |
1584 |
0.00 |
|
|
|
| 4 |
Ag |
1465 |
1406 |
0.00 |
|
|
|
| 5 |
Ag |
1328 |
1274 |
0.00 |
|
|
|
| 6 |
Ag |
933 |
895 |
0.00 |
|
|
|
| 7 |
Ag |
669 |
642 |
0.00 |
|
|
|
| 8 |
Ag |
421 |
404 |
0.00 |
|
|
|
| 9 |
Ag |
342 |
328 |
0.00 |
|
|
|
| 10 |
Au |
758 |
728 |
501.81 |
|
|
|
| 11 |
Au |
662 |
635 |
32.97 |
|
|
|
| 12 |
Au |
405 |
389 |
32.41 |
|
|
|
| 13 |
Au |
70 |
67 |
12.74 |
|
|
|
| 14 |
Bg |
788 |
756 |
0.00 |
|
|
|
| 15 |
Bg |
677 |
650 |
0.00 |
|
|
|
| 16 |
Bg |
663 |
636 |
0.00 |
|
|
|
| 17 |
Bu |
3626 |
3479 |
196.55 |
|
|
|
| 18 |
Bu |
3394 |
3257 |
239.66 |
|
|
|
| 19 |
Bu |
1627 |
1561 |
470.73 |
|
|
|
| 20 |
Bu |
1458 |
1399 |
281.12 |
|
|
|
| 21 |
Bu |
1275 |
1223 |
123.36 |
|
|
|
| 22 |
Bu |
881 |
845 |
55.26 |
|
|
|
| 23 |
Bu |
463 |
444 |
2.93 |
|
|
|
| 24 |
Bu |
294 |
282 |
29.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15430.8 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 14804.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.120 |
|
|
|
| 2 |
C |
0.120 |
|
|
|
| 3 |
S |
-0.115 |
|
|
|
| 4 |
S |
-0.115 |
|
|
|
| 5 |
N |
-0.748 |
|
|
|
| 6 |
N |
-0.748 |
|
|
|
| 7 |
H |
0.377 |
|
|
|
| 8 |
H |
0.366 |
|
|
|
| 9 |
H |
0.377 |
|
|
|
| 10 |
H |
0.366 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.120 |
-12.868 |
0.000 |
| y |
-12.868 |
-50.575 |
0.000 |
| z |
0.000 |
0.000 |
-52.592 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.463 |
-12.868 |
0.000 |
| y |
-12.868 |
-2.219 |
0.000 |
| z |
0.000 |
0.000 |
-5.244 |
|
| Polar |
|---|
| 3z2-r2 | -10.489 |
|---|
| x2-y2 | 6.455 |
|---|
| xy | -12.868 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.296 |
3.148 |
0.000 |
| y |
3.148 |
11.604 |
0.000 |
| z |
0.000 |
0.000 |
3.666 |
<r2> (average value of r
2) Å
2
| <r2> |
256.249 |
| (<r2>)1/2 |
16.008 |