Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -215.122333 |
| Energy at 298.15K | |
| HF Energy | -214.737180 |
| Nuclear repulsion energy | 115.662886 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3295 |
3134 |
4.91 |
39.92 |
0.72 |
0.83 |
| 2 |
A' |
3212 |
3055 |
5.98 |
124.40 |
0.14 |
0.24 |
| 3 |
A' |
3195 |
3039 |
5.15 |
24.79 |
0.70 |
0.82 |
| 4 |
A' |
3083 |
2932 |
32.87 |
114.28 |
0.12 |
0.21 |
| 5 |
A' |
1704 |
1621 |
1.88 |
2.86 |
0.43 |
0.60 |
| 6 |
A' |
1616 |
1538 |
0.61 |
20.72 |
0.67 |
0.80 |
| 7 |
A' |
1519 |
1445 |
5.20 |
7.89 |
0.56 |
0.72 |
| 8 |
A' |
1457 |
1386 |
9.91 |
10.38 |
0.59 |
0.75 |
| 9 |
A' |
1368 |
1301 |
0.10 |
12.23 |
0.45 |
0.62 |
| 10 |
A' |
1146 |
1090 |
12.51 |
2.93 |
0.71 |
0.83 |
| 11 |
A' |
1040 |
990 |
30.73 |
4.90 |
0.71 |
0.83 |
| 12 |
A' |
904 |
860 |
4.15 |
4.95 |
0.17 |
0.29 |
| 13 |
A' |
613 |
583 |
2.39 |
1.48 |
0.75 |
0.86 |
| 14 |
A' |
308 |
293 |
5.26 |
0.59 |
0.50 |
0.66 |
| 15 |
A" |
3118 |
2966 |
39.76 |
79.05 |
0.75 |
0.86 |
| 16 |
A" |
1289 |
1227 |
0.07 |
14.44 |
0.75 |
0.86 |
| 17 |
A" |
1107 |
1053 |
6.05 |
0.49 |
0.75 |
0.86 |
| 18 |
A" |
1051 |
1000 |
27.28 |
0.01 |
0.75 |
0.86 |
| 19 |
A" |
983 |
935 |
40.83 |
0.11 |
0.75 |
0.86 |
| 20 |
A" |
588 |
560 |
12.00 |
14.27 |
0.75 |
0.86 |
| 21 |
A" |
189 |
180 |
3.81 |
5.98 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16391.1 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 15592.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
-0.986 |
0.000 |
| C2 |
-0.971 |
0.176 |
0.000 |
| C3 |
-0.542 |
1.444 |
0.000 |
| F4 |
1.344 |
-0.494 |
0.000 |
| H5 |
-1.225 |
2.285 |
0.000 |
| H6 |
0.521 |
1.657 |
0.000 |
| H7 |
-2.027 |
-0.082 |
0.000 |
| H8 |
-0.147 |
-1.610 |
0.889 |
| H9 |
-0.147 |
-1.610 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5138 | 2.4889 | 1.4314 | 3.4928 | 2.6930 | 2.2188 | 1.0960 | 1.0960 |
C2 | 1.5138 | | 1.3381 | 2.4097 | 2.1246 | 2.1014 | 1.0872 | 2.1584 | 2.1584 | C3 | 2.4889 | 1.3381 | | 2.7037 | 1.0839 | 1.0837 | 2.1292 | 3.2047 | 3.2047 | F4 | 1.4314 | 2.4097 | 2.7037 | | 3.7844 | 2.3025 | 3.3958 | 2.0636 | 2.0636 | H5 | 3.4928 | 2.1246 | 1.0839 | 3.7844 | | 1.8552 | 2.4999 | 4.1383 | 4.1383 | H6 | 2.6930 | 2.1014 | 1.0837 | 2.3025 | 1.8552 | | 3.0843 | 3.4504 | 3.4504 | H7 | 2.2188 | 1.0872 | 2.1292 | 3.3958 | 2.4999 | 3.0843 | | 2.5803 | 2.5803 | H8 | 1.0960 | 2.1584 | 3.2047 | 2.0636 | 4.1383 | 3.4504 | 2.5803 | | 1.7777 | H9 | 1.0960 | 2.1584 | 3.2047 | 2.0636 | 4.1383 | 3.4504 | 2.5803 | 1.7777 | |
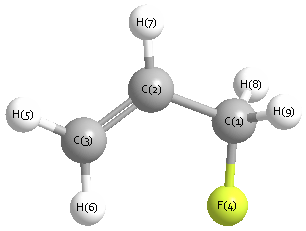 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.434 |
|
C1 |
C2 |
H7 |
116.123 |
| C2 |
C1 |
F4 |
109.773 |
|
C2 |
C1 |
H8 |
110.570 |
| C2 |
C1 |
H9 |
110.570 |
|
C2 |
C3 |
H5 |
122.262 |
| C2 |
C3 |
H6 |
120.023 |
|
C3 |
C2 |
H7 |
122.443 |
| F4 |
C1 |
H8 |
108.748 |
|
F4 |
C1 |
H9 |
108.748 |
| H5 |
C3 |
H6 |
117.715 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -215.118507 |
| Energy at 298.15K | |
| HF Energy | -214.733733 |
| Nuclear repulsion energy | 112.533225 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3264 |
3105 |
12.67 |
50.62 |
0.52 |
0.68 |
| 2 |
A |
3209 |
3053 |
3.36 |
79.21 |
0.32 |
0.48 |
| 3 |
A |
3173 |
3019 |
7.08 |
61.29 |
0.11 |
0.20 |
| 4 |
A |
3128 |
2976 |
35.40 |
59.05 |
0.75 |
0.86 |
| 5 |
A |
3084 |
2934 |
29.82 |
90.31 |
0.11 |
0.19 |
| 6 |
A |
1698 |
1616 |
0.54 |
2.68 |
0.38 |
0.55 |
| 7 |
A |
1613 |
1535 |
0.24 |
14.29 |
0.73 |
0.84 |
| 8 |
A |
1536 |
1462 |
9.50 |
8.12 |
0.48 |
0.65 |
| 9 |
A |
1456 |
1385 |
18.40 |
6.24 |
0.73 |
0.85 |
| 10 |
A |
1367 |
1300 |
0.04 |
9.59 |
0.49 |
0.65 |
| 11 |
A |
1298 |
1234 |
1.80 |
18.16 |
0.74 |
0.85 |
| 12 |
A |
1204 |
1145 |
0.99 |
2.31 |
0.69 |
0.82 |
| 13 |
A |
1073 |
1021 |
40.92 |
6.00 |
0.54 |
0.71 |
| 14 |
A |
1060 |
1008 |
30.28 |
1.17 |
0.56 |
0.72 |
| 15 |
A |
1038 |
988 |
7.50 |
1.08 |
0.75 |
0.86 |
| 16 |
A |
977 |
929 |
46.22 |
0.10 |
0.53 |
0.69 |
| 17 |
A |
937 |
892 |
8.57 |
3.59 |
0.14 |
0.24 |
| 18 |
A |
635 |
604 |
9.52 |
10.01 |
0.73 |
0.84 |
| 19 |
A |
456 |
433 |
1.93 |
4.54 |
0.48 |
0.65 |
| 20 |
A |
302 |
288 |
6.15 |
3.56 |
0.75 |
0.86 |
| 21 |
A |
95 |
91 |
1.80 |
6.72 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 16302.2 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 15508.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.590 |
0.489 |
0.233 |
| C2 |
0.644 |
-0.382 |
0.197 |
| C3 |
1.855 |
0.060 |
-0.170 |
| F4 |
-1.716 |
-0.254 |
-0.257 |
| H5 |
2.732 |
-0.578 |
-0.141 |
| H6 |
2.011 |
1.078 |
-0.520 |
| H7 |
0.499 |
-1.402 |
0.541 |
| H8 |
-0.811 |
0.800 |
1.260 |
| H9 |
-0.446 |
1.383 |
-0.383 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5109 | 2.5150 | 1.4351 | 3.5092 | 2.7714 | 2.2041 | 1.0955 | 1.0953 |
C2 | 1.5109 | | 1.3402 | 2.4069 | 2.1240 | 2.1245 | 1.0867 | 2.1554 | 2.1541 | C3 | 2.5150 | 1.3402 | | 3.5860 | 1.0849 | 1.0878 | 2.1171 | 3.1149 | 2.6632 | F4 | 1.4351 | 2.4069 | 3.5860 | | 4.4613 | 3.9669 | 2.6198 | 2.0565 | 2.0755 | H5 | 3.5092 | 2.1240 | 1.0849 | 4.4613 | | 1.8457 | 2.4756 | 4.0519 | 3.7426 | H6 | 2.7714 | 2.1245 | 1.0878 | 3.9669 | 1.8457 | | 3.0925 | 3.3483 | 2.4803 | H7 | 2.2041 | 1.0867 | 2.1171 | 2.6198 | 2.4756 | 3.0925 | | 2.6619 | 3.0833 | H8 | 1.0955 | 2.1554 | 3.1149 | 2.0565 | 4.0519 | 3.3483 | 2.6619 | | 1.7810 | H9 | 1.0953 | 2.1541 | 2.6632 | 2.0755 | 3.7426 | 2.4803 | 3.0833 | 1.7810 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.692 |
|
C1 |
C2 |
H7 |
115.132 |
| C2 |
C1 |
F4 |
109.545 |
|
C2 |
C1 |
H8 |
110.565 |
| C2 |
C1 |
H9 |
110.478 |
|
C2 |
C3 |
H5 |
121.935 |
| C2 |
C3 |
H6 |
121.750 |
|
C3 |
C2 |
H7 |
121.111 |
| F4 |
C1 |
H8 |
107.958 |
|
F4 |
C1 |
H9 |
109.480 |
| H5 |
C3 |
H6 |
116.314 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
