return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 ((E)-hexa-1,3,5-triene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1Ag
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-231.046075
Energy at 298.15K-231.053157
HF Energy-230.514447
Nuclear repulsion energy192.516074
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3260 3101 0.00      
2 Ag 3172 3017 0.00      
3 Ag 3155 3001 0.00      
4 Ag 3136 2983 0.00      
5 Ag 1682 1600 0.00      
6 Ag 1635 1555 0.00      
7 Ag 1499 1426 0.00      
8 Ag 1390 1323 0.00      
9 Ag 1358 1292 0.00      
10 Ag 1240 1179 0.00      
11 Ag 974 926 0.00      
12 Ag 451 429 0.00      
13 Ag 363 345 0.00      
14 Au 1050 999 86.47      
15 Au 974 926 7.95      
16 Au 938 892 86.92      
17 Au 708 674 11.68      
18 Au 243 231 2.84      
19 Au 90 85 0.31      
20 Bg 1028 978 0.00      
21 Bg 938 892 0.00      
22 Bg 859 817 0.00      
23 Bg 605 575 0.00      
24 Bg 188 179 0.00      
25 Bu 3260 3101 27.09      
26 Bu 3171 3017 7.54      
27 Bu 3156 3003 38.47      
28 Bu 3140 2987 6.35      
29 Bu 1673 1591 6.42      
30 Bu 1538 1463 1.74      
31 Bu 1375 1308 2.75      
32 Bu 1351 1286 0.40      
33 Bu 1167 1110 7.53      
34 Bu 990 942 1.14      
35 Bu 568 540 2.26      
36 Bu 152 145 1.03      

Unscaled Zero Point Vibrational Energy (zpe) 26237.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 24959.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.87351 0.04388 0.04178

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.678 0.000
C2 0.000 -0.678 0.000
C3 1.218 1.498 0.000
C4 -1.218 -1.498 0.000
C5 1.217 2.849 0.000
C6 -1.217 -2.849 0.000
H7 0.953 -1.213 0.000
H8 -0.953 1.213 0.000
H9 2.168 0.961 0.000
H10 -2.168 -0.961 0.000
H11 2.138 3.422 0.000
H12 -2.138 -3.422 0.000
H13 -0.290 -3.417 0.000
H14 0.290 3.417 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.35651.46812.49412.48833.73102.11781.09292.18602.71783.47884.62474.10562.7541
C21.35652.49411.46813.73102.48831.09292.11782.71782.18604.62473.47882.75414.1056
C31.46812.49413.86161.35024.98252.72412.18961.09134.18452.13295.95625.14142.1314
C42.49411.46813.86164.98251.35022.18962.72414.18451.09135.95622.13292.13145.1414
C52.48833.73101.35024.98256.19554.07002.71782.11335.09611.08537.11226.44431.0879
C63.73102.48834.98251.35026.19552.71784.07005.09612.11337.11221.08531.08796.4443
H72.11781.09292.72412.18964.07002.71783.08522.49023.13094.78433.80002.53044.6772
H81.09292.11782.18962.72412.71784.07003.08523.13092.49023.80004.78434.67722.5304
H92.18602.71781.09134.18452.11335.09612.49023.13094.74232.46136.14465.02063.0916
H102.71782.18604.18451.09135.09612.11333.13092.49024.74236.14462.46133.09165.0206
H113.47884.62472.13295.95621.08537.11224.78433.80002.46136.14468.07097.25761.8488
H124.62473.47885.95622.13297.11221.08533.80004.78436.14462.46138.07091.84887.2576
H134.10562.75415.14142.13146.44431.08792.53044.67725.02063.09167.25761.84886.8586
H142.75414.10562.13145.14141.08796.44434.67722.53043.09165.02061.84887.25766.8586

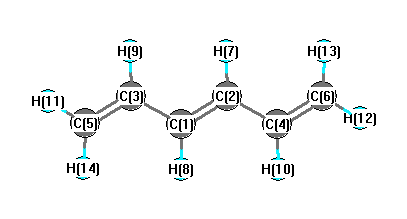
picture of (E)-hexa-1,3,5-triene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 123.961 C1 C2 H7 119.283
C1 C3 C5 123.935 C1 C3 H9 116.548
C2 C1 C3 123.961 C2 C1 H8 119.283
C2 C4 C6 123.935 C2 C4 H10 116.548
C3 C1 H8 116.756 C3 C5 H11 121.887
C3 C5 H14 121.527 C4 C2 H7 116.756
C4 C6 H12 121.887 C4 C6 H13 121.527
C5 C3 H9 119.517 C6 C4 H10 119.517
H11 C5 H14 116.586 H12 C6 H13 116.586
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability