return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O (allenol)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-190.032225
Energy at 298.15K-190.035601
HF Energy-189.647877
Nuclear repulsion energy100.484784
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3514 3343 12.63      
2 A' 3247 3089 4.61      
3 A' 3145 2992 10.97      
4 A' 2033 1934 24.01      
5 A' 1572 1495 4.17      
6 A' 1446 1376 40.97      
7 A' 1336 1271 6.37      
8 A' 1189 1131 87.87      
9 A' 982 934 55.18      
10 A' 936 890 118.31      
11 A' 614 584 22.15      
12 A' 185 176 0.50      
13 A" 3213 3057 9.21      
14 A" 1118 1064 0.08      
15 A" 960 914 30.64      
16 A" 644 612 0.09      
17 A" 421 401 164.22      
18 A" 218 208 5.38      

Unscaled Zero Point Vibrational Energy (zpe) 13385.7 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12733.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
1.40553 0.14108 0.13165

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.677 -0.462 0.000
C2 0.000 0.667 0.000
C3 -0.701 1.783 0.000
O4 0.153 -1.782 0.000
H5 1.760 -0.499 0.000
H6 -1.002 2.277 0.922
H7 -1.002 2.277 -0.922
H8 -0.839 -1.724 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8
C11.31692.63461.42001.08283.34293.34291.9728
C21.31691.31782.45372.11082.10872.10872.5338
C32.63461.31783.66583.35561.08871.08873.5097
O41.42002.45373.66582.05624.31954.31950.9939
H51.08282.11083.35562.05624.02274.02272.8732
H63.34292.10871.08874.31954.02271.84464.1089
H73.34292.10871.08874.31954.02271.84464.1089
H81.97282.53383.50970.99392.87324.10894.1089

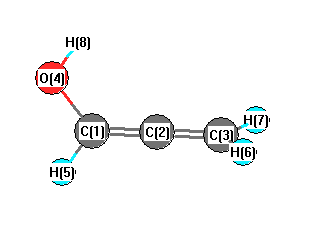
picture of allenol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 178.843 C1 O4 H8 108.327
C2 C1 O4 127.365 C2 C1 H5 122.887
C2 C3 H6 122.095 C2 C3 H7 122.095
O4 C1 H5 109.749 H6 C3 H7 115.805
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2/3-21G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.079      
2 C -0.143      
3 C -0.410      
4 O -0.651      
5 H 0.263      
6 H 0.236      
7 H 0.236      
8 H 0.392      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -20.960 0.784 0.000
y 0.784 -25.502 0.000
z 0.000 0.000 -24.822
Traceless
 xyz
x 4.201 0.784 0.000
y 0.784 -2.611 0.000
z 0.000 0.000 -1.590
Polar
3z2-r2-3.181
x2-y24.542
xy0.784
xz0.000
yz0.000


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000