return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4N2 (2H-Imidazole)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-223.905729
Energy at 298.15K-223.911741
HF Energy-223.434112
Nuclear repulsion energy164.751563
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3289 3129 8.13      
2 A1 3129 2976 0.46      
3 A1 1634 1554 28.79      
4 A1 1508 1434 7.48      
5 A1 1360 1294 0.21      
6 A1 1240 1180 5.86      
7 A1 1023 973 17.02      
8 A1 879 836 5.31      
9 A2 1192 1134 0.00      
10 A2 883 840 0.00      
11 A2 571 543 0.00      
12 B1 3181 3026 0.04      
13 B1 1048 997 18.42      
14 B1 833 792 12.99      
15 B1 368 350 44.17      
16 B2 3273 3114 10.58      
17 B2 1667 1586 0.59      
18 B2 1338 1273 7.95      
19 B2 1284 1221 3.13      
20 B2 1066 1014 26.68      
21 B2 978 930 84.57      

Unscaled Zero Point Vibrational Energy (zpe) 15870.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 15098.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.35668 0.29395 0.16636

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.230
N2 0.000 0.999 0.284
N3 0.000 -0.999 0.284
C4 0.000 0.744 -0.963
C5 0.000 -0.744 -0.963
H6 -0.902 0.000 1.845
H7 0.902 0.000 1.845
H8 0.000 1.481 -1.750
H9 0.000 -1.481 -1.750

Atom - Atom Distances (Å)
  C1 N2 N3 C4 C5 H6 H7 H8 H9
C11.37541.37542.31572.31571.09191.09193.32813.3281
N21.37541.99711.27282.14272.06092.06092.09113.2073
N31.37541.99712.14271.27282.06092.06093.20732.0911
C42.31571.27282.14271.48773.04173.04171.07862.3598
C52.31572.14271.27281.48773.04173.04172.35981.0786
H61.09192.06092.06093.04173.04171.80443.99193.9919
H71.09192.06092.06093.04173.04171.80443.99193.9919
H83.32812.09113.20731.07862.35983.99193.99192.9613
H93.32813.20732.09112.35981.07863.99193.99192.9613

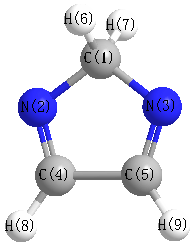
picture of 2H-Imidazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 C4 121.905 C1 N3 C5 121.905
N2 C1 N3 93.105 N2 C1 H6 112.789
N2 C1 H7 112.789 N2 C4 C5 101.543
N2 C4 H8 125.370 N3 C1 H6 112.789
N3 C1 H7 112.789 N3 C5 C4 101.543
N3 C5 H9 125.370 C4 C5 H9 133.087
C5 C4 H8 133.087 H6 C1 H7 111.438
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability