Jump to
S1C2
Vibrational Frequencies calculated at MP2/3-21G*
Geometric Data calculated at MP2/3-21G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -114.775646 |
| Energy at 298.15K | -114.777416 |
| HF Energy | -114.526109 |
| Nuclear repulsion energy | 56.996290 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3362 |
3198 |
0.57 |
|
|
|
| 2 |
A' |
3171 |
3017 |
37.93 |
|
|
|
| 3 |
A' |
1569 |
1492 |
0.15 |
|
|
|
| 4 |
A' |
1181 |
1124 |
15.06 |
|
|
|
| 5 |
A' |
1054 |
1003 |
5.82 |
|
|
|
| 6 |
A' |
957 |
910 |
2.83 |
|
|
|
| 7 |
A' |
629 |
598 |
96.03 |
|
|
|
| 8 |
A" |
5836 |
5552 |
206923.90 |
|
|
|
| 9 |
A" |
3093 |
2942 |
3440.40 |
|
|
|
| 10 |
A" |
1086 |
1034 |
7.33 |
|
|
|
| 11 |
A" |
1030 |
979 |
12.82 |
|
|
|
| 12 |
A" |
870 |
828 |
0.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11918.4 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 11338.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.038 |
0.919 |
0.000 |
| H2 |
0.761 |
1.656 |
0.000 |
| C3 |
-0.038 |
-0.431 |
0.666 |
| C4 |
-0.038 |
-0.431 |
-0.666 |
| H5 |
-0.040 |
-1.002 |
1.573 |
| H6 |
-0.040 |
-1.002 |
-1.573 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
| C1 | | 1.0866 | 1.5050 | 1.5050 | 2.4832 | 2.4832 |
H2 | 1.0866 | | 2.3315 | 2.3315 | 3.1911 | 3.1911 | C3 | 1.5050 | 2.3315 | | 1.3314 | 1.0726 | 2.3108 | C4 | 1.5050 | 2.3315 | 1.3314 | | 2.3108 | 1.0726 | H5 | 2.4832 | 3.1911 | 1.0726 | 2.3108 | | 3.1469 | H6 | 2.4832 | 3.1911 | 2.3108 | 1.0726 | 3.1469 | |
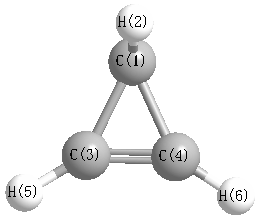 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C4 |
63.749 |
|
C1 |
C3 |
H5 |
148.434 |
| C1 |
C4 |
C3 |
63.749 |
|
C1 |
C4 |
H6 |
148.434 |
| H2 |
C1 |
C3 |
127.475 |
|
H2 |
C1 |
C4 |
127.475 |
| C3 |
C1 |
C4 |
52.502 |
|
C3 |
C4 |
H6 |
147.816 |
| C4 |
C3 |
H5 |
147.816 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
