return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HOCHCCHOH (allenediol)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-264.589790
Energy at 298.15K-264.593852
HF Energy-264.085069
Nuclear repulsion energy149.792430
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3504 3333 7.39      
2 A 3223 3066 8.05      
3 A 1494 1422 15.67      
4 A 1342 1276 0.96      
5 A 1239 1179 29.56      
6 A 997 949 32.97      
7 A 900 856 81.94      
8 A 546 520 94.03      
9 A 510 485 83.99      
10 A 344 327 14.48      
11 A 102 97 0.00      
12 B 3503 3332 20.26      
13 B 3224 3067 19.07      
14 B 2028 1930 228.48      
15 B 1437 1367 51.83      
16 B 1331 1266 15.62      
17 B 1023 973 410.05      
18 B 919 875 11.55      
19 B 625 594 53.55      
20 B 499 475 153.12      
21 B 128 122 1.56      

Unscaled Zero Point Vibrational Energy (zpe) 14458.4 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 13754.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.70661 0.07392 0.07169

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.410
C2 0.000 1.317 0.375
C3 0.000 -1.317 0.375
O4 0.799 2.156 -0.436
O5 -0.799 -2.156 -0.436
H6 -0.632 1.936 1.001
H7 0.632 -1.936 1.001
H8 1.391 1.581 -0.993
H9 -1.391 -1.581 -0.993

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.31711.31712.45022.45022.12082.12082.53032.5303
C21.31712.63331.41453.65461.08463.37231.96813.4932
C31.31712.63333.65461.41453.37231.08463.49321.9681
O42.45021.41453.65464.59902.04084.34040.99494.3675
O52.45023.65461.41454.59904.34042.04084.36750.9949
H62.12081.08463.37232.04084.34044.07342.86274.1138
H72.12083.37231.08464.34042.04084.07344.11382.8627
H82.53031.96813.49320.99494.36752.86274.11384.2116
H92.53033.49321.96814.36750.99494.11382.86274.2116

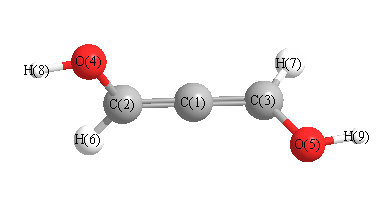
picture of allenediol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 127.490 C1 C2 H6 123.730
C1 C3 O5 127.490 C1 C3 H7 123.730
C2 C1 C3 176.922 C2 O4 H8 108.287
C3 O5 H9 108.287 O4 C2 H6 108.775
O5 C3 H7 108.775
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability