Jump to
S1C2
S1C3
Energy calculated at CISD/3-21G*
| | hartrees |
|---|
| Energy at 0K | -168.245973 |
| Energy at 298.15K | -168.249848 |
| HF Energy | -167.940420 |
| Nuclear repulsion energy | 69.786123 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3656 |
3416 |
22.42 |
|
|
|
| 2 |
A' |
3461 |
3234 |
0.89 |
|
|
|
| 3 |
A' |
3086 |
2883 |
57.84 |
|
|
|
| 4 |
A' |
1807 |
1689 |
105.40 |
|
|
|
| 5 |
A' |
1491 |
1393 |
10.12 |
|
|
|
| 6 |
A' |
1335 |
1247 |
129.23 |
|
|
|
| 7 |
A' |
1212 |
1133 |
82.84 |
|
|
|
| 8 |
A' |
1027 |
960 |
268.61 |
|
|
|
| 9 |
A' |
627 |
586 |
1.25 |
|
|
|
| 10 |
A" |
1111 |
1038 |
7.62 |
|
|
|
| 11 |
A" |
860 |
804 |
64.12 |
|
|
|
| 12 |
A" |
277 |
259 |
100.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9975.6 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 9321.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CISD/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.392 |
0.000 |
| O2 |
-1.009 |
-0.566 |
0.000 |
| N3 |
1.206 |
0.015 |
0.000 |
| H4 |
-0.362 |
1.424 |
0.000 |
| H5 |
-1.890 |
-0.132 |
0.000 |
| H6 |
1.881 |
0.785 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3917 | 1.2639 | 1.0934 | 1.9613 | 1.9220 |
O2 | 1.3917 | | 2.2901 | 2.0927 | 0.9819 | 3.1908 | N3 | 1.2639 | 2.2901 | | 2.1084 | 3.0994 | 1.0245 | H4 | 1.0934 | 2.0927 | 2.1084 | | 2.1808 | 2.3328 | H5 | 1.9613 | 0.9819 | 3.0994 | 2.1808 | | 3.8813 | H6 | 1.9220 | 3.1908 | 1.0245 | 2.3328 | 3.8813 | |
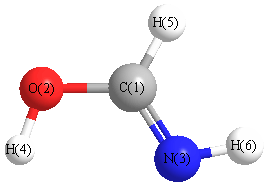 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
110.243 |
|
C1 |
N3 |
H6 |
113.856 |
| O2 |
C1 |
N3 |
119.084 |
|
O2 |
C1 |
H4 |
114.187 |
| N3 |
C1 |
H4 |
126.729 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CISD/3-21G*
| | hartrees |
|---|
| Energy at 0K | -168.259552 |
| Energy at 298.15K | -168.263718 |
| HF Energy | -167.954973 |
| Nuclear repulsion energy | 70.141162 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3606 |
3369 |
18.52 |
|
|
|
| 2 |
A' |
3480 |
3252 |
1.15 |
|
|
|
| 3 |
A' |
3190 |
2980 |
32.30 |
|
|
|
| 4 |
A' |
1761 |
1645 |
149.09 |
|
|
|
| 5 |
A' |
1458 |
1362 |
26.88 |
|
|
|
| 6 |
A' |
1438 |
1344 |
37.21 |
|
|
|
| 7 |
A' |
1205 |
1126 |
74.50 |
|
|
|
| 8 |
A' |
1040 |
972 |
229.53 |
|
|
|
| 9 |
A' |
590 |
551 |
55.06 |
|
|
|
| 10 |
A" |
1117 |
1044 |
0.04 |
|
|
|
| 11 |
A" |
864 |
807 |
55.24 |
|
|
|
| 12 |
A" |
629 |
587 |
239.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10187.5 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 9519.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CISD/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.433 |
0.000 |
| O2 |
-1.123 |
-0.374 |
0.000 |
| N3 |
1.168 |
-0.057 |
0.000 |
| H4 |
-0.305 |
1.475 |
0.000 |
| H5 |
-0.817 |
-1.314 |
0.000 |
| H6 |
1.926 |
0.630 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3825 | 1.2668 | 1.0862 | 1.9286 | 1.9365 |
O2 | 1.3825 | | 2.3128 | 2.0218 | 0.9884 | 3.2101 | N3 | 1.2668 | 2.3128 | | 2.1258 | 2.3501 | 1.0229 | H4 | 1.0862 | 2.0218 | 2.1258 | | 2.8358 | 2.3865 | H5 | 1.9286 | 0.9884 | 2.3501 | 2.8358 | | 3.3625 | H6 | 1.9365 | 3.2101 | 1.0229 | 2.3865 | 3.3625 | |
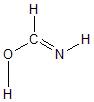 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
107.706 |
|
C1 |
N3 |
H6 |
115.089 |
| O2 |
C1 |
N3 |
121.555 |
|
O2 |
C1 |
H4 |
109.376 |
| N3 |
C1 |
H4 |
129.068 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CISD/3-21G*
| | hartrees |
|---|
| Energy at 0K | -168.255387 |
| Energy at 298.15K | -168.259454 |
| HF Energy | -167.949767 |
| Nuclear repulsion energy | 69.609297 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3604 |
3368 |
5.53 |
|
|
|
| 2 |
A' |
3393 |
3170 |
7.58 |
|
|
|
| 3 |
A' |
3270 |
3055 |
12.86 |
|
|
|
| 4 |
A' |
1744 |
1630 |
135.80 |
|
|
|
| 5 |
A' |
1476 |
1379 |
1.79 |
|
|
|
| 6 |
A' |
1386 |
1295 |
21.09 |
|
|
|
| 7 |
A' |
1167 |
1091 |
247.84 |
|
|
|
| 8 |
A' |
1044 |
975 |
131.43 |
|
|
|
| 9 |
A' |
590 |
551 |
39.99 |
|
|
|
| 10 |
A" |
1137 |
1063 |
85.88 |
|
|
|
| 11 |
A" |
880 |
823 |
55.85 |
|
|
|
| 12 |
A" |
477 |
446 |
136.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10083.4 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 9422.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CISD/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.449 |
0.000 |
| O2 |
-1.115 |
-0.391 |
0.000 |
| N3 |
1.235 |
0.163 |
0.000 |
| H4 |
-0.335 |
1.476 |
0.000 |
| H5 |
-0.846 |
-1.340 |
0.000 |
| H6 |
1.462 |
-0.842 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3963 | 1.2675 | 1.0803 | 1.9793 | 1.9501 |
O2 | 1.3963 | | 2.4147 | 2.0234 | 0.9867 | 2.6165 | N3 | 1.2675 | 2.4147 | | 2.0465 | 2.5674 | 1.0302 | H4 | 1.0803 | 2.0234 | 2.0465 | | 2.8623 | 2.9327 | H5 | 1.9793 | 0.9867 | 2.5674 | 2.8623 | | 2.3615 | H6 | 1.9501 | 2.6165 | 1.0302 | 2.9327 | 2.3615 | |
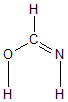 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
111.154 |
|
C1 |
N3 |
H6 |
115.768 |
| O2 |
C1 |
N3 |
129.991 |
|
O2 |
C1 |
H4 |
108.907 |
| N3 |
C1 |
H4 |
121.102 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
