Jump to
S1C2
S1C3
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -168.260914 |
| Energy at 298.15K | -168.264798 |
| HF Energy | -167.938248 |
| Nuclear repulsion energy | 69.322696 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3560 |
3386 |
24.06 |
|
|
|
| 2 |
A' |
3395 |
3230 |
1.82 |
|
|
|
| 3 |
A' |
3051 |
2902 |
68.54 |
|
|
|
| 4 |
A' |
1726 |
1642 |
82.45 |
|
|
|
| 5 |
A' |
1447 |
1377 |
5.07 |
|
|
|
| 6 |
A' |
1307 |
1243 |
115.87 |
|
|
|
| 7 |
A' |
1177 |
1120 |
74.72 |
|
|
|
| 8 |
A' |
1003 |
955 |
278.89 |
|
|
|
| 9 |
A' |
617 |
587 |
1.71 |
|
|
|
| 10 |
A" |
1077 |
1024 |
4.90 |
|
|
|
| 11 |
A" |
845 |
804 |
68.00 |
|
|
|
| 12 |
A" |
307 |
292 |
90.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9755.7 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 9280.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.398 |
0.000 |
| O2 |
-1.017 |
-0.562 |
0.000 |
| N3 |
1.215 |
0.001 |
0.000 |
| H4 |
-0.362 |
1.432 |
0.000 |
| H5 |
-1.896 |
-0.108 |
0.000 |
| H6 |
1.889 |
0.781 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3988 | 1.2784 | 1.0963 | 1.9621 | 1.9273 |
O2 | 1.3988 | | 2.3027 | 2.0999 | 0.9893 | 3.2016 | N3 | 1.2784 | 2.3027 | | 2.1297 | 3.1132 | 1.0303 | H4 | 1.0963 | 2.0999 | 2.1297 | | 2.1738 | 2.3429 | H5 | 1.9621 | 0.9893 | 3.1132 | 2.1738 | | 3.8876 | H6 | 1.9273 | 3.2016 | 1.0303 | 2.3429 | 3.8876 | |
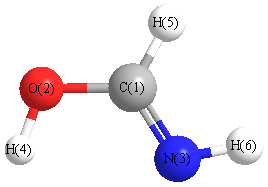 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.292 |
|
C1 |
N3 |
H6 |
112.750 |
| O2 |
C1 |
N3 |
118.596 |
|
O2 |
C1 |
H4 |
114.072 |
| N3 |
C1 |
H4 |
127.332 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -168.274926 |
| Energy at 298.15K | -168.279075 |
| HF Energy | -167.952760 |
| Nuclear repulsion energy | 69.679440 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3489 |
3319 |
18.27 |
|
|
|
| 2 |
A' |
3415 |
3249 |
1.77 |
|
|
|
| 3 |
A' |
3167 |
3012 |
36.62 |
|
|
|
| 4 |
A' |
1677 |
1596 |
119.53 |
|
|
|
| 5 |
A' |
1423 |
1354 |
0.58 |
|
|
|
| 6 |
A' |
1405 |
1337 |
57.55 |
|
|
|
| 7 |
A' |
1171 |
1114 |
72.48 |
|
|
|
| 8 |
A' |
1018 |
968 |
220.18 |
|
|
|
| 9 |
A' |
578 |
550 |
53.02 |
|
|
|
| 10 |
A" |
1089 |
1036 |
0.94 |
|
|
|
| 11 |
A" |
844 |
803 |
55.07 |
|
|
|
| 12 |
A" |
647 |
616 |
222.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9962.2 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 9477.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.438 |
0.000 |
| O2 |
-1.132 |
-0.368 |
0.000 |
| N3 |
1.177 |
-0.069 |
0.000 |
| H4 |
-0.307 |
1.482 |
0.000 |
| H5 |
-0.809 |
-1.311 |
0.000 |
| H6 |
1.935 |
0.626 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3894 | 1.2813 | 1.0882 | 1.9271 | 1.9437 |
O2 | 1.3894 | | 2.3276 | 2.0251 | 0.9974 | 3.2234 | N3 | 1.2813 | 2.3276 | | 2.1468 | 2.3416 | 1.0285 | H4 | 1.0882 | 2.0251 | 2.1468 | | 2.8378 | 2.3999 | H5 | 1.9271 | 0.9974 | 2.3416 | 2.8378 | | 3.3581 | H6 | 1.9437 | 3.2234 | 1.0285 | 2.3999 | 3.3581 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.533 |
|
C1 |
N3 |
H6 |
114.152 |
| O2 |
C1 |
N3 |
121.222 |
|
O2 |
C1 |
H4 |
109.043 |
| N3 |
C1 |
H4 |
129.735 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -168.271353 |
| Energy at 298.15K | -168.275409 |
| HF Energy | -167.947316 |
| Nuclear repulsion energy | 69.056114 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3491 |
3321 |
6.27 |
|
|
|
| 2 |
A' |
3322 |
3160 |
11.39 |
|
|
|
| 3 |
A' |
3251 |
3093 |
11.09 |
|
|
|
| 4 |
A' |
1649 |
1568 |
100.63 |
|
|
|
| 5 |
A' |
1464 |
1393 |
0.55 |
|
|
|
| 6 |
A' |
1353 |
1287 |
18.46 |
|
|
|
| 7 |
A' |
1139 |
1083 |
246.08 |
|
|
|
| 8 |
A' |
1008 |
959 |
127.93 |
|
|
|
| 9 |
A' |
574 |
546 |
36.12 |
|
|
|
| 10 |
A" |
1118 |
1064 |
86.47 |
|
|
|
| 11 |
A" |
853 |
812 |
44.57 |
|
|
|
| 12 |
A" |
504 |
480 |
125.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9863.0 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 9382.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.447 |
0.000 |
| O2 |
-1.129 |
-0.388 |
0.000 |
| N3 |
1.250 |
0.163 |
0.000 |
| H4 |
-0.334 |
1.476 |
0.000 |
| H5 |
-0.846 |
-1.342 |
0.000 |
| H6 |
1.465 |
-0.851 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.4047 | 1.2820 | 1.0822 | 1.9795 | 1.9578 |
O2 | 1.4047 | | 2.4425 | 2.0269 | 0.9955 | 2.6359 | N3 | 1.2820 | 2.4425 | | 2.0580 | 2.5807 | 1.0367 | H4 | 1.0822 | 2.0269 | 2.0580 | | 2.8648 | 2.9421 | H5 | 1.9795 | 0.9955 | 2.5807 | 2.8648 | | 2.3632 | H6 | 1.9578 | 2.6359 | 1.0367 | 2.9421 | 2.3632 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.945 |
|
C1 |
N3 |
H6 |
114.799 |
| O2 |
C1 |
N3 |
130.711 |
|
O2 |
C1 |
H4 |
108.487 |
| N3 |
C1 |
H4 |
120.802 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
