return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C10H8 (naphthalene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-382.079720
Energy at 298.15K-382.087784
HF Energy-381.208782
Nuclear repulsion energy454.101336
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3212 3055 0.00      
2 Ag 3172 3018 0.00      
3 Ag 1570 1494 0.00      
4 Ag 1512 1438 0.00      
5 Ag 1369 1302 0.00      
6 Ag 1235 1175 0.00      
7 Ag 1031 981 0.00      
8 Ag 762 725 0.00      
9 Ag 526 501 0.00      
10 Au 901 857 0.00      
11 Au 813 774 0.00      
12 Au 591 563 0.00      
13 Au 184 175 0.00      
14 B1g 900 857 0.00      
15 B1g 705 671 0.00      
16 B1g 380 362 0.00      
17 B1u 3194 3039 32.59      
18 B1u 3169 3015 16.29      
19 B1u 1608 1529 7.90      
20 B1u 1452 1382 3.96      
21 B1u 1315 1251 10.38      
22 B1u 1172 1115 0.91      
23 B1u 823 783 0.00      
24 B1u 372 354 0.94      
25 B2g 920 875 0.00      
26 B2g 822 782 0.00      
27 B2g 554 527 0.00      
28 B2g 439 418 0.00      
29 B2u 3211 3054 38.70      
30 B2u 3170 3016 0.00      
31 B2u 1533 1459 9.56      
32 B2u 1393 1326 1.64      
33 B2u 1251 1190 0.15      
34 B2u 1208 1149 2.41      
35 B2u 1018 968 2.92      
36 B2u 653 622 3.71      
37 B3g 3194 3038 0.00      
38 B3g 3167 3013 0.00      
39 B3g 1634 1554 0.00      
40 B3g 1501 1428 0.00      
41 B3g 1299 1236 0.00      
42 B3g 1198 1140 0.00      
43 B3g 971 924 0.00      
44 B3g 532 506 0.00      
45 B3u 902 859 0.06      
46 B3u 763 726 146.15      
47 B3u 457 435 16.82      
48 B3u 163 155 2.91      

Unscaled Zero Point Vibrational Energy (zpe) 31961.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30405.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.10241 0.04049 0.02902

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.254 1.415
C2 0.000 2.450 0.715
C3 0.000 2.450 -0.715
C4 0.000 1.254 -1.415
C5 0.000 -1.254 -1.415
C6 0.000 -2.450 -0.715
C7 0.000 -2.450 0.715
C8 0.000 -1.254 1.415
C9 0.000 0.000 0.720
C10 0.000 0.000 -0.720
H11 0.000 1.251 2.504
H12 0.000 3.396 1.250
H13 0.000 3.396 -1.250
H14 0.000 1.251 -2.504
H15 0.000 -1.251 -2.504
H16 0.000 -3.396 -1.250
H17 0.000 -3.396 1.250
H18 0.000 -1.251 2.504

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 C10 H11 H12 H13 H14 H15 H16 H17 H18
C11.38582.44222.82933.78074.27223.76942.50781.43332.47601.08952.14873.41893.91884.65115.35944.65312.7318
C21.38581.42932.44224.27225.10404.89983.76942.44992.83922.15381.08722.18053.43474.90506.16745.87064.1111
C32.44221.42931.38583.76944.89985.10404.27222.83922.44993.43472.18051.08722.15384.11115.87066.16744.9050
C42.82932.44221.38582.50783.76944.27223.78072.47601.43333.91883.41892.14871.08952.73184.65315.35944.6511
C53.78074.27223.76942.50781.38582.44222.82932.47601.43334.65115.35944.65312.73181.08952.14873.41893.9188
C64.27225.10404.89983.76941.38581.42932.44222.83922.44994.90506.16745.87064.11112.15381.08722.18053.4347
C73.76944.89985.10404.27222.44221.42931.38582.44992.83924.11115.87066.16744.90503.43472.18051.08722.1538
C82.50783.76944.27223.78072.82932.44221.38581.43332.47602.73184.65315.35944.65113.91883.41892.14871.0895
C91.43332.44992.83922.47602.47602.83922.44991.43331.44062.17893.43733.92633.45873.45873.92633.43732.1789
C102.47602.83922.44991.43331.43332.44992.83922.47601.44063.45873.92633.43732.17892.17893.43733.92633.4587
H111.08952.15383.43473.91884.65114.90504.11112.73182.17893.45872.48484.32355.00825.59875.97424.81392.5026
H122.14871.08722.18053.41895.35946.16745.87064.65313.43733.92632.48482.49964.32355.97427.23786.79254.8139
H133.41892.18051.08722.14874.65315.87066.16745.35943.92633.43734.32352.49962.48484.81396.79257.23785.9742
H143.91883.43472.15381.08952.73184.11114.90504.65113.45872.17895.00824.32352.48482.50264.81395.97425.5987
H154.65114.90504.11112.73181.08952.15383.43473.91883.45872.17895.59875.97424.81392.50262.48484.32355.0082
H165.35946.16745.87064.65312.14871.08722.18053.41893.92633.43735.97427.23786.79254.81392.48482.49964.3235
H174.65315.87066.16745.35943.41892.18051.08722.14873.43733.92634.81396.79257.23785.97424.32352.49962.4848
H182.73184.11114.90504.65113.91883.43472.15381.08952.17893.45872.50264.81395.97425.59875.00824.32352.4848

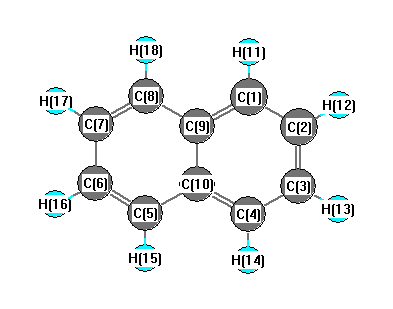
picture of naphthalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.340 C1 C2 H12 120.171
C1 C9 C8 122.048 C1 C9 C10 118.976
C2 C1 C9 120.684 C2 C1 H11 120.475
C2 C3 C4 120.340 C2 C3 H13 119.489
C3 C2 H12 119.489 C3 C4 C10 120.684
C3 C4 H14 120.475 C4 C3 H13 120.171
C4 C10 C5 122.048 C4 C10 C9 118.976
C5 C6 C7 120.340 C5 C6 H16 120.171
C5 C10 C9 118.976 C6 C5 C10 120.684
C6 C5 H15 120.475 C6 C7 C8 120.340
C6 C7 H17 119.489 C7 C6 H16 119.489
C7 C8 C9 120.684 C7 C8 H18 120.475
C8 C7 H17 120.171 C8 C9 C10 118.976
C9 C1 H11 118.841 C9 C8 H18 118.841
C10 C4 H14 118.841 C10 C5 H15 118.841
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability