return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (Ethene, 1,1-dimethoxy-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-304.707878
Energy at 298.15K-304.717143
HF Energy-304.108541
Nuclear repulsion energy243.768285
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3251 3093 0.27      
2 A1 3204 3048 20.20      
3 A1 3073 2924 5.13      
4 A1 1686 1604 127.92      
5 A1 1613 1534 4.66      
6 A1 1556 1480 1.41      
7 A1 1495 1422 24.26      
8 A1 1222 1163 1.65      
9 A1 1108 1054 0.98      
10 A1 834 794 35.46      
11 A1 484 461 0.06      
12 A1 258 245 5.83      
13 A2 3135 2982 0.00      
14 A2 1590 1512 0.00      
15 A2 1191 1133 0.00      
16 A2 685 652 0.00      
17 A2 243 231 0.00      
18 A2 116 110 0.00      
19 B1 3136 2983 70.64      
20 B1 1590 1512 11.74      
21 B1 1195 1137 0.24      
22 B1 790 752 120.19      
23 B1 673 640 7.10      
24 B1 263 251 0.16      
25 B1 87 83 12.87      
26 B2 3334 3171 5.65      
27 B2 3203 3047 12.29      
28 B2 3073 2924 36.13      
29 B2 1610 1532 18.11      
30 B2 1540 1465 7.11      
31 B2 1339 1274 388.72      
32 B2 1214 1155 53.81      
33 B2 1043 992 121.69      
34 B2 919 875 0.22      
35 B2 557 530 0.17      
36 B2 366 349 10.71      

Unscaled Zero Point Vibrational Energy (zpe) 26338.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 25055.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.29550 0.07309 0.05994

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.427
C2 0.000 0.000 0.079
H3 0.000 0.919 1.993
H4 0.000 -0.919 1.993
O5 0.000 1.106 -0.761
O6 0.000 -1.106 -0.761
C7 0.000 2.420 -0.095
C8 0.000 -2.420 -0.095
H9 0.000 3.143 -0.910
H10 0.000 -3.143 -0.910
H11 -0.893 2.548 0.527
H12 0.893 2.548 0.527
H13 0.893 -2.548 0.527
H14 -0.893 -2.548 0.527

Atom - Atom Distances (Å)
  C1 C2 H3 H4 O5 O6 C7 C8 H9 H10 H11 H12 H13 H14
C11.34791.07961.07962.45152.45152.85842.85843.91623.91622.84592.84592.84592.8459
C21.34792.12312.12311.38881.38882.42582.42583.29463.29462.73672.73672.73672.7367
H31.07962.12311.83892.76003.41842.57083.93803.65614.99262.36592.36593.86893.8689
H41.07962.12311.83893.41842.76003.93802.57084.99263.65613.86893.86892.36592.3659
O52.45151.38882.76003.41842.21191.47283.58792.04214.25132.12952.12953.97573.9757
O62.45151.38883.41842.76002.21193.58791.47284.25132.04213.97573.97572.12952.1295
C72.85842.42582.57083.93801.47283.58794.83921.08945.62171.09551.09555.08525.0852
C82.85842.42583.93802.57083.58791.47284.83925.62171.08945.08525.08521.09551.0955
H93.91623.29463.65614.99262.04214.25131.08945.62176.28541.79301.79305.93665.9366
H103.91623.29464.99263.65614.25132.04215.62171.08946.28545.93665.93661.79301.7930
H112.84592.73672.36593.86892.12953.97571.09555.08521.79305.93661.78565.39955.0958
H122.84592.73672.36593.86892.12953.97571.09555.08521.79305.93661.78565.09585.3995
H132.84592.73673.86892.36593.97572.12955.08521.09555.93661.79305.39955.09581.7856
H142.84592.73673.86892.36593.97572.12955.08521.09555.93661.79305.09585.39951.7856

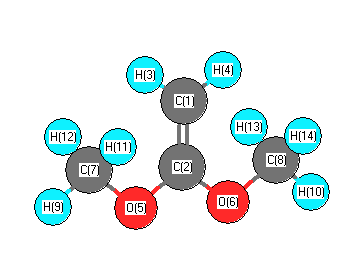
picture of Ethene, 1,1-dimethoxy- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O5 127.217 C1 C2 O6 127.217
C2 C1 H3 121.608 C2 C1 H4 121.608
C2 O5 C7 115.897 C2 O6 C8 115.897
H3 C1 H4 116.784 O5 C2 O6 105.565
O5 C7 H9 104.699 O5 C7 H11 111.168
O5 C7 H12 111.168 O6 C8 H10 104.699
O6 C8 H13 111.168 O6 C8 H14 111.168
H9 C7 H11 110.292 H9 C7 H12 110.292
H10 C8 H13 110.292 H10 C8 H14 110.292
H11 C7 H12 109.166 H13 C8 H14 109.166
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability