return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 (3-Methylenecyclopentene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-231.070722
Energy at 298.15K-231.079009
HF Energy-230.537785
Nuclear repulsion energy214.425467
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3247 3089 13.36      
2 A 3240 3082 12.95      
3 A 3210 3054 2.19      
4 A 3165 3010 5.84      
5 A 3164 3010 20.13      
6 A 3131 2978 7.69      
7 A 3114 2963 15.33      
8 A 3093 2943 22.28      
9 A 1692 1610 5.93      
10 A 1619 1540 0.06      
11 A 1586 1509 2.55      
12 A 1569 1493 1.08      
13 A 1523 1449 0.34      
14 A 1401 1333 0.80      
15 A 1360 1293 3.23      
16 A 1326 1262 0.96      
17 A 1306 1242 0.20      
18 A 1261 1200 2.49      
19 A 1235 1175 0.39      
20 A 1172 1115 1.25      
21 A 1070 1017 2.70      
22 A 1003 954 1.15      
23 A 950 904 1.34      
24 A 945 899 0.86      
25 A 899 855 43.49      
26 A 891 848 3.93      
27 A 873 831 9.90      
28 A 835 794 0.37      
29 A 794 755 18.15      
30 A 793 755 0.66      
31 A 678 645 21.05      
32 A 633 602 0.23      
33 A 508 483 14.11      
34 A 368 350 0.20      
35 A 279 265 1.23      
36 A 47 44 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 26987.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 25673.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.23757 0.11317 0.07894

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.094 -1.216 0.000
H2 -0.077 -1.831 -0.890
H3 -0.077 -1.831 0.890
C4 1.556 -0.615 -0.000
H5 2.117 -0.930 -0.888
H6 2.117 -0.930 0.887
C7 -2.192 -0.016 -0.000
H8 -2.783 0.898 -0.000
H9 -2.746 -0.951 -0.000
C10 -0.847 0.013 0.000
C11 1.325 0.901 -0.000
H12 2.147 1.609 -0.000
C13 0.012 1.222 0.000
H14 -0.390 2.231 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 H6 C7 H8 H9 C10 C11 H12 C13 H14
C11.09461.09461.58052.22752.22752.58263.57002.85271.54852.44943.49242.44013.4812
H21.09461.77932.22142.37162.96352.92603.94422.94792.18743.19704.19153.18134.1698
H31.09461.77932.22142.96352.37152.92613.94422.94812.18743.19694.19143.18124.1697
C41.58052.22142.22141.09611.09613.79594.59474.31512.48381.53392.30132.40003.4478
H52.22752.37162.96351.09611.77494.49385.30414.94353.23472.18352.68953.13864.1310
H62.22752.96352.37151.09611.77494.49385.30424.94363.23472.18352.68953.13864.1309
C72.58262.92602.92613.79594.49384.49381.08731.08701.34553.63494.63372.52822.8800
H83.57003.94423.94424.59475.30415.30421.08731.84882.12804.10784.98102.81352.7390
H92.85272.94792.94814.31514.94354.94361.08701.84882.12974.47275.52243.51153.9591
C101.54852.18742.18742.48383.23473.23471.34552.12802.12972.34683.39301.48332.2643
C112.44943.19703.19691.53392.18352.18353.63494.10784.47272.34681.08471.35182.1703
H123.49244.19154.19142.30132.68952.68954.63374.98105.52243.39301.08472.16992.6126
C132.44013.18133.18122.40003.13863.13862.52822.81353.51151.48331.35182.16991.0858
H143.48124.16984.16973.44784.13104.13092.88002.73903.95912.26432.17032.61261.0858

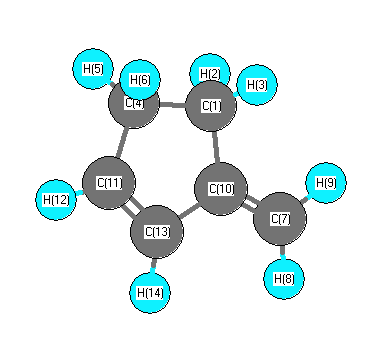
picture of 3-Methylenecyclopentene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 111.364 C1 C4 H6 111.362
C1 C4 C11 103.703 C1 C10 C7 126.212
C1 C10 C13 107.169 H2 C1 H3 108.735
H2 C1 C4 110.973 H2 C1 C10 110.532
H3 C1 C4 110.972 H3 C1 C10 110.531
C4 C1 C10 105.080 C4 C11 H12 122.064
C4 C11 C13 112.395 H5 C4 H6 108.118
H5 C4 C11 111.152 H6 C4 C11 111.150
C7 C10 C13 126.619 H8 C7 H9 116.487
H8 C7 C10 121.659 H9 C7 C10 121.854
C10 C13 C11 111.652 C10 C13 H14 122.864
C11 C13 H14 125.484 H12 C11 C13 125.541
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability