return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,2-dichlorobenzene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-1143.845901
Energy at 298.15K-1143.850093
HF Energy-1143.052869
Nuclear repulsion energy465.652344
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3232 3075 5.01      
2 A1 3214 3058 2.87      
3 A1 1572 1495 0.25      
4 A1 1503 1430 52.10      
5 A1 1307 1243 0.22      
6 A1 1244 1183 0.01      
7 A1 1149 1094 34.39      
8 A1 1046 995 6.64      
9 A1 689 656 12.00      
10 A1 478 455 8.26      
11 A1 204 194 0.00      
12 A2 928 883 0.00      
13 A2 850 808 0.00      
14 A2 663 630 0.00      
15 A2 519 493 0.00      
16 A2 140 133 0.00      
17 B1 918 874 2.15      
18 B1 762 724 82.32      
19 B1 449 427 6.29      
20 B1 236 224 1.40      
21 B2 3224 3067 1.42      
22 B2 3203 3047 0.98      
23 B2 1583 1506 3.21      
24 B2 1483 1411 17.14      
25 B2 1317 1253 5.60      
26 B2 1163 1106 4.75      
27 B2 1086 1034 29.18      
28 B2 766 728 16.64      
29 B2 437 416 0.71      
30 B2 344 327 0.36      

Unscaled Zero Point Vibrational Energy (zpe) 17852.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 16983.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.06295 0.04680 0.02684

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.703 -0.027
C2 0.000 -0.703 -0.027
C3 0.000 1.404 1.191
C4 0.000 -1.404 1.191
C5 0.000 0.703 2.407
C6 0.000 -0.703 2.407
Cl7 0.000 1.609 -1.526
Cl8 0.000 -1.609 -1.526
H9 0.000 2.489 1.172
H10 0.000 -2.489 1.172
H11 0.000 1.253 3.343
H12 0.000 -1.253 3.343

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C11.40531.40502.43282.43382.81071.75182.75542.15143.40943.41493.8967
C21.40532.43281.40502.81072.43382.75541.75183.40942.15143.89673.4149
C31.40502.43282.80701.40322.43262.72454.05681.08583.89272.15773.4193
C42.43281.40502.80702.43261.40324.05682.72453.89271.08583.41932.1577
C52.43382.81071.40322.43261.40684.03574.56232.17123.42301.08602.1692
C62.81072.43382.43261.40321.40684.56234.03573.42302.17122.16921.0860
Cl71.75182.75542.72454.05684.03574.56233.21862.83754.90664.88225.6483
Cl82.75541.75184.05682.72454.56234.03573.21864.90662.83755.64834.8822
H92.15143.40941.08583.89272.17123.42302.83754.90664.97822.49854.3267
H103.40942.15143.89271.08583.42302.17124.90662.83754.97824.32672.4985
H113.41493.89672.15773.41931.08602.16924.88225.64832.49854.32672.5065
H123.89673.41493.41932.15772.16921.08605.64834.88224.32672.49852.5065

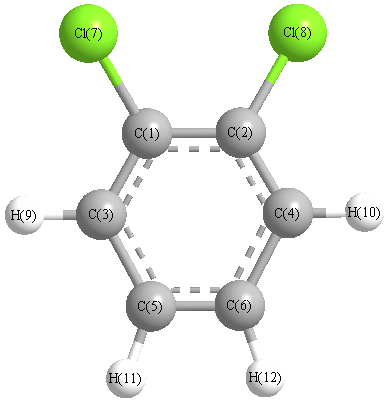
picture of 1,2-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 119.923 C1 C2 Cl8 121.168
C1 C3 C5 120.147 C1 C3 H9 118.922
C2 C1 C3 119.923 C2 C1 Cl7 121.168
C2 C4 C6 120.147 C2 C4 H10 118.922
C3 C1 Cl7 118.909 C3 C5 C6 119.930
C3 C5 H11 119.649 C4 C2 Cl8 118.909
C4 C6 C5 119.930 C4 C6 H12 119.649
C5 C3 H9 120.931 C5 C6 H12 120.421
C6 C4 H10 120.931 C6 C5 H11 120.421
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability