return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H14 (Pentane, 3-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-234.625645
Energy at 298.15K-234.640871
HF Energy-234.067736
Nuclear repulsion energy254.500525
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3167 3013 65.11      
2 A' 3151 2998 71.07      
3 A' 3148 2995 7.09      
4 A' 3111 2959 33.36      
5 A' 3083 2933 4.97      
6 A' 3077 2927 14.71      
7 A' 3068 2918 47.32      
8 A' 3053 2905 3.80      
9 A' 1619 1540 6.39      
10 A' 1607 1528 9.45      
11 A' 1598 1520 12.64      
12 A' 1587 1510 0.55      
13 A' 1496 1423 5.11      
14 A' 1481 1409 5.94      
15 A' 1444 1374 0.67      
16 A' 1420 1350 1.47      
17 A' 1369 1302 0.66      
18 A' 1260 1199 0.06      
19 A' 1207 1149 1.44      
20 A' 1053 1002 0.28      
21 A' 1022 973 1.53      
22 A' 982 934 3.67      
23 A' 856 814 2.85      
24 A' 777 739 3.59      
25 A' 447 425 0.22      
26 A' 396 376 0.14      
27 A' 234 223 0.04      
28 A' 206 196 0.02      
29 A' 87 83 0.00      
30 A" 3170 3015 8.12      
31 A" 3157 3003 19.84      
32 A" 3149 2995 21.47      
33 A" 3102 2951 0.08      
34 A" 3082 2932 29.29      
35 A" 3062 2912 0.69      
36 A" 1609 1531 5.50      
37 A" 1602 1524 0.27      
38 A" 1597 1520 0.52      
39 A" 1578 1501 0.02      
40 A" 1491 1418 5.39      
41 A" 1425 1355 0.14      
42 A" 1377 1310 0.18      
43 A" 1346 1281 1.02      
44 A" 1223 1164 4.65      
45 A" 1108 1054 2.64      
46 A" 1067 1015 7.24      
47 A" 1011 962 3.71      
48 A" 886 843 0.01      
49 A" 809 770 0.35      
50 A" 471 448 0.53      
51 A" 334 318 0.04      
52 A" 276 263 0.02      
53 A" 222 212 0.04      
54 A" 90 86 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 42623.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 40547.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.21511 0.06077 0.05250

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.207 0.000
C2 -1.546 0.108 0.000
H3 0.289 1.270 0.000
C4 0.603 -0.453 1.271
C5 0.603 -0.453 -1.271
C6 0.230 0.271 2.591
C7 0.230 0.271 -2.591
H8 -1.843 -0.949 0.000
H9 -1.973 0.590 0.885
H10 -1.973 0.590 -0.885
H11 0.268 -1.499 1.312
H12 0.268 -1.499 -1.312
H13 1.697 -0.464 1.170
H14 1.697 -0.464 -1.170
H15 0.735 -0.198 3.443
H16 0.735 -0.198 -3.443
H17 0.540 1.322 2.543
H18 -0.848 0.236 2.772
H19 0.540 1.322 -2.543
H20 -0.848 0.236 -2.772

Atom - Atom Distances (Å)
  C1 C2 H3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.54931.10111.55401.55402.60162.60162.17572.19652.19652.16922.16922.16752.16753.54343.54342.82862.89892.82862.8989
C21.54932.17162.55892.55893.14553.14551.09761.09471.09472.75602.75603.49463.49464.14104.14103.50592.86143.50592.8614
H31.10112.17162.16402.16402.77692.77693.07672.52242.52243.06423.06422.52132.52133.76903.76902.55583.16932.55583.1693
C41.55402.55892.16402.54221.55073.94672.80072.80583.51761.09912.80711.09882.67502.19034.72232.18472.19814.20744.3504
C51.55402.55892.16402.54223.94671.55072.80073.51762.80582.80711.09912.67501.09884.72232.19034.20744.35042.18472.1981
C62.60163.14552.77691.55073.94675.18123.53562.80474.12792.18424.28592.17014.10271.09596.07241.09621.09415.24905.4701
C72.60163.14552.77693.94671.55075.18123.53564.12792.80474.28592.18424.10272.17016.07241.09595.24905.47011.09621.0941
H82.17571.09763.07672.80072.80073.53563.53561.77971.77972.54602.54603.75983.75984.36594.36594.15953.17464.15953.1746
H92.19651.09472.52242.80583.51762.80474.12791.77971.77063.09343.77043.82904.33643.80735.16593.09832.22524.31333.8429
H102.19651.09472.52243.51762.80584.12792.80471.77971.77063.77043.09344.33643.82905.16593.80734.31333.84293.09832.2252
H112.16922.75603.06421.09912.80712.18424.28592.54603.09343.77042.62451.77043.04552.53944.95163.09002.52714.78494.5756
H122.16922.75603.06422.80711.09914.28592.18422.54603.77043.09342.62453.04551.77044.95162.53944.78494.57563.09002.5271
H132.16753.49462.52131.09882.67502.17014.10273.75983.82904.33641.77043.04552.33982.48204.71912.53243.08744.27944.7438
H142.16753.49462.52132.67501.09884.10272.17013.75984.33643.82903.04551.77042.33984.71912.48204.27944.74382.53243.0874
H153.54344.14103.76902.19034.72231.09596.07244.36593.80735.16592.53944.95162.48204.71916.88501.77731.77316.17866.4276
H163.54344.14103.76904.72232.19036.07241.09594.36595.16593.80734.95162.53944.71912.48206.88506.17866.42761.77731.7731
H172.82863.50592.55582.18474.20741.09625.24904.15953.09834.31333.09004.78492.53244.27941.77736.17861.77705.08595.5995
H182.89892.86143.16932.19814.35041.09415.47013.17462.22523.84292.52714.57563.08744.74381.77316.42761.77705.59955.5440
H192.82863.50592.55584.20742.18475.24901.09624.15954.31333.09834.78493.09004.27942.53246.17861.77735.08595.59951.7770
H202.89892.86143.16934.35042.19815.47011.09413.17463.84292.22524.57562.52714.74383.08746.42761.77315.59955.54401.7770

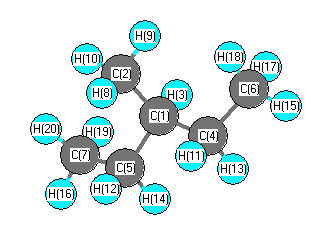
picture of Pentane, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H8 109.379 C1 C2 H9 111.178
C1 C2 H10 111.178 C1 C4 C6 113.849
C1 C4 H11 108.474 C1 C4 H13 108.357
C1 C5 C7 113.849 C1 C5 H12 108.474
C1 C5 H14 108.357 C2 C1 H3 108.861
C2 C1 C4 111.089 C2 C1 C5 111.089
H3 C1 C4 107.966 H3 C1 C5 107.966
C4 C1 C5 109.760 C4 C6 H15 110.524
C4 C6 H17 110.065 C4 C6 H18 111.253
C5 C7 H16 110.524 C5 C7 H19 110.065
C5 C7 H20 111.253 C6 C4 H11 109.861
C6 C4 H13 108.777 C7 C5 H12 109.861
C7 C5 H14 108.777 H8 C2 H9 108.544
H8 C2 H10 108.544 H9 C2 H10 107.937
H11 C4 H13 107.314 H12 C5 H14 107.314
H15 C6 H17 108.343 H15 C6 H18 108.117
H16 C7 H19 108.343 H16 C7 H20 108.117
H17 C6 H18 108.452 H19 C7 H20 108.452
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability