return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (2-methylbutyraldehyde)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-269.099765
Energy at 298.15K-269.110636
HF Energy-268.513265
Nuclear repulsion energy237.687112
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3179 3024 16.54      
2 A 3169 3015 19.45      
3 A 3166 3012 29.70      
4 A 3153 3000 26.30      
5 A 3113 2962 14.24      
6 A 3096 2945 9.85      
7 A 3084 2934 15.62      
8 A 3068 2918 22.42      
9 A 3055 2906 8.09      
10 A 2934 2791 119.21      
11 A 1634 1555 43.90      
12 A 1610 1531 3.68      
13 A 1605 1526 8.66      
14 A 1598 1520 4.82      
15 A 1594 1516 7.11      
16 A 1581 1504 0.06      
17 A 1499 1426 8.16      
18 A 1489 1417 6.92      
19 A 1464 1392 1.05      
20 A 1423 1354 4.85      
21 A 1399 1331 0.73      
22 A 1367 1300 2.90      
23 A 1344 1278 1.08      
24 A 1248 1187 1.20      
25 A 1185 1127 3.27      
26 A 1161 1105 5.37      
27 A 1057 1006 5.24      
28 A 1048 997 3.06      
29 A 1017 968 17.08      
30 A 969 921 0.34      
31 A 906 862 19.60      
32 A 825 785 5.26      
33 A 788 750 3.59      
34 A 675 642 5.02      
35 A 420 399 0.62      
36 A 398 378 0.50      
37 A 303 288 6.57      
38 A 271 258 0.55      
39 A 237 226 2.01      
40 A 197 188 0.60      
41 A 101 96 1.35      
42 A 85 81 2.71      

Unscaled Zero Point Vibrational Energy (zpe) 31756.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 30210.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.23575 0.06820 0.05730

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.370 -0.723 0.203
C2 -0.075 0.057 0.414
C3 1.078 -0.684 -0.324
C4 2.478 -0.116 0.023
C5 -0.257 1.515 -0.056
O6 -2.434 -0.231 -0.232
H7 -1.296 -1.796 0.467
H8 0.148 0.032 1.493
H9 1.048 -1.749 -0.055
H10 0.902 -0.611 -1.405
H11 3.260 -0.689 -0.488
H12 2.562 0.930 -0.285
H13 2.653 -0.177 1.103
H14 -0.426 1.533 -1.138
H15 -1.130 1.962 0.428
H16 0.626 2.117 0.177

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.52682.50503.89982.51271.25051.10762.12992.63922.78574.68164.29364.15812.78902.70523.4712
C21.52681.55632.58821.54212.46302.22011.10212.17742.16963.53412.86422.82282.17002.17772.1884
C32.50501.55631.54922.58583.54272.73882.16281.09921.09792.18772.19252.18372.79993.52722.8803
C43.89982.58821.54923.18504.91984.15482.75942.17202.18311.09561.09391.09603.53544.18322.9050
C52.51271.54212.58583.18502.79613.50932.18213.51462.77134.17232.88793.55951.09521.09401.0935
O61.25052.46303.54274.91982.79612.05783.11603.80243.55635.71815.12965.25922.82172.63553.8783
H71.10762.22012.73884.15483.50932.05782.54512.40153.12064.78504.78384.31473.79673.76224.3691
H82.12991.10212.16282.75942.18213.11602.54512.52523.06243.75913.13022.54393.08282.54792.5117
H92.63922.17741.09922.17203.51463.80242.40152.52521.77152.49133.08622.52713.75704.32983.8955
H102.78572.16961.09792.18312.77133.55633.12063.06241.77152.53102.52723.08882.53603.75613.1650
H114.68163.53412.18771.09564.17235.71814.78503.75912.49132.53101.77451.77824.35245.20923.9050
H124.29362.86422.19251.09392.88795.12964.78383.13023.08622.52721.77451.77783.16563.89922.3171
H134.15812.82282.18371.09603.55955.25924.31472.54392.52713.08881.77821.77784.17434.39773.1981
H142.78902.17002.79993.53541.09522.82173.79673.08283.75702.53604.35243.16564.17431.76971.7820
H152.70522.17773.52724.18321.09402.63553.76222.54794.32983.75615.20923.89924.39771.76971.7805
H163.47122.18842.88032.90501.09353.87834.36912.51173.89553.16503.90502.31713.19811.78201.7805

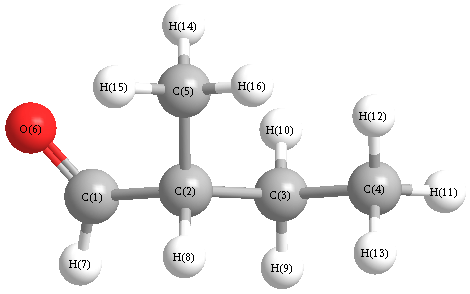
picture of 2-methylbutyraldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 108.677 C1 C2 C5 109.923
C1 C2 H8 107.123 C2 C1 O6 124.663
C2 C1 H7 113.916 C2 C3 C4 112.908
C2 C3 H9 108.948 C2 C3 H10 108.419
C2 C5 H14 109.564 C2 C5 H15 110.243
C2 C5 H16 111.121 C3 C2 C5 113.138
C3 C2 H8 107.666 C3 C4 H11 110.448
C3 C4 H12 110.926 C3 C4 H13 110.108
C4 C3 H9 109.004 C4 C3 H10 109.945
C5 C2 H8 110.109 O6 C1 H7 121.421
H9 C3 H10 107.467 H11 C4 H12 108.282
H11 C4 H13 108.462 H12 C4 H13 108.544
H14 C5 H15 107.870 H14 C5 H16 109.008
H15 C5 H16 108.964
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability