return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (Cyclopentane, methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-233.456265
Energy at 298.15K-233.470113
HF Energy-232.907989
Nuclear repulsion energy250.629824
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3169 3015 64.01      
2 A 3149 2996 7.84      
3 A 3146 2993 28.41      
4 A 3144 2991 23.97      
5 A 3139 2986 6.54      
6 A 3133 2980 21.85      
7 A 3111 2960 30.82      
8 A 3099 2948 18.92      
9 A 3086 2935 39.65      
10 A 3078 2928 17.13      
11 A 3070 2920 6.59      
12 A 3069 2920 13.75      
13 A 1606 1527 3.35      
14 A 1603 1525 6.26      
15 A 1598 1520 2.92      
16 A 1586 1509 4.49      
17 A 1585 1508 1.71      
18 A 1575 1498 0.32      
19 A 1489 1416 3.90      
20 A 1429 1359 1.67      
21 A 1410 1342 0.56      
22 A 1387 1319 0.88      
23 A 1386 1319 0.51      
24 A 1372 1305 0.77      
25 A 1360 1293 0.01      
26 A 1318 1254 0.67      
27 A 1300 1236 0.31      
28 A 1253 1192 0.63      
29 A 1246 1185 0.59      
30 A 1184 1127 2.81      
31 A 1118 1063 0.43      
32 A 1085 1032 1.57      
33 A 1021 971 2.79      
34 A 1019 970 0.22      
35 A 1009 960 0.39      
36 A 959 912 0.21      
37 A 918 873 0.63      
38 A 880 837 2.19      
39 A 854 813 0.60      
40 A 837 796 0.29      
41 A 777 739 0.90      
42 A 645 614 0.48      
43 A 540 514 0.56      
44 A 434 412 0.22      
45 A 312 297 0.11      
46 A 253 240 0.00      
47 A 180 172 0.02      
48 A 46 44 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 37981.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 36132.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.20411 0.09598 0.07234

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.697 -0.002 -1.476
C2 0.764 -0.001 -0.379
H3 0.196 -2.147 -0.342
H4 0.173 -1.317 1.235
C5 -0.049 -1.207 0.165
H6 0.194 2.146 -0.350
H7 0.177 1.323 1.230
C8 -0.049 1.208 0.161
H9 -1.920 1.166 -0.984
H10 -2.179 1.192 0.770
C11 -1.545 0.790 -0.027
H12 -2.184 -1.198 0.758
H13 -1.911 -1.162 -0.994
C14 -1.544 -0.791 -0.031
H15 2.761 0.889 -0.326
H16 2.761 -0.890 -0.324
H17 2.306 0.001 1.149
C18 2.243 -0.000 0.053

Atom - Atom Distances (Å)
  H1 C2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 H13 C14 H15 H16 H17 C18
H11.09952.47803.05822.16852.47713.05802.16892.90843.83972.78503.83712.89532.78112.52542.52523.07902.1750
C21.09952.22102.16451.55352.22102.16441.55362.98893.37672.46593.37882.98062.46442.18682.18682.17071.5410
H32.47802.22101.78191.09544.29303.80933.40153.98374.24533.42882.78802.41562.22793.97502.85743.36052.9933
H43.05822.16451.78191.09833.80782.63922.75223.93313.46982.99672.40733.05492.19693.74143.05122.50922.7231
C52.16851.55351.09541.09833.40092.75442.41523.23323.26442.50252.21542.19321.56403.54022.87042.82462.5934
H62.47712.22104.29303.80783.40091.78171.09542.41462.79162.22824.24963.97313.42702.85873.97573.36252.9945
H73.05802.16443.80932.63922.75441.78171.09833.05372.40382.19723.48523.93413.00293.04773.74002.50742.7210
C82.16891.55363.40152.75222.41521.09541.09832.19442.21511.56443.27113.22762.50332.87003.54022.82452.5933
H92.90842.98893.98373.93313.23322.41463.05372.19441.77311.09512.94872.32822.20914.73565.15554.87544.4465
H103.83973.37674.24533.46983.26442.79162.40382.21511.77311.09442.38952.95292.23025.06925.47114.65594.6356
C112.78502.46593.42882.99672.50252.22822.19721.56441.09511.09442.23072.20881.58074.31764.63174.10303.8705
H123.83713.37882.78802.40732.21544.24963.48523.27112.94872.38952.23071.77321.09445.47555.07144.66384.6400
H132.89532.98062.41563.05492.19323.97313.93413.22762.32822.95292.20881.77321.09515.14624.72824.87144.4392
C142.78112.46442.22792.19691.56403.42703.00292.50332.20912.23021.58071.09441.09514.63074.31644.10413.8699
H152.52542.18683.97503.74143.54022.85873.04772.87004.73565.06924.31765.47555.14624.63071.77841.78001.0964
H162.52522.18682.85743.05122.87043.97573.74003.54025.15555.47114.63175.07144.72824.31641.77841.78011.0964
H173.07902.17073.36052.50922.82463.36252.50742.82454.87544.65594.10304.66384.87144.10411.78001.78011.0970
C182.17501.54102.99332.72312.59342.99452.72102.59334.44654.63563.87054.64004.43923.86991.09641.09641.0970

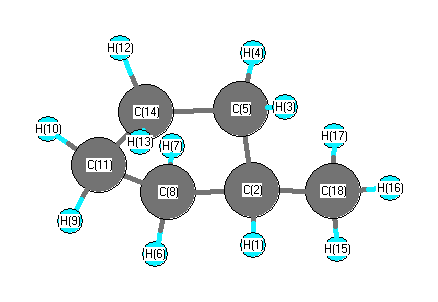
picture of Cyclopentane, methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C5 108.435 H1 C2 C8 108.453
H1 C2 C18 109.780 C2 C5 H3 112.818
C2 C5 H4 108.194 C2 C5 C14 104.466
C2 C8 H6 112.803 C2 C8 H7 108.182
C2 C8 C11 104.527 C2 C18 H15 110.898
C2 C18 H16 110.898 C2 C18 H17 109.589
H3 C5 H4 108.634 H3 C5 C14 112.617
H4 C5 C14 109.981 C5 C2 C8 102.032
C5 C2 C18 113.875 C5 C14 C11 105.458
C5 C14 H12 111.678 C5 C14 H13 109.881
H6 C8 H7 108.619 H6 C8 C11 112.609
H7 C8 C11 109.971 C8 C2 C18 113.859
C8 C11 H9 109.944 C8 C11 H10 111.613
C8 C11 C14 105.482 H9 C11 H10 108.157
H9 C11 C14 109.969 H10 C11 C14 111.669
C11 C14 H12 111.705 C11 C14 H13 109.942
H12 C14 H13 108.166 H15 C18 H16 108.397
H15 C18 H17 108.494 H16 C18 H17 108.495
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability