return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3COC6H5 (acetophenone)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-381.162499
Energy at 298.15K-381.170884
HF Energy-380.333229
Nuclear repulsion energy397.802420
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3222 3065 9.72      
2 A' 3215 3058 9.57      
3 A' 3207 3050 11.46      
4 A' 3196 3041 3.68      
5 A' 3190 3035 10.48      
6 A' 3187 3032 1.32      
7 A' 3090 2939 1.22      
8 A' 1642 1562 19.57      
9 A' 1606 1528 4.48      
10 A' 1587 1510 2.99      
11 A' 1559 1483 23.26      
12 A' 1535 1460 1.86      
13 A' 1495 1422 14.70      
14 A' 1461 1390 27.34      
15 A' 1386 1319 6.78      
16 A' 1331 1266 0.60      
17 A' 1271 1210 151.23      
18 A' 1248 1187 13.72      
19 A' 1238 1178 8.40      
20 A' 1125 1070 3.67      
21 A' 1112 1058 1.69      
22 A' 1054 1003 7.25      
23 A' 1020 970 0.82      
24 A' 960 913 30.36      
25 A' 742 705 0.32      
26 A' 656 624 0.50      
27 A' 586 558 20.47      
28 A' 470 447 0.15      
29 A' 370 352 1.09      
30 A' 224 213 4.71      
31 A" 3161 3007 6.36      
32 A" 1582 1505 11.09      
33 A" 1106 1052 1.96      
34 A" 949 903 1.08      
35 A" 933 887 0.28      
36 A" 901 857 1.68      
37 A" 850 808 0.57      
38 A" 757 720 57.01      
39 A" 689 656 27.73      
40 A" 614 584 11.41      
41 A" 426 405 1.04      
42 A" 417 397 0.00      
43 A" 202 192 0.55      
44 A" 156 148 0.05      
45 A" 75 71 2.42      

Unscaled Zero Point Vibrational Energy (zpe) 30399.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 28918.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.12009 0.04010 0.03023

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.524 2.319 0.000
C2 0.117 1.715 0.000
C3 0.000 0.211 0.000
O4 -0.911 2.437 0.000
C5 1.112 -0.654 0.000
C6 -1.302 -0.331 0.000
C7 0.919 -2.047 0.000
C8 -1.494 -1.718 0.000
C9 -0.381 -2.582 0.000
H10 1.421 3.407 0.000
H11 2.079 2.003 0.890
H12 2.079 2.003 -0.890
H13 2.123 -0.254 0.000
H14 -2.140 0.360 0.000
H15 1.779 -2.712 0.000
H16 -2.501 -2.128 0.000
H17 -0.526 -3.659 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17
C11.53092.60152.43753.00153.87384.40795.04085.25841.09261.09561.09562.64214.15505.03745.99826.3200
C21.53091.50831.25662.56872.48943.84603.79214.32522.13642.17352.17352.81002.63294.72794.65005.4119
C32.60151.50832.40511.40861.40992.43772.43992.81853.49802.88602.88602.17302.14573.42153.42423.9054
O42.43751.25662.40513.69372.79484.84284.19565.04642.52583.14973.14974.05502.41315.80894.83406.1078
C53.00152.56871.40863.69372.43541.40642.81532.43874.07262.96452.96451.08683.40702.16333.90223.4228
C63.87382.48941.40992.79482.43542.80701.40072.43214.62434.20384.20383.42541.08673.89402.16083.4175
C74.40793.84602.43774.84281.40642.80702.43591.40635.47704.30614.30612.15933.89331.08703.42112.1653
C85.04083.79212.43994.19562.81531.40072.43591.40885.89635.23565.23563.90212.17663.42121.08692.1690
C95.25844.32522.81855.04642.43872.43211.40631.40886.25425.27935.27933.41893.42812.16472.16751.0869
H101.09262.13643.49802.52584.07264.62435.47705.89636.25421.78761.78763.72774.68696.12926.78397.3295
H111.09562.17352.88603.14972.96454.20384.30615.23565.27931.78761.77982.42694.61504.80766.23226.2963
H121.09562.17352.88603.14972.96454.20384.30615.23565.27931.78761.77982.42694.61504.80766.23226.2963
H132.64212.81002.17304.05501.08683.42542.15933.90213.41893.72772.42692.42694.30732.48144.98904.3140
H144.15502.63292.14572.41313.40701.08673.89332.17663.42814.68694.61504.61504.30734.98032.51474.3315
H155.03744.72793.42155.80892.16333.89401.08703.42122.16476.12924.80764.80762.48144.98034.31982.4927
H165.99824.65003.42424.83403.90222.16083.42111.08692.16756.78396.23226.23224.98902.51474.31982.4984
H176.32005.41193.90546.10783.42283.41752.16532.16901.08697.32956.29636.29634.31404.33152.49272.4984

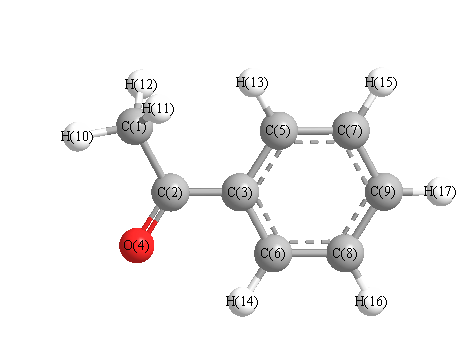
picture of acetophenone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 117.736 C1 C2 O4 121.650
C2 C1 H10 107.873 C2 C1 H11 110.602
C2 C1 H12 110.602 C2 C3 C5 123.394
C2 C3 C6 117.048 C3 C2 O4 120.614
C3 C5 C7 119.984 C3 C5 H13 120.557
C3 C6 C8 120.479 C3 C6 H14 117.930
C5 C3 C6 119.557 C5 C7 C9 120.232
C5 C7 H15 119.819 C6 C8 C9 119.916
C6 C8 H16 120.072 C7 C5 H13 119.459
C7 C9 C8 119.831 C7 C9 H17 120.017
C8 C6 H14 121.591 C8 C9 H17 120.151
C9 C7 H15 119.949 C9 C8 H16 120.013
H10 C1 H11 109.558 H10 C1 H12 109.558
H11 C1 H12 108.636
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability