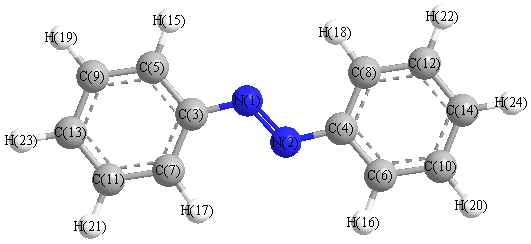
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3252 |
3252 |
0.00 |
|
|
|
| 2 |
Ag |
3241 |
3241 |
0.00 |
|
|
|
| 3 |
Ag |
3228 |
3228 |
0.00 |
|
|
|
| 4 |
Ag |
3217 |
3217 |
0.00 |
|
|
|
| 5 |
Ag |
3205 |
3205 |
0.00 |
|
|
|
| 6 |
Ag |
1651 |
1651 |
0.00 |
|
|
|
| 7 |
Ag |
1639 |
1639 |
0.00 |
|
|
|
| 8 |
Ag |
1538 |
1538 |
0.00 |
|
|
|
| 9 |
Ag |
1520 |
1520 |
0.00 |
|
|
|
| 10 |
Ag |
1423 |
1423 |
0.00 |
|
|
|
| 11 |
Ag |
1386 |
1386 |
0.00 |
|
|
|
| 12 |
Ag |
1376 |
1376 |
0.00 |
|
|
|
| 13 |
Ag |
1234 |
1234 |
0.00 |
|
|
|
| 14 |
Ag |
1227 |
1227 |
0.00 |
|
|
|
| 15 |
Ag |
1184 |
1184 |
0.00 |
|
|
|
| 16 |
Ag |
1118 |
1118 |
0.00 |
|
|
|
| 17 |
Ag |
1058 |
1058 |
0.00 |
|
|
|
| 18 |
Ag |
1035 |
1035 |
0.00 |
|
|
|
| 19 |
Ag |
934 |
934 |
0.00 |
|
|
|
| 20 |
Ag |
699 |
699 |
0.00 |
|
|
|
| 21 |
Ag |
645 |
645 |
0.00 |
|
|
|
| 22 |
Ag |
307 |
307 |
0.00 |
|
|
|
| 23 |
Ag |
221 |
221 |
0.00 |
|
|
|
| 24 |
Au |
985 |
985 |
0.35 |
|
|
|
| 25 |
Au |
976 |
976 |
0.11 |
|
|
|
| 26 |
Au |
942 |
942 |
8.79 |
|
|
|
| 27 |
Au |
869 |
869 |
0.00 |
|
|
|
| 28 |
Au |
788 |
788 |
117.51 |
|
|
|
| 29 |
Au |
693 |
693 |
48.53 |
|
|
|
| 30 |
Au |
556 |
556 |
9.85 |
|
|
|
| 31 |
Au |
421 |
421 |
0.02 |
|
|
|
| 32 |
Au |
310 |
310 |
1.08 |
|
|
|
| 33 |
Au |
64 |
64 |
1.53 |
|
|
|
| 34 |
Au |
30 |
30 |
0.05 |
|
|
|
| 35 |
Bg |
985 |
985 |
0.00 |
|
|
|
| 36 |
Bg |
976 |
976 |
0.00 |
|
|
|
| 37 |
Bg |
940 |
940 |
0.00 |
|
|
|
| 38 |
Bg |
870 |
870 |
0.00 |
|
|
|
| 39 |
Bg |
774 |
774 |
0.00 |
|
|
|
| 40 |
Bg |
681 |
681 |
0.00 |
|
|
|
| 41 |
Bg |
490 |
490 |
0.00 |
|
|
|
| 42 |
Bg |
424 |
424 |
0.00 |
|
|
|
| 43 |
Bg |
262 |
262 |
0.00 |
|
|
|
| 44 |
Bg |
113 |
113 |
0.00 |
|
|
|
| 45 |
Bu |
3252 |
3252 |
13.65 |
|
|
|
| 46 |
Bu |
3241 |
3241 |
40.18 |
|
|
|
| 47 |
Bu |
3228 |
3228 |
58.62 |
|
|
|
| 48 |
Bu |
3217 |
3217 |
26.46 |
|
|
|
| 49 |
Bu |
3205 |
3205 |
1.29 |
|
|
|
| 50 |
Bu |
1652 |
1652 |
2.57 |
|
|
|
| 51 |
Bu |
1636 |
1636 |
5.86 |
|
|
|
| 52 |
Bu |
1544 |
1544 |
14.21 |
|
|
|
| 53 |
Bu |
1512 |
1512 |
14.23 |
|
|
|
| 54 |
Bu |
1384 |
1384 |
8.59 |
|
|
|
| 55 |
Bu |
1374 |
1374 |
3.08 |
|
|
|
| 56 |
Bu |
1266 |
1266 |
17.51 |
|
|
|
| 57 |
Bu |
1227 |
1227 |
0.50 |
|
|
|
| 58 |
Bu |
1210 |
1210 |
48.68 |
|
|
|
| 59 |
Bu |
1121 |
1121 |
10.96 |
|
|
|
| 60 |
Bu |
1057 |
1057 |
15.58 |
|
|
|
| 61 |
Bu |
1036 |
1036 |
1.24 |
|
|
|
| 62 |
Bu |
842 |
842 |
0.12 |
|
|
|
| 63 |
Bu |
650 |
650 |
0.48 |
|
|
|
| 64 |
Bu |
558 |
558 |
5.59 |
|
|
|
| 65 |
Bu |
534 |
534 |
25.28 |
|
|
|
| 66 |
Bu |
85 |
85 |
1.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42156.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 42156.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.292 |
|
|
|
| 2 |
N |
-0.292 |
|
|
|
| 3 |
C |
0.140 |
|
|
|
| 4 |
C |
0.140 |
|
|
|
| 5 |
C |
-0.108 |
|
|
|
| 6 |
C |
-0.108 |
|
|
|
| 7 |
C |
-0.074 |
|
|
|
| 8 |
C |
-0.074 |
|
|
|
| 9 |
C |
-0.154 |
|
|
|
| 10 |
C |
-0.154 |
|
|
|
| 11 |
C |
-0.162 |
|
|
|
| 12 |
C |
-0.162 |
|
|
|
| 13 |
C |
-0.122 |
|
|
|
| 14 |
C |
-0.122 |
|
|
|
| 15 |
H |
0.158 |
|
|
|
| 16 |
H |
0.158 |
|
|
|
| 17 |
H |
0.176 |
|
|
|
| 18 |
H |
0.176 |
|
|
|
| 19 |
H |
0.147 |
|
|
|
| 20 |
H |
0.147 |
|
|
|
| 21 |
H |
0.147 |
|
|
|
| 22 |
H |
0.147 |
|
|
|
| 23 |
H |
0.145 |
|
|
|
| 24 |
H |
0.145 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-66.778 |
5.518 |
0.000 |
| y |
5.518 |
-69.487 |
0.000 |
| z |
0.000 |
0.000 |
-86.427 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
11.179 |
5.518 |
0.000 |
| y |
5.518 |
7.115 |
0.000 |
| z |
0.000 |
0.000 |
-18.295 |
|
| Polar |
|---|
| 3z2-r2 | -36.589 |
|---|
| x2-y2 | 2.709 |
|---|
| xy | 5.518 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
32.201 |
9.937 |
0.000 |
| y |
9.937 |
28.682 |
0.000 |
| z |
0.000 |
0.000 |
5.950 |
<r2> (average value of r
2) Å
2
| <r2> |
1115.471 |
| (<r2>)1/2 |
33.399 |