Jump to
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -5220.610078 |
| Energy at 298.15K | |
| HF Energy | -5220.523596 |
| Nuclear repulsion energy | 408.950391 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3168 |
3168 |
0.00 |
|
|
|
| 2 |
Ag |
1545 |
1545 |
0.00 |
|
|
|
| 3 |
Ag |
1328 |
1328 |
0.00 |
|
|
|
| 4 |
Ag |
1100 |
1100 |
0.00 |
|
|
|
| 5 |
Ag |
645 |
645 |
0.00 |
|
|
|
| 6 |
Ag |
185 |
185 |
0.00 |
|
|
|
| 7 |
Au |
3264 |
3264 |
2.47 |
|
|
|
| 8 |
Au |
1128 |
1128 |
2.55 |
|
|
|
| 9 |
Au |
781 |
781 |
6.71 |
|
|
|
| 10 |
Au |
103 |
103 |
5.37 |
|
|
|
| 11 |
Bg |
3242 |
3242 |
0.00 |
|
|
|
| 12 |
Bg |
1337 |
1337 |
0.00 |
|
|
|
| 13 |
Bg |
947 |
947 |
0.00 |
|
|
|
| 14 |
Bu |
3176 |
3176 |
6.57 |
|
|
|
| 15 |
Bu |
1542 |
1542 |
12.43 |
|
|
|
| 16 |
Bu |
1259 |
1259 |
56.89 |
|
|
|
| 17 |
Bu |
567 |
567 |
77.42 |
|
|
|
| 18 |
Bu |
180 |
180 |
8.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12748.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12748.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.506 |
0.558 |
0.000 |
| C2 |
-0.506 |
-0.558 |
0.000 |
| Br3 |
-0.506 |
2.311 |
0.000 |
| Br4 |
0.506 |
-2.311 |
0.000 |
| H5 |
1.121 |
0.572 |
0.896 |
| H6 |
1.121 |
0.572 |
-0.896 |
| H7 |
-1.121 |
-0.572 |
0.896 |
| H8 |
-1.121 |
-0.572 |
-0.896 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5068 | 2.0243 | 2.8689 | 1.0869 | 1.0869 | 2.1747 | 2.1747 |
C2 | 1.5068 | | 2.8689 | 2.0243 | 2.1747 | 2.1747 | 1.0869 | 1.0869 | Br3 | 2.0243 | 2.8689 | | 4.7314 | 2.5444 | 2.5444 | 3.0814 | 3.0814 | Br4 | 2.8689 | 2.0243 | 4.7314 | | 3.0814 | 3.0814 | 2.5444 | 2.5444 | H5 | 1.0869 | 2.1747 | 2.5444 | 3.0814 | | 1.7923 | 2.5176 | 3.0904 | H6 | 1.0869 | 2.1747 | 2.5444 | 3.0814 | 1.7923 | | 3.0904 | 2.5176 | H7 | 2.1747 | 1.0869 | 3.0814 | 2.5444 | 2.5176 | 3.0904 | | 1.7923 | H8 | 2.1747 | 1.0869 | 3.0814 | 2.5444 | 3.0904 | 2.5176 | 1.7923 | |
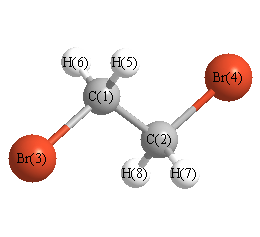 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
107.773 |
|
C1 |
C2 |
H7 |
112.955 |
| C1 |
C2 |
H8 |
112.955 |
|
C2 |
C1 |
Br3 |
107.773 |
| C2 |
C1 |
H5 |
112.955 |
|
C2 |
C1 |
H6 |
112.955 |
| Br3 |
C1 |
H5 |
105.748 |
|
Br3 |
C1 |
H6 |
105.748 |
| Br4 |
C2 |
H7 |
105.748 |
|
Br4 |
C2 |
H8 |
105.748 |
| H5 |
C1 |
H6 |
111.072 |
|
H7 |
C2 |
H8 |
111.072 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.462 |
|
|
|
| 2 |
C |
-0.462 |
|
|
|
| 3 |
Br |
-0.009 |
|
|
|
| 4 |
Br |
-0.009 |
|
|
|
| 5 |
H |
0.236 |
|
|
|
| 6 |
H |
0.236 |
|
|
|
| 7 |
H |
0.236 |
|
|
|
| 8 |
H |
0.236 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.307 |
-2.824 |
0.000 |
| y |
-2.824 |
11.722 |
0.000 |
| z |
0.000 |
0.000 |
3.837 |
<r2> (average value of r
2) Å
2
| <r2> |
440.821 |
| (<r2>)1/2 |
20.996 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -5220.605846 |
| Energy at 298.15K | |
| HF Energy | -5220.519140 |
| Nuclear repulsion energy | 447.056537 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3220 |
3220 |
0.09 |
110.23 |
0.75 |
0.86 |
| 2 |
A |
3147 |
3147 |
14.28 |
221.35 |
0.01 |
0.03 |
| 3 |
A |
1526 |
1526 |
0.41 |
8.65 |
0.71 |
0.83 |
| 4 |
A |
1352 |
1352 |
24.32 |
5.58 |
0.59 |
0.74 |
| 5 |
A |
1217 |
1217 |
2.10 |
30.58 |
0.74 |
0.85 |
| 6 |
A |
1071 |
1071 |
2.01 |
4.45 |
0.67 |
0.80 |
| 7 |
A |
925 |
925 |
8.35 |
17.65 |
0.56 |
0.72 |
| 8 |
A |
528 |
528 |
11.70 |
21.44 |
0.12 |
0.21 |
| 9 |
A |
227 |
227 |
1.28 |
2.07 |
0.48 |
0.65 |
| 10 |
A |
75 |
75 |
0.26 |
2.22 |
0.74 |
0.85 |
| 11 |
B |
3233 |
3233 |
2.47 |
33.32 |
0.75 |
0.86 |
| 12 |
B |
3139 |
3139 |
2.27 |
62.50 |
0.75 |
0.86 |
| 13 |
B |
1524 |
1524 |
14.12 |
23.64 |
0.75 |
0.86 |
| 14 |
B |
1327 |
1327 |
59.38 |
2.60 |
0.75 |
0.86 |
| 15 |
B |
1152 |
1152 |
2.12 |
11.62 |
0.75 |
0.86 |
| 16 |
B |
864 |
864 |
21.09 |
1.25 |
0.75 |
0.86 |
| 17 |
B |
566 |
566 |
16.01 |
16.64 |
0.75 |
0.86 |
| 18 |
B |
353 |
353 |
8.89 |
5.78 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12722.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12722.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.319 |
0.684 |
1.243 |
| C2 |
-0.319 |
-0.684 |
1.243 |
| Br3 |
-0.319 |
1.794 |
-0.307 |
| Br4 |
0.319 |
-1.794 |
-0.307 |
| H5 |
0.030 |
1.246 |
2.131 |
| H6 |
1.402 |
0.642 |
1.151 |
| H7 |
-0.030 |
-1.246 |
2.131 |
| H8 |
-1.402 |
-0.642 |
1.151 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5089 | 2.0102 | 2.9221 | 1.0904 | 1.0877 | 2.1530 | 2.1744 |
C2 | 1.5089 | | 2.9221 | 2.0102 | 2.1530 | 2.1744 | 1.0904 | 1.0877 | Br3 | 2.0102 | 2.9221 | | 3.6437 | 2.5226 | 2.5327 | 3.9075 | 3.0382 | Br4 | 2.9221 | 2.0102 | 3.6437 | | 3.9075 | 3.0382 | 2.5226 | 2.5327 | H5 | 1.0904 | 2.1530 | 2.5226 | 3.9075 | | 1.7914 | 2.4935 | 2.5642 | H6 | 1.0877 | 2.1744 | 2.5327 | 3.0382 | 1.7914 | | 2.5642 | 3.0840 | H7 | 2.1530 | 1.0904 | 3.9075 | 2.5226 | 2.4935 | 2.5642 | | 1.7914 | H8 | 2.1744 | 1.0877 | 3.0382 | 2.5327 | 2.5642 | 3.0840 | 1.7914 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
111.476 |
|
C1 |
C2 |
H7 |
110.816 |
| C1 |
C2 |
H8 |
112.725 |
|
C2 |
C1 |
Br3 |
111.476 |
| C2 |
C1 |
H5 |
110.816 |
|
C2 |
C1 |
H6 |
112.725 |
| Br3 |
C1 |
H5 |
104.984 |
|
Br3 |
C1 |
H6 |
105.796 |
| Br4 |
C2 |
H7 |
104.984 |
|
Br4 |
C2 |
H8 |
105.796 |
| H5 |
C1 |
H6 |
110.665 |
|
H7 |
C2 |
H8 |
110.665 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.454 |
|
|
|
| 2 |
C |
-0.454 |
|
|
|
| 3 |
Br |
0.019 |
|
|
|
| 4 |
Br |
0.019 |
|
|
|
| 5 |
H |
0.213 |
|
|
|
| 6 |
H |
0.222 |
|
|
|
| 7 |
H |
0.213 |
|
|
|
| 8 |
H |
0.222 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
3.011 |
3.011 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.385 |
1.119 |
0.000 |
| y |
1.119 |
-54.396 |
0.000 |
| z |
0.000 |
0.000 |
-45.144 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.385 |
1.119 |
0.000 |
| y |
1.119 |
-7.132 |
0.000 |
| z |
0.000 |
0.000 |
6.747 |
|
| Polar |
|---|
| 3z2-r2 | 13.495 |
|---|
| x2-y2 | 5.011 |
|---|
| xy | 1.119 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.423 |
-1.115 |
0.000 |
| y |
-1.115 |
7.418 |
0.000 |
| z |
0.000 |
0.000 |
7.362 |
<r2> (average value of r
2) Å
2
| <r2> |
314.895 |
| (<r2>)1/2 |
17.745 |
