Jump to
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -191.591924 |
| Energy at 298.15K | |
| HF Energy | -191.455835 |
| Nuclear repulsion energy | 101.972501 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3279 |
3279 |
10.46 |
67.63 |
0.52 |
0.68 |
| 2 |
A' |
3222 |
3222 |
2.23 |
89.14 |
0.32 |
0.48 |
| 3 |
A' |
3179 |
3179 |
6.02 |
69.49 |
0.10 |
0.19 |
| 4 |
A' |
2981 |
2981 |
92.87 |
90.49 |
0.27 |
0.42 |
| 5 |
A' |
1705 |
1705 |
54.81 |
57.60 |
0.29 |
0.45 |
| 6 |
A' |
1651 |
1651 |
89.42 |
15.59 |
0.56 |
0.72 |
| 7 |
A' |
1497 |
1497 |
3.72 |
9.47 |
0.45 |
0.62 |
| 8 |
A' |
1426 |
1426 |
6.84 |
18.36 |
0.44 |
0.61 |
| 9 |
A' |
1337 |
1337 |
1.78 |
12.66 |
0.33 |
0.50 |
| 10 |
A' |
1210 |
1210 |
30.70 |
12.67 |
0.63 |
0.78 |
| 11 |
A' |
955 |
955 |
23.03 |
0.99 |
0.10 |
0.18 |
| 12 |
A' |
576 |
576 |
6.52 |
7.26 |
0.38 |
0.55 |
| 13 |
A' |
326 |
326 |
10.12 |
0.47 |
0.26 |
0.41 |
| 14 |
A" |
1045 |
1045 |
22.64 |
0.50 |
0.75 |
0.86 |
| 15 |
A" |
1029 |
1029 |
6.37 |
6.05 |
0.75 |
0.86 |
| 16 |
A" |
1013 |
1013 |
40.50 |
1.82 |
0.75 |
0.86 |
| 17 |
A" |
619 |
619 |
11.35 |
6.21 |
0.75 |
0.86 |
| 18 |
A" |
178 |
178 |
5.24 |
1.22 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13613.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13613.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.132 |
-0.747 |
0.000 |
| C2 |
0.000 |
0.718 |
0.000 |
| C3 |
1.209 |
1.311 |
0.000 |
| O4 |
-1.227 |
-1.348 |
0.000 |
| H5 |
0.819 |
-1.311 |
0.000 |
| H6 |
-0.923 |
1.291 |
0.000 |
| H7 |
1.327 |
2.389 |
0.000 |
| H8 |
2.125 |
0.725 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4710 | 2.4564 | 1.2496 | 1.1049 | 2.1859 | 3.4584 | 2.6944 |
C2 | 1.4710 | | 1.3470 | 2.4032 | 2.1879 | 1.0861 | 2.1336 | 2.1255 | C3 | 2.4564 | 1.3470 | | 3.6066 | 2.6508 | 2.1326 | 1.0841 | 1.0876 | O4 | 1.2496 | 2.4032 | 3.6066 | | 2.0460 | 2.6562 | 4.5264 | 3.9416 | H5 | 1.1049 | 2.1879 | 2.6508 | 2.0460 | | 3.1306 | 3.7343 | 2.4189 | H6 | 2.1859 | 1.0861 | 2.1326 | 2.6562 | 3.1306 | | 2.5039 | 3.1006 | H7 | 3.4584 | 2.1336 | 1.0841 | 4.5264 | 3.7343 | 2.5039 | | 1.8456 | H8 | 2.6944 | 2.1255 | 1.0876 | 3.9416 | 2.4189 | 3.1006 | 1.8456 | |
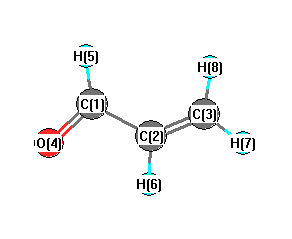 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.245 |
|
C1 |
C2 |
H6 |
116.672 |
| C2 |
C1 |
O4 |
123.888 |
|
C2 |
C1 |
H5 |
115.552 |
| C2 |
C3 |
H7 |
122.348 |
|
C2 |
C3 |
H8 |
121.261 |
| C3 |
C2 |
H6 |
122.083 |
|
O4 |
C1 |
H5 |
120.559 |
| H7 |
C3 |
H8 |
116.390 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.198 |
|
|
|
| 2 |
C |
-0.108 |
|
|
|
| 3 |
C |
-0.288 |
|
|
|
| 4 |
O |
-0.405 |
|
|
|
| 5 |
H |
0.121 |
|
|
|
| 6 |
H |
0.169 |
|
|
|
| 7 |
H |
0.159 |
|
|
|
| 8 |
H |
0.153 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.634 |
2.221 |
0.000 |
3.446 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.911 |
-3.003 |
-0.002 |
| y |
-3.003 |
-24.458 |
-0.002 |
| z |
-0.002 |
-0.002 |
-24.735 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.314 |
-3.003 |
-0.002 |
| y |
-3.003 |
0.365 |
-0.002 |
| z |
-0.002 |
-0.002 |
-0.051 |
|
| Polar |
|---|
| 3z2-r2 | -0.102 |
|---|
| x2-y2 | -0.453 |
|---|
| xy | -3.003 |
|---|
| xz | -0.002 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.670 |
1.841 |
0.000 |
| y |
1.841 |
6.014 |
0.000 |
| z |
0.000 |
0.000 |
1.804 |
<r2> (average value of r
2) Å
2
| <r2> |
85.065 |
| (<r2>)1/2 |
9.223 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -191.589870 |
| Energy at 298.15K | |
| HF Energy | -191.453745 |
| Nuclear repulsion energy | 103.534525 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3289 |
3289 |
5.19 |
54.75 |
0.61 |
0.76 |
| 2 |
A' |
3210 |
3210 |
12.08 |
120.82 |
0.22 |
0.37 |
| 3 |
A' |
3188 |
3188 |
5.18 |
44.41 |
0.15 |
0.26 |
| 4 |
A' |
3016 |
3016 |
128.51 |
144.14 |
0.31 |
0.48 |
| 5 |
A' |
1707 |
1707 |
9.14 |
7.47 |
0.20 |
0.33 |
| 6 |
A' |
1636 |
1636 |
99.95 |
46.76 |
0.33 |
0.50 |
| 7 |
A' |
1489 |
1489 |
37.64 |
11.73 |
0.34 |
0.50 |
| 8 |
A' |
1448 |
1448 |
4.73 |
11.89 |
0.43 |
0.60 |
| 9 |
A' |
1356 |
1356 |
2.65 |
15.34 |
0.45 |
0.62 |
| 10 |
A' |
1107 |
1107 |
6.06 |
5.03 |
0.75 |
0.86 |
| 11 |
A' |
962 |
962 |
56.26 |
5.41 |
0.22 |
0.37 |
| 12 |
A' |
695 |
695 |
9.90 |
1.36 |
0.71 |
0.83 |
| 13 |
A' |
292 |
292 |
8.39 |
3.61 |
0.51 |
0.67 |
| 14 |
A" |
1044 |
1044 |
40.33 |
2.64 |
0.75 |
0.86 |
| 15 |
A" |
1037 |
1037 |
17.32 |
8.37 |
0.75 |
0.86 |
| 16 |
A" |
1023 |
1023 |
2.54 |
0.65 |
0.75 |
0.86 |
| 17 |
A" |
580 |
580 |
11.07 |
7.76 |
0.75 |
0.86 |
| 18 |
A" |
191 |
191 |
8.68 |
0.88 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13635.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13635.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.897 |
-0.273 |
0.000 |
| C2 |
0.000 |
0.903 |
0.000 |
| C3 |
1.341 |
0.775 |
0.000 |
| O4 |
-0.501 |
-1.460 |
0.000 |
| H5 |
-1.977 |
-0.055 |
0.000 |
| H6 |
-0.476 |
1.881 |
0.000 |
| H7 |
2.003 |
1.634 |
0.000 |
| H8 |
1.796 |
-0.211 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4796 | 2.4714 | 1.2514 | 1.1015 | 2.1946 | 3.4711 | 2.6939 |
C2 | 1.4796 | | 1.3469 | 2.4163 | 2.1971 | 1.0868 | 2.1322 | 2.1135 | C3 | 2.4714 | 1.3469 | | 2.8967 | 3.4201 | 2.1262 | 1.0842 | 1.0861 | O4 | 1.2514 | 2.4163 | 2.8967 | | 2.0377 | 3.3410 | 3.9805 | 2.6150 | H5 | 1.1015 | 2.1971 | 3.4201 | 2.0377 | | 2.4497 | 4.3237 | 3.7761 | H6 | 2.1946 | 1.0868 | 2.1262 | 3.3410 | 2.4497 | | 2.4913 | 3.0877 | H7 | 3.4711 | 2.1322 | 1.0842 | 3.9805 | 4.3237 | 2.4913 | | 1.8558 | H8 | 2.6939 | 2.1135 | 1.0861 | 2.6150 | 3.7761 | 3.0877 | 1.8558 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.869 |
|
C1 |
C2 |
H6 |
116.717 |
| C2 |
C1 |
O4 |
124.228 |
|
C2 |
C1 |
H5 |
115.913 |
| C2 |
C3 |
H7 |
122.217 |
|
C2 |
C3 |
H8 |
120.236 |
| C3 |
C2 |
H6 |
121.414 |
|
O4 |
C1 |
H5 |
119.859 |
| H7 |
C3 |
H8 |
117.546 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.196 |
|
|
|
| 2 |
C |
-0.138 |
|
|
|
| 3 |
C |
-0.258 |
|
|
|
| 4 |
O |
-0.401 |
|
|
|
| 5 |
H |
0.122 |
|
|
|
| 6 |
H |
0.149 |
|
|
|
| 7 |
H |
0.152 |
|
|
|
| 8 |
H |
0.178 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.221 |
2.903 |
0.000 |
2.912 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.224 |
-0.739 |
-0.000 |
| y |
-0.739 |
-26.884 |
-0.002 |
| z |
-0.000 |
-0.002 |
-24.711 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.574 |
-0.739 |
-0.000 |
| y |
-0.739 |
-4.417 |
-0.002 |
| z |
-0.000 |
-0.002 |
-1.157 |
|
| Polar |
|---|
| 3z2-r2 | -2.313 |
|---|
| x2-y2 | 6.660 |
|---|
| xy | -0.739 |
|---|
| xz | -0.000 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.657 |
0.472 |
0.000 |
| y |
0.472 |
5.547 |
0.000 |
| z |
0.000 |
0.000 |
1.801 |
<r2> (average value of r
2) Å
2
| <r2> |
76.215 |
| (<r2>)1/2 |
8.730 |
