Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP/6-31G
Geometric Data calculated at B2PLYP/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -998.443683 |
| Energy at 298.15K | -998.448412 |
| HF Energy | -998.352660 |
| Nuclear repulsion energy | 195.703095 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3218 |
3218 |
0.16 |
|
|
|
| 2 |
A |
3146 |
3146 |
13.44 |
|
|
|
| 3 |
A |
1527 |
1527 |
0.85 |
|
|
|
| 4 |
A |
1373 |
1373 |
20.72 |
|
|
|
| 5 |
A |
1247 |
1247 |
0.76 |
|
|
|
| 6 |
A |
1082 |
1082 |
1.87 |
|
|
|
| 7 |
A |
970 |
970 |
11.26 |
|
|
|
| 8 |
A |
606 |
606 |
22.59 |
|
|
|
| 9 |
A |
259 |
259 |
1.34 |
|
|
|
| 10 |
A |
105 |
105 |
1.50 |
|
|
|
| 11 |
B |
3230 |
3230 |
2.39 |
|
|
|
| 12 |
B |
3138 |
3138 |
2.13 |
|
|
|
| 13 |
B |
1528 |
1528 |
14.68 |
|
|
|
| 14 |
B |
1359 |
1359 |
44.98 |
|
|
|
| 15 |
B |
1181 |
1181 |
1.01 |
|
|
|
| 16 |
B |
911 |
911 |
19.24 |
|
|
|
| 17 |
B |
630 |
630 |
31.82 |
|
|
|
| 18 |
B |
400 |
400 |
11.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12954.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12954.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.314 |
0.686 |
0.930 |
| C2 |
-0.314 |
-0.686 |
0.930 |
| Cl3 |
-0.314 |
1.757 |
-0.485 |
| Cl4 |
0.314 |
-1.757 |
-0.485 |
| H5 |
0.040 |
1.225 |
1.838 |
| H6 |
1.396 |
0.652 |
0.820 |
| H7 |
-0.040 |
-1.225 |
1.838 |
| H8 |
-1.396 |
-0.652 |
0.820 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5096 | 1.8829 | 2.8239 | 1.0908 | 1.0878 | 2.1457 | 2.1743 |
C2 | 1.5096 | | 2.8239 | 1.8829 | 2.1457 | 2.1743 | 1.0908 | 1.0878 | Cl3 | 1.8829 | 2.8239 | | 3.5705 | 2.4093 | 2.4186 | 3.7906 | 2.9458 | Cl4 | 2.8239 | 1.8829 | 3.5705 | | 3.7906 | 2.9458 | 2.4093 | 2.4186 | H5 | 1.0908 | 2.1457 | 2.4093 | 3.7906 | | 1.7899 | 2.4520 | 2.5735 | H6 | 1.0878 | 2.1743 | 2.4186 | 2.9458 | 1.7899 | | 2.5735 | 3.0813 | H7 | 2.1457 | 1.0908 | 3.7906 | 2.4093 | 2.4520 | 2.5735 | | 1.7899 | H8 | 2.1743 | 1.0878 | 2.9458 | 2.4186 | 2.5735 | 3.0813 | 1.7899 | |
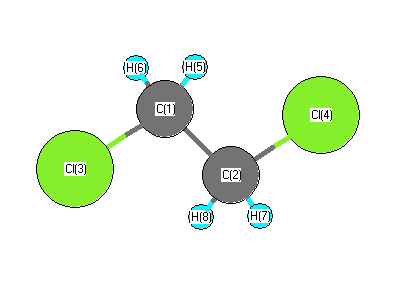 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.226 |
|
C1 |
C2 |
H7 |
110.163 |
| C1 |
C2 |
H8 |
112.658 |
|
C2 |
C1 |
Cl3 |
112.226 |
| C2 |
C1 |
H5 |
110.163 |
|
C2 |
C1 |
H6 |
112.658 |
| Cl3 |
C1 |
H5 |
105.100 |
|
Cl3 |
C1 |
H6 |
105.880 |
| Cl4 |
C2 |
H7 |
105.100 |
|
Cl4 |
C2 |
H8 |
105.880 |
| H5 |
C1 |
H6 |
110.486 |
|
H7 |
C2 |
H8 |
110.486 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.399 |
|
|
|
| 2 |
C |
-0.399 |
|
|
|
| 3 |
Cl |
-0.034 |
|
|
|
| 4 |
Cl |
-0.034 |
|
|
|
| 5 |
H |
0.210 |
|
|
|
| 6 |
H |
0.223 |
|
|
|
| 7 |
H |
0.210 |
|
|
|
| 8 |
H |
0.223 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
3.512 |
3.512 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.848 |
1.741 |
0.000 |
| y |
1.741 |
-44.166 |
0.000 |
| z |
0.000 |
0.000 |
-35.789 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.129 |
1.741 |
0.000 |
| y |
1.741 |
-7.347 |
0.000 |
| z |
0.000 |
0.000 |
5.218 |
|
| Polar |
|---|
| 3z2-r2 | 10.437 |
|---|
| x2-y2 | 6.318 |
|---|
| xy | 1.741 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.185 |
-0.874 |
0.000 |
| y |
-0.874 |
6.506 |
0.000 |
| z |
0.000 |
0.000 |
6.266 |
<r2> (average value of r
2) Å
2
| <r2> |
173.963 |
| (<r2>)1/2 |
13.189 |
