Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP/6-31G
Geometric Data calculated at B2PLYP/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -612.953719 |
| Energy at 298.15K | -612.957116 |
| HF Energy | -612.831986 |
| Nuclear repulsion energy | 143.506774 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3118 |
3118 |
10.42 |
|
|
|
| 2 |
A' |
2992 |
2992 |
78.01 |
|
|
|
| 3 |
A' |
1705 |
1705 |
73.97 |
|
|
|
| 4 |
A' |
1503 |
1503 |
31.41 |
|
|
|
| 5 |
A' |
1436 |
1436 |
13.32 |
|
|
|
| 6 |
A' |
1326 |
1326 |
16.72 |
|
|
|
| 7 |
A' |
971 |
971 |
5.55 |
|
|
|
| 8 |
A' |
758 |
758 |
4.47 |
|
|
|
| 9 |
A' |
591 |
591 |
43.48 |
|
|
|
| 10 |
A' |
215 |
215 |
2.12 |
|
|
|
| 11 |
A" |
3181 |
3181 |
0.71 |
|
|
|
| 12 |
A" |
1224 |
1224 |
2.68 |
|
|
|
| 13 |
A" |
1042 |
1042 |
1.16 |
|
|
|
| 14 |
A" |
734 |
734 |
4.60 |
|
|
|
| 15 |
A" |
145 |
145 |
0.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10470.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10470.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.960 |
0.000 |
| C2 |
1.377 |
0.342 |
0.000 |
| Cl3 |
-1.369 |
-0.302 |
0.000 |
| O4 |
1.638 |
-0.868 |
0.000 |
| H5 |
-0.136 |
1.580 |
0.889 |
| H6 |
-0.136 |
1.580 |
-0.889 |
| H7 |
2.181 |
1.099 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5095 | 1.8618 | 2.4543 | 1.0926 | 1.0926 | 2.1858 |
C2 | 1.5095 | | 2.8210 | 1.2374 | 2.1476 | 2.1476 | 1.1043 | Cl3 | 1.8618 | 2.8210 | | 3.0606 | 2.4190 | 2.4190 | 3.8170 | O4 | 2.4543 | 1.2374 | 3.0606 | | 3.1508 | 3.1508 | 2.0399 | H5 | 1.0926 | 2.1476 | 2.4190 | 3.1508 | | 1.7788 | 2.5282 | H6 | 1.0926 | 2.1476 | 2.4190 | 3.1508 | 1.7788 | | 2.5282 | H7 | 2.1858 | 1.1043 | 3.8170 | 2.0399 | 2.5282 | 2.5282 | |
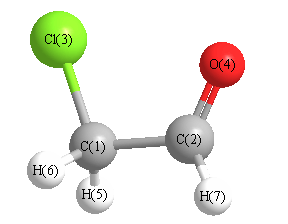 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
126.349 |
|
C1 |
C2 |
H7 |
112.574 |
| C2 |
C1 |
Cl3 |
113.191 |
|
C2 |
C1 |
H5 |
110.223 |
| C2 |
C1 |
H6 |
110.223 |
|
Cl3 |
C1 |
H5 |
107.030 |
| Cl3 |
C1 |
H6 |
107.030 |
|
O4 |
C2 |
H7 |
121.077 |
| H5 |
C1 |
H6 |
108.991 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.526 |
|
|
|
| 2 |
C |
0.292 |
|
|
|
| 3 |
Cl |
0.001 |
|
|
|
| 4 |
O |
-0.392 |
|
|
|
| 5 |
H |
0.233 |
|
|
|
| 6 |
H |
0.233 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.839 |
4.318 |
0.000 |
4.399 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.476 |
4.099 |
0.000 |
| y |
4.099 |
-29.702 |
0.000 |
| z |
0.000 |
0.000 |
-29.517 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.867 |
4.099 |
0.000 |
| y |
4.099 |
1.794 |
0.000 |
| z |
0.000 |
0.000 |
2.072 |
|
| Polar |
|---|
| 3z2-r2 | 4.144 |
|---|
| x2-y2 | -3.774 |
|---|
| xy | 4.099 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.599 |
0.865 |
0.000 |
| y |
0.865 |
5.371 |
0.000 |
| z |
0.000 |
0.000 |
2.708 |
<r2> (average value of r
2) Å
2
| <r2> |
110.406 |
| (<r2>)1/2 |
10.507 |
