Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-31G
Geometric Data calculated at MP2=FULL/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -208.216977 |
| Energy at 298.15K | |
| HF Energy | -207.782652 |
| Nuclear repulsion energy | 116.753043 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3633 |
3451 |
43.81 |
|
|
|
| 2 |
A' |
3206 |
3046 |
10.43 |
|
|
|
| 3 |
A' |
3191 |
3032 |
5.65 |
|
|
|
| 4 |
A' |
3068 |
2915 |
13.72 |
|
|
|
| 5 |
A' |
1664 |
1581 |
0.14 |
|
|
|
| 6 |
A' |
1554 |
1477 |
10.39 |
|
|
|
| 7 |
A' |
1481 |
1407 |
14.03 |
|
|
|
| 8 |
A' |
1393 |
1323 |
0.87 |
|
|
|
| 9 |
A' |
1309 |
1244 |
66.12 |
|
|
|
| 10 |
A' |
1176 |
1117 |
20.41 |
|
|
|
| 11 |
A' |
943 |
896 |
18.89 |
|
|
|
| 12 |
A' |
766 |
728 |
47.18 |
|
|
|
| 13 |
A' |
645 |
613 |
25.97 |
|
|
|
| 14 |
A' |
308 |
293 |
3.06 |
|
|
|
| 15 |
A" |
3135 |
2979 |
16.31 |
|
|
|
| 16 |
A" |
1565 |
1487 |
13.55 |
|
|
|
| 17 |
A" |
1130 |
1073 |
0.68 |
|
|
|
| 18 |
A" |
882 |
838 |
12.62 |
|
|
|
| 19 |
A" |
466 |
443 |
33.49 |
|
|
|
| 20 |
A" |
358 |
340 |
154.82 |
|
|
|
| 21 |
A" |
82i |
78i |
0.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15894.9 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 15101.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.457 |
0.504 |
0.000 |
| C2 |
0.000 |
0.893 |
0.000 |
| N3 |
1.051 |
0.117 |
0.000 |
| O4 |
0.637 |
-1.328 |
0.000 |
| H5 |
1.509 |
-1.778 |
0.000 |
| H6 |
-1.566 |
-0.584 |
0.000 |
| H7 |
-1.959 |
0.916 |
0.887 |
| H8 |
-1.959 |
0.916 |
-0.887 |
| H9 |
0.263 |
1.952 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5084 | 2.5385 | 2.7827 | 3.7430 | 1.0934 | 1.0995 | 1.0995 | 2.2487 |
C2 | 1.5084 | | 1.3069 | 2.3106 | 3.0682 | 2.1526 | 2.1507 | 2.1507 | 1.0909 | N3 | 2.5385 | 1.3069 | | 1.5030 | 1.9497 | 2.7095 | 3.2386 | 3.2386 | 1.9970 | O4 | 2.7827 | 2.3106 | 1.5030 | | 0.9815 | 2.3254 | 3.5445 | 3.5445 | 3.3009 | H5 | 3.7430 | 3.0682 | 1.9497 | 0.9815 | | 3.2990 | 4.4807 | 4.4807 | 3.9326 | H6 | 1.0934 | 2.1526 | 2.7095 | 2.3254 | 3.2990 | | 1.7867 | 1.7867 | 3.1267 | H7 | 1.0995 | 2.1507 | 3.2386 | 3.5445 | 4.4807 | 1.7867 | | 1.7747 | 2.6074 | H8 | 1.0995 | 2.1507 | 3.2386 | 3.5445 | 4.4807 | 1.7867 | 1.7747 | | 2.6074 | H9 | 2.2487 | 1.0909 | 1.9970 | 3.3009 | 3.9326 | 3.1267 | 2.6074 | 2.6074 | |
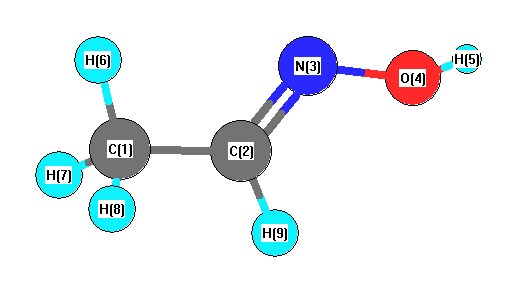 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.618 |
|
C1 |
C2 |
H9 |
118.914 |
| C2 |
C1 |
H6 |
110.641 |
|
C2 |
C1 |
H7 |
110.128 |
| C2 |
C1 |
H8 |
110.128 |
|
C2 |
N3 |
O4 |
110.441 |
| N3 |
C2 |
H9 |
112.468 |
|
N3 |
O4 |
H5 |
101.318 |
| H6 |
C1 |
H7 |
109.131 |
|
H6 |
C1 |
H8 |
109.131 |
| H7 |
C1 |
H8 |
107.619 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
