return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(NH2)H2CH2CH2CH3 (1-Butanamine)

using model chemistry: MP2=FULL/6-31G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G
 hartrees
Energy at 0K-212.713353
Energy at 298.15K-212.726028
Nuclear repulsion energy183.457269
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3530 3354 0.44      
2 A' 3139 2983 41.21      
3 A' 3059 2906 57.09      
4 A' 3054 2901 31.17      
5 A' 3041 2889 30.35      
6 A' 3025 2874 15.06      
7 A' 1748 1660 26.66      
8 A' 1583 1504 4.24      
9 A' 1572 1493 0.69      
10 A' 1564 1486 0.44      
11 A' 1555 1478 0.58      
12 A' 1485 1411 2.51      
13 A' 1438 1366 4.49      
14 A' 1428 1357 2.07      
15 A' 1345 1278 1.91      
16 A' 1170 1112 6.99      
17 A' 1107 1051 7.82      
18 A' 1091 1037 3.33      
19 A' 1021 970 20.20      
20 A' 930 884 6.16      
21 A' 738 701 320.82      
22 A' 436 415 8.26      
23 A' 399 379 0.54      
24 A' 185 176 2.67      
25 A" 3651 3469 0.11      
26 A" 3138 2981 63.02      
27 A" 3114 2959 59.31      
28 A" 3087 2933 18.69      
29 A" 3063 2911 2.34      
30 A" 1574 1495 6.52      
31 A" 1416 1346 0.41      
32 A" 1369 1301 0.19      
33 A" 1355 1288 0.24      
34 A" 1294 1230 0.07      
35 A" 1074 1020 0.07      
36 A" 955 907 0.06      
37 A" 815 774 0.57      
38 A" 753 715 2.77      
39 A" 291 276 61.50      
40 A" 252 239 0.06      
41 A" 125 119 0.36      
42 A" 108 102 3.12      

Unscaled Zero Point Vibrational Energy (zpe) 33537.0 cm-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 31863.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G
ABC
0.59070 0.06307 0.06013

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 2.568 0.376 0.000
C2 1.337 -0.448 0.000
C3 0.000 0.334 0.000
C4 -1.243 -0.585 0.000
C5 -2.575 0.199 0.000
H6 2.680 0.940 0.839
H7 2.680 0.940 -0.839
H8 1.376 -1.104 0.883
H9 1.376 -1.104 -0.883
H10 -0.029 0.993 -0.886
H11 -0.029 0.993 0.886
H12 -1.203 -1.244 0.884
H13 -1.203 -1.244 -0.884
H14 -3.438 -0.482 0.000
H15 -2.648 0.841 0.889
H16 -2.648 0.841 -0.889

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.48072.56793.93025.14591.01711.01712.09492.09492.81222.81224.19784.19786.06685.31105.3110
C21.48071.54952.58443.96592.10562.10561.10041.10042.17472.17472.80492.80494.77584.28204.2820
C32.56791.54951.54642.57892.87282.87282.17762.17761.10421.10422.17222.17223.53382.83882.8388
C43.93022.58441.54641.54554.29214.29212.81262.81262.17912.17911.10281.10282.19742.19052.1905
C55.14593.96592.57891.54555.37335.37334.25344.25342.81032.81032.17822.17821.09891.09971.0997
H61.01712.10562.87284.29215.37331.67772.42472.97353.21212.71024.45524.77646.33715.32895.6019
H71.01712.10562.87284.29215.37331.67772.97352.42472.71023.21214.77644.45526.33715.60195.3289
H82.09491.10042.17762.81264.25342.42472.97351.76593.08222.52422.58303.12934.93424.46944.8080
H92.09491.10042.17762.81264.25342.97352.42471.76592.52423.08223.12932.58304.93424.80804.4694
H102.81222.17471.10422.17912.81033.21212.71023.08222.52421.77183.08402.52593.81843.16722.6228
H112.81222.17471.10422.17912.81032.71023.21212.52423.08221.77182.52593.08403.81842.62283.1672
H124.19782.80492.17221.10282.17824.45524.77642.58303.12933.08402.52591.76712.52152.53673.0949
H134.19782.80492.17221.10282.17824.77644.45523.12932.58302.52593.08401.76712.52153.09492.5367
H146.06684.77583.53382.19741.09896.33716.33714.93424.93423.81843.81842.52152.52151.77951.7795
H155.31104.28202.83882.19051.09975.32895.60194.46944.80803.16722.62282.53673.09491.77951.7789
H165.31104.28202.83882.19051.09975.60195.32894.80804.46942.62283.16723.09492.53671.77951.7789

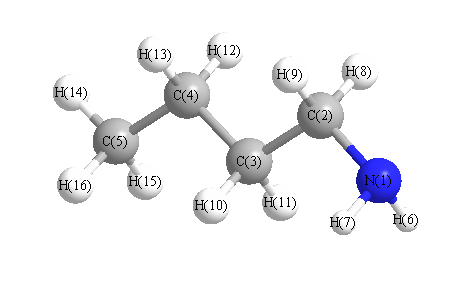
picture of 1-Butanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 115.847 N1 C2 H8 107.596
N1 C2 H9 107.596 C2 N1 H6 113.614
C2 N1 H7 113.614 C2 C3 C4 113.185
C2 C3 H10 108.908 C2 C3 H11 108.908
C3 C2 H8 109.349 C3 C2 H9 109.349
C3 C4 C5 113.043 C3 C4 H12 109.010
C3 C4 H13 109.010 C4 C3 H10 109.467
C4 C3 H11 109.467 C4 C5 H14 111.271
C4 C5 H15 110.676 C4 C5 H16 110.676
C5 C4 H12 109.537 C5 C4 H13 109.537
H6 N1 H7 111.125 H8 C2 H9 106.714
H10 C3 H11 106.698 H12 C4 H13 106.491
H14 C5 H15 108.068 H14 C5 H16 108.068
H15 C5 H16 107.960
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability