Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-31G
Geometric Data calculated at MP2=FULL/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -250.647818 |
| Energy at 298.15K | -250.662183 |
| Nuclear repulsion energy | 255.602627 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3503 |
3328 |
1.27 |
|
|
|
| 2 |
A' |
3116 |
2960 |
67.25 |
|
|
|
| 3 |
A' |
3097 |
2943 |
58.38 |
|
|
|
| 4 |
A' |
3088 |
2934 |
38.11 |
|
|
|
| 5 |
A' |
3049 |
2897 |
43.08 |
|
|
|
| 6 |
A' |
3035 |
2884 |
31.18 |
|
|
|
| 7 |
A' |
3035 |
2883 |
18.81 |
|
|
|
| 8 |
A' |
1570 |
1492 |
4.22 |
|
|
|
| 9 |
A' |
1563 |
1485 |
8.10 |
|
|
|
| 10 |
A' |
1554 |
1476 |
2.47 |
|
|
|
| 11 |
A' |
1438 |
1366 |
1.57 |
|
|
|
| 12 |
A' |
1419 |
1348 |
0.42 |
|
|
|
| 13 |
A' |
1367 |
1298 |
0.36 |
|
|
|
| 14 |
A' |
1313 |
1248 |
6.75 |
|
|
|
| 15 |
A' |
1238 |
1176 |
3.40 |
|
|
|
| 16 |
A' |
1077 |
1023 |
1.73 |
|
|
|
| 17 |
A' |
1038 |
986 |
4.92 |
|
|
|
| 18 |
A' |
932 |
885 |
21.13 |
|
|
|
| 19 |
A' |
873 |
829 |
8.57 |
|
|
|
| 20 |
A' |
827 |
786 |
7.98 |
|
|
|
| 21 |
A' |
712 |
677 |
159.32 |
|
|
|
| 22 |
A' |
565 |
537 |
4.04 |
|
|
|
| 23 |
A' |
444 |
422 |
2.16 |
|
|
|
| 24 |
A' |
389 |
370 |
8.34 |
|
|
|
| 25 |
A' |
242 |
230 |
6.18 |
|
|
|
| 26 |
A" |
3113 |
2958 |
26.10 |
|
|
|
| 27 |
A" |
3088 |
2934 |
61.85 |
|
|
|
| 28 |
A" |
3047 |
2895 |
18.24 |
|
|
|
| 29 |
A" |
3034 |
2883 |
31.50 |
|
|
|
| 30 |
A" |
1558 |
1480 |
0.15 |
|
|
|
| 31 |
A" |
1550 |
1473 |
5.56 |
|
|
|
| 32 |
A" |
1527 |
1451 |
5.10 |
|
|
|
| 33 |
A" |
1426 |
1355 |
1.66 |
|
|
|
| 34 |
A" |
1411 |
1340 |
0.47 |
|
|
|
| 35 |
A" |
1405 |
1335 |
1.74 |
|
|
|
| 36 |
A" |
1335 |
1268 |
0.70 |
|
|
|
| 37 |
A" |
1241 |
1179 |
11.03 |
|
|
|
| 38 |
A" |
1158 |
1100 |
1.13 |
|
|
|
| 39 |
A" |
1125 |
1069 |
13.95 |
|
|
|
| 40 |
A" |
1100 |
1045 |
8.23 |
|
|
|
| 41 |
A" |
984 |
935 |
5.50 |
|
|
|
| 42 |
A" |
884 |
840 |
0.00 |
|
|
|
| 43 |
A" |
830 |
789 |
0.26 |
|
|
|
| 44 |
A" |
459 |
436 |
1.22 |
|
|
|
| 45 |
A" |
242 |
230 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35499.2 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 33727.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.653 |
1.334 |
0.000 |
| H2 |
0.592 |
2.433 |
0.000 |
| H3 |
1.725 |
1.071 |
0.000 |
| C4 |
-0.008 |
0.756 |
1.279 |
| C5 |
-0.008 |
0.756 |
-1.279 |
| C6 |
-0.008 |
-0.795 |
-1.239 |
| C7 |
-0.008 |
-0.795 |
1.239 |
| N8 |
-0.615 |
-1.356 |
0.000 |
| H9 |
-1.631 |
-1.245 |
0.000 |
| H10 |
0.515 |
1.113 |
2.180 |
| H11 |
0.515 |
1.113 |
-2.180 |
| H12 |
-1.051 |
1.113 |
1.345 |
| H13 |
-1.051 |
1.113 |
-1.345 |
| H14 |
1.030 |
-1.161 |
-1.286 |
| H15 |
1.030 |
-1.161 |
1.286 |
| H16 |
-0.544 |
-1.214 |
-2.103 |
| H17 |
-0.544 |
-1.214 |
2.103 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.1009 | 1.1042 | 1.5516 | 1.5516 | 2.5508 | 2.5508 | 2.9734 | 3.4449 | 2.1959 | 2.1959 | 2.1813 | 2.1813 | 2.8320 | 2.8320 | 3.5135 | 3.5135 |
H2 | 1.1009 | | 1.7720 | 2.1933 | 2.1933 | 3.5099 | 3.5099 | 3.9764 | 4.2978 | 2.5501 | 2.5501 | 2.4999 | 2.4999 | 3.8423 | 3.8423 | 4.3604 | 4.3604 | H3 | 1.1042 | 1.7720 | | 2.1772 | 2.1772 | 2.8326 | 2.8326 | 3.3710 | 4.0778 | 2.4939 | 2.4939 | 3.0846 | 3.0846 | 2.6678 | 2.6678 | 3.8458 | 3.8458 | C4 | 1.5516 | 2.1933 | 2.1772 | | 2.5583 | 2.9575 | 1.5515 | 2.5420 | 2.8762 | 1.1019 | 3.5172 | 1.1039 | 2.8457 | 3.3661 | 2.1798 | 3.9500 | 2.2010 | C5 | 1.5516 | 2.1933 | 2.1772 | 2.5583 | | 1.5515 | 2.9575 | 2.5420 | 2.8762 | 3.5172 | 1.1019 | 2.8457 | 1.1039 | 2.1798 | 3.3661 | 2.2010 | 3.9500 | C6 | 2.5508 | 3.5099 | 2.8326 | 2.9575 | 1.5515 | | 2.4780 | 1.4889 | 2.0907 | 3.9509 | 2.1916 | 3.3768 | 2.1769 | 1.1021 | 2.7545 | 1.0990 | 3.4100 | C7 | 2.5508 | 3.5099 | 2.8326 | 1.5515 | 2.9575 | 2.4780 | | 1.4889 | 2.0907 | 2.1916 | 3.9509 | 2.1769 | 3.3768 | 2.7545 | 1.1021 | 3.4100 | 1.0990 | N8 | 2.9734 | 3.9764 | 3.3710 | 2.5420 | 2.5420 | 1.4889 | 1.4889 | | 1.0225 | 3.4823 | 3.4823 | 2.8446 | 2.8446 | 2.0969 | 2.0969 | 2.1086 | 2.1086 | H9 | 3.4449 | 4.2978 | 4.0778 | 2.8762 | 2.8762 | 2.0907 | 2.0907 | 1.0225 | | 3.8630 | 3.8630 | 2.7756 | 2.7756 | 2.9570 | 2.9570 | 2.3673 | 2.3673 | H10 | 2.1959 | 2.5501 | 2.4939 | 1.1019 | 3.5172 | 3.9509 | 2.1916 | 3.4823 | 3.8630 | | 4.3609 | 1.7751 | 3.8572 | 4.1775 | 2.4974 | 4.9882 | 2.5580 | H11 | 2.1959 | 2.5501 | 2.4939 | 3.5172 | 1.1019 | 2.1916 | 3.9509 | 3.4823 | 3.8630 | 4.3609 | | 3.8572 | 1.7751 | 2.4974 | 4.1775 | 2.5580 | 4.9882 | H12 | 2.1813 | 2.4999 | 3.0846 | 1.1039 | 2.8457 | 3.3768 | 2.1769 | 2.8446 | 2.7756 | 1.7751 | 3.8572 | | 2.6892 | 4.0520 | 3.0828 | 4.1897 | 2.4991 | H13 | 2.1813 | 2.4999 | 3.0846 | 2.8457 | 1.1039 | 2.1769 | 3.3768 | 2.8446 | 2.7756 | 3.8572 | 1.7751 | 2.6892 | | 3.0828 | 4.0520 | 2.4991 | 4.1897 | H14 | 2.8320 | 3.8423 | 2.6678 | 3.3661 | 2.1798 | 1.1021 | 2.7545 | 2.0969 | 2.9570 | 4.1775 | 2.4974 | 4.0520 | 3.0828 | | 2.5716 | 1.7742 | 3.7366 | H15 | 2.8320 | 3.8423 | 2.6678 | 2.1798 | 3.3661 | 2.7545 | 1.1021 | 2.0969 | 2.9570 | 2.4974 | 4.1775 | 3.0828 | 4.0520 | 2.5716 | | 3.7366 | 1.7742 | H16 | 3.5135 | 4.3604 | 3.8458 | 3.9500 | 2.2010 | 1.0990 | 3.4100 | 2.1086 | 2.3673 | 4.9882 | 2.5580 | 4.1897 | 2.4991 | 1.7742 | 3.7366 | | 4.2052 | H17 | 3.5135 | 4.3604 | 3.8458 | 2.2010 | 3.9500 | 3.4100 | 1.0990 | 2.1086 | 2.3673 | 2.5580 | 4.9882 | 2.4991 | 4.1897 | 3.7366 | 1.7742 | 4.2052 | |
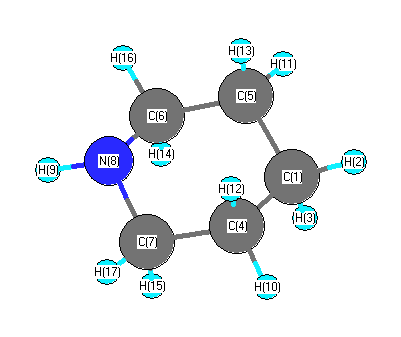 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.570 |
|
C1 |
C4 |
H10 |
110.547 |
| C1 |
C4 |
H12 |
109.299 |
|
C1 |
C5 |
C6 |
110.570 |
| C1 |
C5 |
H11 |
110.547 |
|
C1 |
C5 |
H13 |
109.299 |
| H2 |
C1 |
H3 |
106.948 |
|
H2 |
C1 |
C4 |
110.406 |
| H2 |
C1 |
C5 |
110.406 |
|
H3 |
C1 |
C4 |
108.960 |
| H3 |
C1 |
C5 |
108.960 |
|
C4 |
C1 |
C5 |
111.051 |
| C4 |
C7 |
N8 |
113.439 |
|
C4 |
C7 |
H15 |
109.291 |
| C4 |
C7 |
H17 |
111.128 |
|
C5 |
C6 |
N8 |
113.439 |
| C5 |
C6 |
H14 |
109.291 |
|
C5 |
C6 |
H16 |
111.128 |
| C6 |
C5 |
H11 |
110.214 |
|
C6 |
C5 |
H13 |
108.961 |
| C6 |
N8 |
C7 |
112.646 |
|
C6 |
N8 |
H9 |
111.352 |
| C7 |
C4 |
H10 |
110.214 |
|
C7 |
C4 |
H12 |
108.961 |
| C7 |
N8 |
H9 |
111.352 |
|
N8 |
C6 |
H14 |
107.117 |
| N8 |
C6 |
H16 |
108.190 |
|
N8 |
C7 |
H15 |
107.117 |
| N8 |
C7 |
H17 |
108.190 |
|
H10 |
C4 |
H12 |
107.168 |
| H11 |
C5 |
H13 |
107.168 |
|
H14 |
C6 |
H16 |
107.430 |
| H15 |
C7 |
H17 |
107.430 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
