Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -208.283324 |
| Energy at 298.15K | -208.289418 |
| HF Energy | -207.858591 |
| Nuclear repulsion energy | 117.127406 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3624 |
3443 |
28.31 |
|
|
|
| 2 |
A' |
3161 |
3004 |
21.21 |
|
|
|
| 3 |
A' |
3064 |
2911 |
48.64 |
|
|
|
| 4 |
A' |
3037 |
2886 |
79.45 |
|
|
|
| 5 |
A' |
1712 |
1627 |
320.99 |
|
|
|
| 6 |
A' |
1598 |
1518 |
13.66 |
|
|
|
| 7 |
A' |
1530 |
1454 |
10.32 |
|
|
|
| 8 |
A' |
1494 |
1419 |
3.15 |
|
|
|
| 9 |
A' |
1445 |
1373 |
21.30 |
|
|
|
| 10 |
A' |
1335 |
1269 |
101.22 |
|
|
|
| 11 |
A' |
1178 |
1119 |
48.86 |
|
|
|
| 12 |
A' |
1026 |
975 |
42.35 |
|
|
|
| 13 |
A' |
600 |
570 |
13.33 |
|
|
|
| 14 |
A' |
348 |
330 |
7.61 |
|
|
|
| 15 |
A" |
3139 |
2982 |
34.58 |
|
|
|
| 16 |
A" |
1557 |
1479 |
6.37 |
|
|
|
| 17 |
A" |
1189 |
1129 |
2.53 |
|
|
|
| 18 |
A" |
1030 |
979 |
1.64 |
|
|
|
| 19 |
A" |
669 |
636 |
203.72 |
|
|
|
| 20 |
A" |
204 |
194 |
0.95 |
|
|
|
| 21 |
A" |
109 |
103 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16523.5 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 15699.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.275 |
-0.765 |
0.000 |
| O2 |
1.433 |
-1.264 |
0.000 |
| N3 |
0.000 |
0.585 |
0.000 |
| C4 |
-1.356 |
1.154 |
0.000 |
| H5 |
-0.634 |
-1.389 |
0.000 |
| H6 |
0.800 |
1.209 |
0.000 |
| H7 |
-2.078 |
0.329 |
0.000 |
| H8 |
-1.532 |
1.766 |
0.894 |
| H9 |
-1.532 |
1.766 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2609 | 1.3773 | 2.5183 | 1.1030 | 2.0425 | 2.5951 | 3.2356 | 3.2356 |
O2 | 1.2609 | | 2.3390 | 3.6910 | 2.0707 | 2.5528 | 3.8556 | 4.3324 | 4.3324 | N3 | 1.3773 | 2.3390 | | 1.4705 | 2.0736 | 1.0146 | 2.0942 | 2.1311 | 2.1311 | C4 | 2.5183 | 3.6910 | 1.4705 | | 2.6441 | 2.1562 | 1.0971 | 1.0978 | 1.0978 | H5 | 1.1030 | 2.0707 | 2.0736 | 2.6441 | | 2.9680 | 2.2447 | 3.4005 | 3.4005 | H6 | 2.0425 | 2.5528 | 1.0146 | 2.1562 | 2.9680 | | 3.0098 | 2.5585 | 2.5585 | H7 | 2.5951 | 3.8556 | 2.0942 | 1.0971 | 2.2447 | 3.0098 | | 1.7789 | 1.7789 | H8 | 3.2356 | 4.3324 | 2.1311 | 1.0978 | 3.4005 | 2.5585 | 1.7789 | | 1.7885 | H9 | 3.2356 | 4.3324 | 2.1311 | 1.0978 | 3.4005 | 2.5585 | 1.7789 | 1.7885 | |
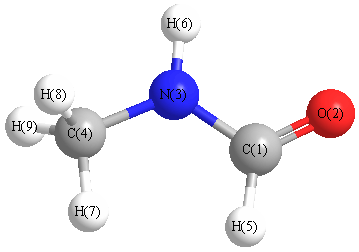 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.301 |
|
C1 |
N3 |
H6 |
116.463 |
| O2 |
C1 |
N3 |
124.838 |
|
O2 |
C1 |
H5 |
122.181 |
| N3 |
C1 |
H5 |
112.980 |
|
N3 |
C4 |
H7 |
108.420 |
| N3 |
C4 |
H8 |
111.328 |
|
N3 |
C4 |
H9 |
111.328 |
| C4 |
N3 |
H6 |
119.236 |
|
H7 |
C4 |
H8 |
108.284 |
| H7 |
C4 |
H9 |
108.284 |
|
H8 |
C4 |
H9 |
109.098 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -208.283324 |
| Energy at 298.15K | -208.289418 |
| HF Energy | -207.858591 |
| Nuclear repulsion energy | 117.127406 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -208.285117 |
| Energy at 298.15K | -208.291076 |
| HF Energy | -207.860637 |
| Nuclear repulsion energy | 119.588627 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3664 |
3481 |
35.56 |
|
|
|
| 2 |
A |
3186 |
3027 |
13.92 |
|
|
|
| 3 |
A |
3155 |
2998 |
26.06 |
|
|
|
| 4 |
A |
3075 |
2922 |
24.59 |
|
|
|
| 5 |
A |
3052 |
2899 |
110.05 |
|
|
|
| 6 |
A |
1687 |
1603 |
196.43 |
|
|
|
| 7 |
A |
1598 |
1518 |
42.23 |
|
|
|
| 8 |
A |
1571 |
1493 |
60.64 |
|
|
|
| 9 |
A |
1553 |
1475 |
29.49 |
|
|
|
| 10 |
A |
1513 |
1437 |
29.92 |
|
|
|
| 11 |
A |
1432 |
1361 |
12.93 |
|
|
|
| 12 |
A |
1255 |
1192 |
61.78 |
|
|
|
| 13 |
A |
1203 |
1143 |
1.51 |
|
|
|
| 14 |
A |
1198 |
1138 |
4.75 |
|
|
|
| 15 |
A |
1010 |
959 |
0.42 |
|
|
|
| 16 |
A |
962 |
914 |
12.50 |
|
|
|
| 17 |
A |
771 |
732 |
4.34 |
|
|
|
| 18 |
A |
499 |
474 |
94.56 |
|
|
|
| 19 |
A |
329 |
312 |
93.15 |
|
|
|
| 20 |
A |
280 |
266 |
11.61 |
|
|
|
| 21 |
A |
15 |
14 |
2.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16502.4 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 15678.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.868 |
0.460 |
0.001 |
| O2 |
1.408 |
-0.683 |
0.001 |
| N3 |
-0.489 |
0.669 |
-0.004 |
| C4 |
-1.438 |
-0.459 |
0.001 |
| H5 |
1.461 |
1.389 |
-0.002 |
| H6 |
-0.841 |
1.616 |
0.030 |
| H7 |
-2.431 |
-0.084 |
-0.268 |
| H8 |
-1.118 |
-1.207 |
-0.734 |
| H9 |
-1.485 |
-0.941 |
0.987 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2637 | 1.3735 | 2.4829 | 1.1017 | 2.0639 | 3.3545 | 2.6947 | 2.9107 |
O2 | 1.2637 | | 2.3296 | 2.8551 | 2.0718 | 3.2163 | 3.8946 | 2.6823 | 3.0673 | N3 | 1.3735 | 2.3296 | | 1.4743 | 2.0789 | 1.0111 | 2.0993 | 2.1083 | 2.1365 | C4 | 2.4829 | 2.8551 | 1.4743 | | 3.4381 | 2.1601 | 1.0948 | 1.0956 | 1.0980 | H5 | 1.1017 | 2.0718 | 2.0789 | 3.4381 | | 2.3135 | 4.1699 | 3.7311 | 3.8836 | H6 | 2.0639 | 3.2163 | 1.0111 | 2.1601 | 2.3135 | | 2.3473 | 2.9376 | 2.8050 | H7 | 3.3545 | 3.8946 | 2.0993 | 1.0948 | 4.1699 | 2.3473 | | 1.7887 | 1.7897 | H8 | 2.6947 | 2.6823 | 2.1083 | 1.0956 | 3.7311 | 2.9376 | 1.7887 | | 1.7789 | H9 | 2.9107 | 3.0673 | 2.1365 | 1.0980 | 3.8836 | 2.8050 | 1.7897 | 1.7789 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.307 |
|
C1 |
N3 |
H6 |
119.092 |
| O2 |
C1 |
N3 |
124.046 |
|
O2 |
C1 |
H5 |
122.149 |
| N3 |
C1 |
H5 |
113.804 |
|
N3 |
C4 |
H7 |
108.698 |
| N3 |
C4 |
H8 |
109.358 |
|
N3 |
C4 |
H9 |
111.482 |
| C4 |
N3 |
H6 |
119.545 |
|
H7 |
C4 |
H8 |
109.492 |
| H7 |
C4 |
H9 |
109.415 |
|
H8 |
C4 |
H9 |
108.375 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
