Jump to
S1C2
S1C3
Energy calculated at B3PW91/6-31G
| | hartrees |
|---|
| Energy at 0K | -166.404229 |
| Energy at 298.15K | |
| HF Energy | -166.404229 |
| Nuclear repulsion energy | 49.459715 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G
Geometric Data calculated at B3PW91/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at B3PW91/6-31G
| | hartrees |
|---|
| Energy at 0K | -166.404175 |
| Energy at 298.15K | |
| HF Energy | -166.404175 |
| Nuclear repulsion energy | 49.615148 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G
Geometric Data calculated at B3PW91/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B3PW91/6-31G
| | hartrees |
|---|
| Energy at 0K | -166.404229 |
| Energy at 298.15K | -166.404789 |
| HF Energy | -166.404229 |
| Nuclear repulsion energy | 49.457544 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
4110 |
3937 |
0.00 |
|
|
|
| 2 |
Σg |
789 |
755 |
0.00 |
|
|
|
| 3 |
Σu |
4109 |
3935 |
364.95 |
|
|
|
| 4 |
Σu |
1687 |
1615 |
431.39 |
|
|
|
| 5 |
Πg |
153 |
146 |
0.00 |
|
|
|
| 5 |
Πg |
153 |
146 |
0.00 |
|
|
|
| 6 |
Πu |
403 |
386 |
104.21 |
|
|
|
| 6 |
Πu |
403 |
386 |
104.21 |
|
|
|
| 7 |
Πu |
169 |
162 |
494.11 |
|
|
|
| 7 |
Πu |
169 |
162 |
494.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6072.6 cm
-1
Scaled (by 0.9577) Zero Point Vibrational Energy (zpe) 5815.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
0.000 |
| O2 |
0.000 |
0.000 |
1.398 |
| O3 |
0.000 |
0.000 |
-1.398 |
| H4 |
0.000 |
0.000 |
2.345 |
| H5 |
0.000 |
0.000 |
-2.345 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.3981 | 1.3981 | 2.3450 | 2.3450 |
O2 | 1.3981 | | 2.7963 | 0.9468 | 3.7431 | O3 | 1.3981 | 2.7963 | | 3.7431 | 0.9468 | H4 | 2.3450 | 0.9468 | 3.7431 | | 4.6899 | H5 | 2.3450 | 3.7431 | 0.9468 | 4.6899 | |
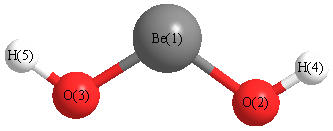 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
180.000 |
|
Be1 |
O3 |
H5 |
180.000 |
| O2 |
Be1 |
O3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.667 |
|
|
|
| 2 |
O |
-0.734 |
|
|
|
| 3 |
O |
-0.734 |
|
|
|
| 4 |
H |
0.401 |
|
|
|
| 5 |
H |
0.401 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.346 |
0.000 |
0.000 |
| y |
0.000 |
-16.346 |
0.000 |
| z |
0.000 |
0.000 |
-7.064 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.641 |
0.000 |
0.000 |
| y |
0.000 |
-4.641 |
0.000 |
| z |
0.000 |
0.000 |
9.282 |
|
| Polar |
|---|
| 3z2-r2 | 18.563 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.265 |
0.000 |
0.000 |
| y |
0.000 |
1.265 |
0.000 |
| z |
0.000 |
0.000 |
3.643 |
<r2> (average value of r
2) Å
2
| <r2> |
50.551 |
| (<r2>)1/2 |
7.110 |
