Jump to
S1C2
Energy calculated at B3LYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -454.665413 |
| Energy at 298.15K | -454.668992 |
| HF Energy | -454.665413 |
| Nuclear repulsion energy | 55.504970 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3545 |
3410 |
6.65 |
|
|
|
| 2 |
A' |
2450 |
2357 |
80.42 |
|
|
|
| 3 |
A' |
1648 |
1585 |
16.41 |
|
|
|
| 4 |
A' |
964 |
928 |
8.14 |
|
|
|
| 5 |
A' |
813 |
782 |
65.66 |
|
|
|
| 6 |
A' |
425 |
409 |
149.89 |
|
|
|
| 7 |
A" |
3692 |
3552 |
14.61 |
|
|
|
| 8 |
A" |
1099 |
1057 |
4.07 |
|
|
|
| 9 |
A" |
472 |
454 |
80.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7554.1 cm
-1
Scaled (by 0.962) Zero Point Vibrational Energy (zpe) 7267.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.023 |
1.160 |
0.000 |
| S2 |
0.023 |
-0.649 |
0.000 |
| H3 |
-1.350 |
-0.823 |
0.000 |
| H4 |
0.413 |
1.543 |
0.852 |
| H5 |
0.413 |
1.543 |
-0.852 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.8084 | 2.4111 | 1.0126 | 1.0126 |
S2 | 1.8084 | | 1.3837 | 2.3836 | 2.3836 | H3 | 2.4111 | 1.3837 | | 3.0706 | 3.0706 | H4 | 1.0126 | 2.3836 | 3.0706 | | 1.7045 | H5 | 1.0126 | 2.3836 | 3.0706 | 1.7045 | |
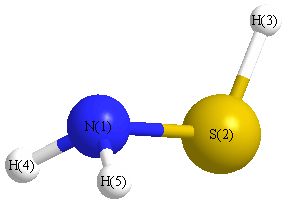 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
97.215 |
|
S2 |
N1 |
H4 |
112.238 |
| S2 |
N1 |
H5 |
112.238 |
|
H4 |
N1 |
H5 |
114.630 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.838 |
|
|
|
| 2 |
S |
0.133 |
|
|
|
| 3 |
H |
0.067 |
|
|
|
| 4 |
H |
0.319 |
|
|
|
| 5 |
H |
0.319 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.447 |
0.922 |
0.000 |
1.025 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.675 |
4.107 |
0.000 |
| y |
4.107 |
-18.739 |
0.000 |
| z |
0.000 |
0.000 |
-20.064 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.273 |
4.107 |
0.000 |
| y |
4.107 |
1.130 |
0.000 |
| z |
0.000 |
0.000 |
-0.857 |
|
| Polar |
|---|
| 3z2-r2 | -1.714 |
|---|
| x2-y2 | -0.936 |
|---|
| xy | 4.107 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.957 |
0.639 |
0.000 |
| y |
0.639 |
4.292 |
0.000 |
| z |
0.000 |
0.000 |
2.322 |
<r2> (average value of r
2) Å
2
| <r2> |
37.387 |
| (<r2>)1/2 |
6.115 |
Jump to
S1C1
Energy calculated at B3LYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -454.665413 |
| Energy at 298.15K | -454.668987 |
| HF Energy | -454.665413 |
| Nuclear repulsion energy | 55.518330 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3547 |
3412 |
6.76 |
|
|
|
| 2 |
A' |
2451 |
2357 |
80.98 |
|
|
|
| 3 |
A' |
1647 |
1584 |
16.32 |
|
|
|
| 4 |
A' |
965 |
928 |
7.94 |
|
|
|
| 5 |
A' |
811 |
781 |
64.83 |
|
|
|
| 6 |
A' |
419 |
403 |
154.05 |
|
|
|
| 7 |
A" |
3695 |
3555 |
14.96 |
|
|
|
| 8 |
A" |
1098 |
1057 |
4.19 |
|
|
|
| 9 |
A" |
473 |
455 |
80.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7553.0 cm
-1
Scaled (by 0.962) Zero Point Vibrational Energy (zpe) 7266.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.023 |
1.159 |
0.000 |
| S2 |
0.023 |
-0.649 |
0.000 |
| H3 |
-1.350 |
-0.823 |
0.000 |
| H4 |
0.410 |
1.544 |
0.853 |
| H5 |
0.410 |
1.544 |
-0.853 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.8077 | 2.4108 | 1.0124 | 1.0124 |
S2 | 1.8077 | | 1.3836 | 2.3840 | 2.3840 | H3 | 2.4108 | 1.3836 | | 3.0698 | 3.0698 | H4 | 1.0124 | 2.3840 | 3.0698 | | 1.7054 | H5 | 1.0124 | 2.3840 | 3.0698 | 1.7054 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
97.232 |
|
S2 |
N1 |
H4 |
112.329 |
| S2 |
N1 |
H5 |
112.329 |
|
H4 |
N1 |
H5 |
114.770 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.839 |
|
|
|
| 2 |
S |
0.134 |
|
|
|
| 3 |
H |
0.067 |
|
|
|
| 4 |
H |
0.319 |
|
|
|
| 5 |
H |
0.319 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.437 |
0.927 |
0.000 |
1.025 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.686 |
4.090 |
0.000 |
| y |
4.090 |
-18.720 |
0.000 |
| z |
0.000 |
0.000 |
-20.057 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.298 |
4.090 |
0.000 |
| y |
4.090 |
1.151 |
0.000 |
| z |
0.000 |
0.000 |
-0.853 |
|
| Polar |
|---|
| 3z2-r2 | -1.707 |
|---|
| x2-y2 | -0.966 |
|---|
| xy | 4.090 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.954 |
0.638 |
0.000 |
| y |
0.638 |
4.287 |
0.000 |
| z |
0.000 |
0.000 |
2.321 |
<r2> (average value of r
2) Å
2
| <r2> |
37.375 |
| (<r2>)1/2 |
6.113 |
