Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B2PLYP/6-31G
Geometric Data calculated at B2PLYP/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B2PLYP/6-31G
Geometric Data calculated at B2PLYP/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -210.012700 |
| Energy at 298.15K | -210.021837 |
| HF Energy | -209.873270 |
| Nuclear repulsion energy | 133.476124 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3677 |
3677 |
3.91 |
|
|
|
| 2 |
A |
3558 |
3558 |
0.09 |
|
|
|
| 3 |
A |
3537 |
3537 |
38.25 |
|
|
|
| 4 |
A |
3145 |
3145 |
44.03 |
|
|
|
| 5 |
A |
3124 |
3124 |
28.14 |
|
|
|
| 6 |
A |
3022 |
3022 |
93.30 |
|
|
|
| 7 |
A |
3009 |
3009 |
54.94 |
|
|
|
| 8 |
A |
1730 |
1730 |
33.23 |
|
|
|
| 9 |
A |
1576 |
1576 |
0.30 |
|
|
|
| 10 |
A |
1558 |
1558 |
3.24 |
|
|
|
| 11 |
A |
1453 |
1453 |
64.80 |
|
|
|
| 12 |
A |
1448 |
1448 |
5.58 |
|
|
|
| 13 |
A |
1399 |
1399 |
15.81 |
|
|
|
| 14 |
A |
1354 |
1354 |
4.96 |
|
|
|
| 15 |
A |
1273 |
1273 |
25.55 |
|
|
|
| 16 |
A |
1198 |
1198 |
17.34 |
|
|
|
| 17 |
A |
1113 |
1113 |
12.98 |
|
|
|
| 18 |
A |
1043 |
1043 |
57.44 |
|
|
|
| 19 |
A |
1016 |
1016 |
2.69 |
|
|
|
| 20 |
A |
905 |
905 |
19.38 |
|
|
|
| 21 |
A |
889 |
889 |
37.55 |
|
|
|
| 22 |
A |
732 |
732 |
247.74 |
|
|
|
| 23 |
A |
621 |
621 |
164.48 |
|
|
|
| 24 |
A |
544 |
544 |
16.23 |
|
|
|
| 25 |
A |
328 |
328 |
0.37 |
|
|
|
| 26 |
A |
267 |
267 |
10.48 |
|
|
|
| 27 |
A |
185 |
185 |
8.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21853.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21853.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.363 |
-0.544 |
0.130 |
| C2 |
-0.636 |
0.677 |
-0.268 |
| C3 |
0.800 |
0.582 |
0.261 |
| O4 |
1.393 |
-0.671 |
-0.164 |
| H5 |
-1.669 |
-0.560 |
1.097 |
| H6 |
-2.097 |
-0.835 |
-0.501 |
| H7 |
-1.111 |
1.610 |
0.075 |
| H8 |
-0.606 |
0.700 |
-1.361 |
| H9 |
1.423 |
1.385 |
-0.141 |
| H10 |
0.800 |
0.662 |
1.359 |
| H11 |
0.671 |
-1.337 |
-0.088 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4759 | 2.4428 | 2.7748 | 1.0137 | 1.0114 | 2.1696 | 2.0848 | 3.3996 | 2.7649 | 2.1940 |
C2 | 1.4759 | | 1.5337 | 2.4388 | 2.1116 | 2.1159 | 1.1013 | 1.0941 | 2.1813 | 2.1704 | 2.4078 | C3 | 2.4428 | 1.5337 | | 1.4504 | 2.8461 | 3.3149 | 2.1777 | 2.1499 | 1.0920 | 1.1013 | 1.9552 | O4 | 2.7748 | 2.4388 | 1.4504 | | 3.3128 | 3.5107 | 3.3957 | 2.7038 | 2.0566 | 2.1092 | 0.9851 | H5 | 1.0137 | 2.1116 | 2.8461 | 3.3128 | | 1.6774 | 2.4629 | 2.9598 | 3.8565 | 2.7672 | 2.7348 | H6 | 1.0114 | 2.1159 | 3.3149 | 3.5107 | 1.6774 | | 2.6992 | 2.3070 | 4.1777 | 3.7550 | 2.8436 | H7 | 2.1696 | 1.1013 | 2.1777 | 3.3957 | 2.4629 | 2.6992 | | 1.7737 | 2.5528 | 2.4898 | 3.4475 | H8 | 2.0848 | 1.0941 | 2.1499 | 2.7038 | 2.9598 | 2.3070 | 1.7737 | | 2.4648 | 3.0625 | 2.7204 | H9 | 3.3996 | 2.1813 | 1.0920 | 2.0566 | 3.8565 | 4.1777 | 2.5528 | 2.4648 | | 1.7774 | 2.8244 | H10 | 2.7649 | 2.1704 | 1.1013 | 2.1092 | 2.7672 | 3.7550 | 2.4898 | 3.0625 | 1.7774 | | 2.4712 | H11 | 2.1940 | 2.4078 | 1.9552 | 0.9851 | 2.7348 | 2.8436 | 3.4475 | 2.7204 | 2.8244 | 2.4712 | |
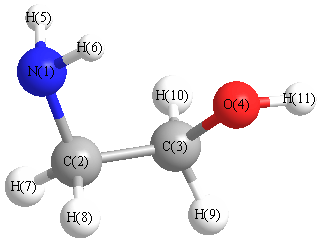 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
108.502 |
|
N1 |
C2 |
H7 |
113.880 |
| N1 |
C2 |
H8 |
107.498 |
|
C2 |
N1 |
H5 |
114.758 |
| C2 |
N1 |
H6 |
115.301 |
|
C2 |
C3 |
O4 |
109.590 |
| C2 |
C3 |
H9 |
111.234 |
|
C2 |
C3 |
H10 |
109.820 |
| C3 |
C2 |
H7 |
110.395 |
|
C3 |
C2 |
H8 |
108.635 |
| C3 |
O4 |
H11 |
105.188 |
|
O4 |
C3 |
H9 |
107.139 |
| O4 |
C3 |
H10 |
110.758 |
|
H5 |
N1 |
H6 |
111.844 |
| H7 |
C2 |
H8 |
107.778 |
|
H9 |
C3 |
H10 |
108.267 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.751 |
|
|
|
| 2 |
C |
-0.148 |
|
|
|
| 3 |
C |
0.020 |
|
|
|
| 4 |
O |
-0.646 |
|
|
|
| 5 |
H |
0.296 |
|
|
|
| 6 |
H |
0.303 |
|
|
|
| 7 |
H |
0.123 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.115 |
|
|
|
| 11 |
H |
0.381 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.515 |
1.132 |
0.601 |
3.742 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.318 |
3.109 |
0.646 |
| y |
3.109 |
-25.026 |
-0.266 |
| z |
0.646 |
-0.266 |
-24.416 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.597 |
3.109 |
0.646 |
| y |
3.109 |
0.841 |
-0.266 |
| z |
0.646 |
-0.266 |
1.756 |
|
| Polar |
|---|
| 3z2-r2 | 3.511 |
|---|
| x2-y2 | -2.292 |
|---|
| xy | 3.109 |
|---|
| xz | 0.646 |
|---|
| yz | -0.266 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.832 |
0.331 |
-0.059 |
| y |
0.331 |
4.877 |
0.106 |
| z |
-0.059 |
0.106 |
4.340 |
<r2> (average value of r
2) Å
2
| <r2> |
87.748 |
| (<r2>)1/2 |
9.367 |
