Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -254.336893 |
| Energy at 298.15K | -254.336402 |
| HF Energy | -254.112254 |
| Nuclear repulsion energy | 45.766122 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.670 |
| N2 |
0.000 |
0.000 |
-1.884 |
| Na3 |
0.000 |
0.000 |
1.564 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.2135 | 2.2343 |
N2 | 1.2135 | | 3.4477 | Na3 | 2.2343 | 3.4477 | |
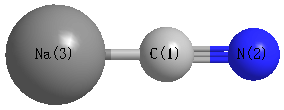 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
0.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -254.332882 |
| Energy at 298.15K | -254.332537 |
| HF Energy | -254.111553 |
| Nuclear repulsion energy | 50.391473 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.100 |
0.618 |
0.000 |
| N2 |
0.000 |
1.156 |
0.000 |
| Na3 |
-0.600 |
-1.072 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.2241 | 2.3965 |
N2 | 1.2241 | | 2.3072 | Na3 | 2.3965 | 2.3072 | |
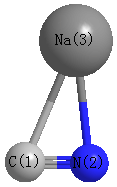 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
78.992 |
|
C1 |
Na3 |
N2 |
30.092 |
| N2 |
C1 |
Na3 |
70.915 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -254.331289 |
| Energy at 298.15K | -254.330639 |
| HF Energy | -254.121908 |
| Nuclear repulsion energy | 48.012543 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.882 |
| N2 |
0.000 |
0.000 |
-0.666 |
| Na3 |
0.000 |
0.000 |
1.450 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.2158 | 3.3324 |
N2 | 1.2158 | | 2.1165 | Na3 | 3.3324 | 2.1165 | |
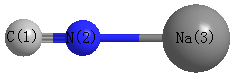 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
180.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
