Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -691.609605 |
| Energy at 298.15K | -691.609079 |
| HF Energy | -691.387945 |
| Nuclear repulsion energy | 59.080507 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.348 |
| C2 |
0.000 |
0.000 |
-1.315 |
| N3 |
0.000 |
0.000 |
-2.531 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.6632 | 3.8788 |
C2 | 2.6632 | | 1.2156 | N3 | 3.8788 | 1.2156 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -691.606394 |
| Energy at 298.15K | -691.605609 |
| HF Energy | -691.397202 |
| Nuclear repulsion energy | 62.316384 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.252 |
| C2 |
0.000 |
0.000 |
-2.486 |
| N3 |
0.000 |
0.000 |
-1.269 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.7381 | 2.5209 |
C2 | 3.7381 | | 1.2172 | N3 | 2.5209 | 1.2172 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -691.610405 |
| Energy at 298.15K | -691.610214 |
| HF Energy | -691.391307 |
| Nuclear repulsion energy | 65.808098 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.087 |
0.000 |
| C2 |
0.658 |
-1.619 |
0.000 |
| N3 |
-0.564 |
-1.563 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.7847 | 2.7100 |
C2 | 2.7847 | | 1.2229 | N3 | 2.7100 | 1.2229 | |
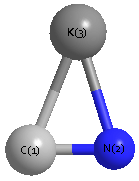 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
73.749 |
|
K1 |
N3 |
C2 |
80.577 |
| C2 |
K1 |
N3 |
25.674 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
