Jump to
S1C2
S1C3
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -100.212182 |
| Energy at 298.15K | -100.211330 |
| HF Energy | -100.135354 |
| Nuclear repulsion energy | 27.174177 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.078 |
| C2 |
0.000 |
0.000 |
-0.163 |
| N3 |
0.000 |
0.000 |
1.030 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.9145 | 3.1079 |
C2 | 1.9145 | | 1.1934 | N3 | 3.1079 | 1.1934 | |
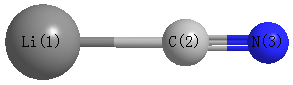 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.545 |
|
|
|
| 2 |
C |
-0.172 |
|
|
|
| 3 |
N |
-0.373 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-8.845 |
8.845 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.278 |
0.000 |
0.000 |
| y |
0.000 |
-14.278 |
0.000 |
| z |
0.000 |
0.000 |
0.864 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.571 |
0.000 |
0.000 |
| y |
0.000 |
-7.571 |
0.000 |
| z |
0.000 |
0.000 |
15.143 |
|
| Polar |
|---|
| 3z2-r2 | 30.285 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.120 |
0.000 |
0.000 |
| y |
0.000 |
2.120 |
0.000 |
| z |
0.000 |
0.000 |
4.362 |
<r2> (average value of r
2) Å
2
| <r2> |
26.305 |
| (<r2>)1/2 |
5.129 |
Jump to
S1C1
S1C3
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -100.211397 |
| Energy at 298.15K | |
| HF Energy | -100.137162 |
| Nuclear repulsion energy | 27.064207 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
1.470 |
-0.561 |
0.000 |
| C2 |
-0.735 |
-0.383 |
0.000 |
| N3 |
0.000 |
0.568 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.2115 | 1.8533 |
C2 | 2.2115 | | 1.2017 | N3 | 1.8533 | 1.2017 | |
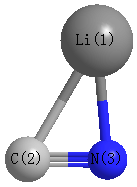 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
56.934 |
|
Li1 |
N3 |
C2 |
90.153 |
| C2 |
Li1 |
N3 |
32.913 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.632 |
|
|
|
| 2 |
C |
-0.036 |
|
|
|
| 3 |
N |
-0.595 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.556 |
8.681 |
0.000 |
8.819 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.874 |
-2.385 |
0.000 |
| y |
-2.385 |
-1.750 |
0.000 |
| z |
0.000 |
0.000 |
-14.319 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.840 |
-2.385 |
0.000 |
| y |
-2.385 |
12.346 |
0.000 |
| z |
0.000 |
0.000 |
-6.507 |
|
| Polar |
|---|
| 3z2-r2 | -13.013 |
|---|
| x2-y2 | -12.124 |
|---|
| xy | -2.385 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.120 |
0.000 |
0.000 |
| y |
0.000 |
2.120 |
0.000 |
| z |
0.000 |
0.000 |
4.362 |
<r2> (average value of r
2) Å
2
| <r2> |
25.386 |
| (<r2>)1/2 |
5.038 |
Jump to
S1C1
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -100.214329 |
| Energy at 298.15K | -100.213351 |
| HF Energy | -100.143124 |
| Nuclear repulsion energy | 27.925938 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
1.895 |
| C2 |
0.000 |
0.000 |
-1.085 |
| N3 |
0.000 |
0.000 |
0.118 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.9799 | 1.7770 |
C2 | 2.9799 | | 1.2029 | N3 | 1.7770 | 1.2029 | |
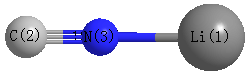 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.000 |
|
Li1 |
N3 |
C2 |
180.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.628 |
|
|
|
| 2 |
C |
-0.042 |
|
|
|
| 3 |
N |
-0.586 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
8.598 |
8.598 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.196 |
0.000 |
0.000 |
| y |
0.000 |
-14.196 |
0.000 |
| z |
0.000 |
0.000 |
-2.236 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.980 |
0.000 |
0.000 |
| y |
0.000 |
-5.980 |
0.000 |
| z |
0.000 |
0.000 |
11.960 |
|
| Polar |
|---|
| 3z2-r2 | 23.921 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.001 |
0.000 |
0.000 |
| y |
0.000 |
2.001 |
0.000 |
| z |
0.000 |
0.000 |
4.505 |
<r2> (average value of r
2) Å
2
| <r2> |
24.310 |
| (<r2>)1/2 |
4.930 |
