Jump to
S1C2
Energy calculated at B3LYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -418.291395 |
| Energy at 298.15K | -418.295341 |
| HF Energy | -418.291395 |
| Nuclear repulsion energy | 58.230581 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3621 |
3483 |
10.70 |
|
|
|
| 2 |
A' |
2152 |
2071 |
186.99 |
|
|
|
| 3 |
A' |
1133 |
1090 |
3.17 |
|
|
|
| 4 |
A' |
1101 |
1060 |
72.22 |
|
|
|
| 5 |
A' |
859 |
827 |
45.83 |
|
|
|
| 6 |
A' |
678 |
653 |
71.77 |
|
|
|
| 7 |
A" |
2177 |
2095 |
243.92 |
|
|
|
| 8 |
A" |
810 |
779 |
10.73 |
|
|
|
| 9 |
A" |
353 |
340 |
153.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6442.6 cm
-1
Scaled (by 0.962) Zero Point Vibrational Energy (zpe) 6197.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| P1 |
-0.109 |
-0.611 |
0.000 |
| O2 |
-0.109 |
1.167 |
0.000 |
| H3 |
0.776 |
1.588 |
0.000 |
| H4 |
0.867 |
-0.878 |
1.060 |
| H5 |
0.867 |
-0.878 |
-1.060 |
Atom - Atom Distances (Å)
| |
P1 |
O2 |
H3 |
H4 |
H5 |
| P1 | | 1.7786 | 2.3708 | 1.4656 | 1.4656 |
O2 | 1.7786 | | 0.9801 | 2.5023 | 2.5023 | H3 | 2.3708 | 0.9801 | | 2.6860 | 2.6860 | H4 | 1.4656 | 2.5023 | 2.6860 | | 2.1193 | H5 | 1.4656 | 2.5023 | 2.6860 | 2.1193 | |
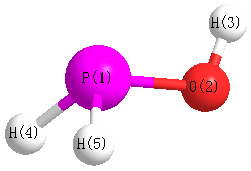 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| P1 |
O2 |
H3 |
115.424 |
|
O2 |
P1 |
H4 |
100.501 |
| O2 |
P1 |
H5 |
100.501 |
|
H4 |
P1 |
H5 |
92.608 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
P |
0.482 |
|
|
|
| 2 |
O |
-0.744 |
|
|
|
| 3 |
H |
0.372 |
|
|
|
| 4 |
H |
-0.055 |
|
|
|
| 5 |
H |
-0.055 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.790 |
-0.682 |
0.000 |
2.872 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.654 |
1.955 |
0.000 |
| y |
1.955 |
-20.245 |
0.000 |
| z |
0.000 |
0.000 |
-20.292 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.386 |
1.955 |
0.000 |
| y |
1.955 |
0.228 |
0.000 |
| z |
0.000 |
0.000 |
0.157 |
|
| Polar |
|---|
| 3z2-r2 | 0.315 |
|---|
| x2-y2 | -0.410 |
|---|
| xy | 1.955 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.265 |
-0.295 |
0.000 |
| y |
-0.295 |
4.049 |
0.000 |
| z |
0.000 |
0.000 |
3.495 |
<r2> (average value of r
2) Å
2
| <r2> |
37.937 |
| (<r2>)1/2 |
6.159 |
Jump to
S1C1
Energy calculated at B3LYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -418.295703 |
| Energy at 298.15K | -418.299528 |
| HF Energy | -418.295703 |
| Nuclear repulsion energy | 58.145542 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3661 |
3522 |
41.65 |
|
|
|
| 2 |
A' |
2241 |
2156 |
138.70 |
|
|
|
| 3 |
A' |
1142 |
1098 |
99.03 |
|
|
|
| 4 |
A' |
1127 |
1084 |
28.79 |
|
|
|
| 5 |
A' |
862 |
829 |
15.20 |
|
|
|
| 6 |
A' |
685 |
659 |
54.25 |
|
|
|
| 7 |
A" |
2257 |
2171 |
173.26 |
|
|
|
| 8 |
A" |
861 |
828 |
0.17 |
|
|
|
| 9 |
A" |
235 |
226 |
148.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6534.7 cm
-1
Scaled (by 0.962) Zero Point Vibrational Energy (zpe) 6286.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| P1 |
0.040 |
-0.620 |
0.000 |
| O2 |
0.040 |
1.174 |
0.000 |
| H3 |
0.962 |
1.501 |
0.000 |
| H4 |
-0.941 |
-0.795 |
1.055 |
| H5 |
-0.941 |
-0.795 |
-1.055 |
Atom - Atom Distances (Å)
| |
P1 |
O2 |
H3 |
H4 |
H5 |
| P1 | | 1.7943 | 2.3133 | 1.4512 | 1.4512 |
O2 | 1.7943 | | 0.9784 | 2.4394 | 2.4394 | H3 | 2.3133 | 0.9784 | | 3.1632 | 3.1632 | H4 | 1.4512 | 2.4394 | 3.1632 | | 2.1101 | H5 | 1.4512 | 2.4394 | 3.1632 | 2.1101 | |
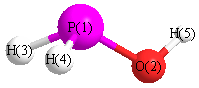 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| P1 |
O2 |
H3 |
109.544 |
|
O2 |
P1 |
H4 |
96.898 |
| O2 |
P1 |
H5 |
96.898 |
|
H4 |
P1 |
H5 |
93.278 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
P |
0.420 |
|
|
|
| 2 |
O |
-0.734 |
|
|
|
| 3 |
H |
0.375 |
|
|
|
| 4 |
H |
-0.030 |
|
|
|
| 5 |
H |
-0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.738 |
-0.823 |
0.000 |
1.106 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.474 |
4.616 |
0.000 |
| y |
4.616 |
-20.946 |
0.000 |
| z |
0.000 |
0.000 |
-20.078 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.038 |
4.616 |
0.000 |
| y |
4.616 |
-1.170 |
0.000 |
| z |
0.000 |
0.000 |
0.132 |
|
| Polar |
|---|
| 3z2-r2 | 0.264 |
|---|
| x2-y2 | 1.472 |
|---|
| xy | 4.616 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.442 |
0.792 |
0.000 |
| y |
0.792 |
4.106 |
0.000 |
| z |
0.000 |
0.000 |
3.457 |
<r2> (average value of r
2) Å
2
| <r2> |
37.868 |
| (<r2>)1/2 |
6.154 |
