Jump to
S1C2
S2C1
S2C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -131.207197 |
| Energy at 298.15K | -131.206544 |
| HF Energy | -131.125558 |
| Nuclear repulsion energy | 46.890632 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3401 |
3401 |
20.74 |
|
|
|
| 2 |
A' |
1797 |
1797 |
15.68 |
|
|
|
| 3 |
A' |
1186 |
1186 |
11.33 |
|
|
|
| 4 |
A' |
564 |
564 |
30.64 |
|
|
|
| 5 |
A' |
387 |
387 |
8.89 |
|
|
|
| 6 |
A" |
421 |
421 |
0.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3877.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 3877.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.060 |
-1.239 |
0.000 |
| C2 |
0.000 |
0.089 |
0.000 |
| N3 |
-0.148 |
1.288 |
0.000 |
| H4 |
0.682 |
-2.113 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3293 | 2.5354 | 1.0725 |
C2 | 1.3293 | | 1.2081 | 2.3046 | N3 | 2.5354 | 1.2081 | | 3.5002 | H4 | 1.0725 | 2.3046 | 3.5002 | |
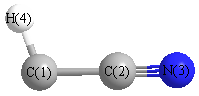 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
175.508 |
|
C2 |
C1 |
H4 |
147.088 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -131.205914 |
| Energy at 298.15K | |
| HF Energy | -131.125831 |
| Nuclear repulsion energy | 46.980678 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3476 |
3476 |
58.71 |
|
|
|
| 2 |
Σ |
1617 |
1617 |
27.71 |
|
|
|
| 3 |
Σ |
1261 |
1261 |
6.68 |
|
|
|
| 4 |
Π |
406 |
406 |
0.12 |
|
|
|
| 4 |
Π |
406 |
406 |
0.12 |
|
|
|
| 5 |
Π |
373i |
373i |
60.31 |
|
|
|
| 5 |
Π |
373i |
373i |
60.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3210.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 3210.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.219 |
| C2 |
0.000 |
0.000 |
0.079 |
| N3 |
0.000 |
0.000 |
1.304 |
| H4 |
0.000 |
0.000 |
-2.286 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.2986 | 2.5231 | 1.0664 |
C2 | 1.2986 | | 1.2246 | 2.3649 | N3 | 2.5231 | 1.2246 | | 3.5895 | H4 | 1.0664 | 2.3649 | 3.5895 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.019 |
|
|
|
| 2 |
C |
-0.056 |
|
|
|
| 3 |
N |
-0.215 |
|
|
|
| 4 |
H |
0.289 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.438 |
3.438 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.114 |
0.000 |
0.000 |
| y |
0.000 |
-17.114 |
0.000 |
| z |
0.000 |
0.000 |
-16.088 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.513 |
0.000 |
0.000 |
| y |
0.000 |
-0.513 |
0.000 |
| z |
0.000 |
0.000 |
1.026 |
|
| Polar |
|---|
| 3z2-r2 | 2.052 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.463 |
0.000 |
0.000 |
| y |
0.000 |
1.463 |
0.000 |
| z |
0.000 |
0.000 |
6.013 |
<r2> (average value of r
2) Å
2
| <r2> |
36.558 |
| (<r2>)1/2 |
6.046 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -131.177742 |
| Energy at 298.15K | -131.177144 |
| HF Energy | -131.078237 |
| Nuclear repulsion energy | 46.353514 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3082 |
3082 |
18.46 |
|
|
|
| 2 |
A' |
2030 |
2030 |
17.21 |
|
|
|
| 3 |
A' |
1086 |
1086 |
45.73 |
|
|
|
| 4 |
A' |
879 |
879 |
62.25 |
|
|
|
| 5 |
A' |
404 |
404 |
18.34 |
|
|
|
| 6 |
A" |
334 |
334 |
2.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3907.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 3907.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.402 |
-1.227 |
0.000 |
| C2 |
0.000 |
0.098 |
0.000 |
| N3 |
-0.558 |
1.165 |
0.000 |
| H4 |
1.494 |
-1.379 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3847 | 2.5780 | 1.1026 |
C2 | 1.3847 | | 1.2046 | 2.1009 | N3 | 2.5780 | 1.2046 | | 3.2689 | H4 | 1.1026 | 2.1009 | 3.2689 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.017 |
|
|
|
| 2 |
C |
0.025 |
|
|
|
| 3 |
N |
-0.209 |
|
|
|
| 4 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.271 |
-1.043 |
0.000 |
2.499 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.139 |
-1.227 |
0.000 |
| y |
-1.227 |
-22.293 |
0.000 |
| z |
0.000 |
0.000 |
-15.674 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.844 |
-1.227 |
0.000 |
| y |
-1.227 |
-6.387 |
0.000 |
| z |
0.000 |
0.000 |
3.542 |
|
| Polar |
|---|
| 3z2-r2 | 7.084 |
|---|
| x2-y2 | 6.154 |
|---|
| xy | -1.227 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.925 |
-1.710 |
0.000 |
| y |
-1.710 |
5.512 |
0.000 |
| z |
0.000 |
0.000 |
2.171 |
<r2> (average value of r
2) Å
2
| <r2> |
37.149 |
| (<r2>)1/2 |
6.095 |
Jump to
S1C1
S1C2
S2C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -131.177742 |
| Energy at 298.15K | -131.177144 |
| HF Energy | -131.078237 |
| Nuclear repulsion energy | 46.353514 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
