Jump to
S1C2
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -343.656443 |
| Energy at 298.15K | -343.673337 |
| Nuclear repulsion energy | 417.799058 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3072 |
2919 |
0.00 |
|
|
|
| 2 |
A1' |
1560 |
1482 |
0.00 |
|
|
|
| 3 |
A1' |
1373 |
1304 |
0.00 |
|
|
|
| 4 |
A1' |
953 |
906 |
0.00 |
|
|
|
| 5 |
A1' |
780 |
741 |
0.00 |
|
|
|
| 6 |
A1' |
607 |
577 |
0.00 |
|
|
|
| 7 |
A1" |
3109 |
2954 |
0.00 |
|
|
|
| 8 |
A1" |
1294 |
1230 |
0.00 |
|
|
|
| 9 |
A1" |
1018 |
967 |
0.00 |
|
|
|
| 10 |
A1" |
30 |
28 |
0.00 |
|
|
|
| 11 |
A2' |
3131 |
2975 |
0.00 |
|
|
|
| 12 |
A2' |
1204 |
1143 |
0.00 |
|
|
|
| 13 |
A2' |
826 |
785 |
0.00 |
|
|
|
| 14 |
A2" |
3059 |
2906 |
101.68 |
|
|
|
| 15 |
A2" |
1552 |
1474 |
4.54 |
|
|
|
| 16 |
A2" |
1404 |
1334 |
1.27 |
|
|
|
| 17 |
A2" |
985 |
936 |
20.99 |
|
|
|
| 18 |
A2" |
693 |
659 |
65.64 |
|
|
|
| 19 |
E' |
3140 |
2983 |
76.45 |
|
|
|
| 19 |
E' |
3140 |
2983 |
76.45 |
|
|
|
| 20 |
E' |
3067 |
2914 |
88.17 |
|
|
|
| 20 |
E' |
3067 |
2914 |
88.17 |
|
|
|
| 21 |
E' |
1549 |
1472 |
11.23 |
|
|
|
| 21 |
E' |
1549 |
1472 |
11.23 |
|
|
|
| 22 |
E' |
1369 |
1301 |
6.12 |
|
|
|
| 22 |
E' |
1369 |
1301 |
6.12 |
|
|
|
| 23 |
E' |
1345 |
1278 |
0.00 |
|
|
|
| 23 |
E' |
1345 |
1278 |
0.00 |
|
|
|
| 24 |
E' |
1053 |
1001 |
27.33 |
|
|
|
| 24 |
E' |
1053 |
1001 |
27.33 |
|
|
|
| 25 |
E' |
899 |
854 |
4.19 |
|
|
|
| 25 |
E' |
899 |
854 |
4.19 |
|
|
|
| 26 |
E' |
845 |
803 |
6.92 |
|
|
|
| 26 |
E' |
845 |
803 |
6.92 |
|
|
|
| 27 |
E' |
424 |
403 |
0.06 |
|
|
|
| 27 |
E' |
424 |
403 |
0.06 |
|
|
|
| 28 |
E" |
3113 |
2958 |
0.00 |
|
|
|
| 28 |
E" |
3113 |
2958 |
0.00 |
|
|
|
| 29 |
E" |
3058 |
2905 |
0.00 |
|
|
|
| 29 |
E" |
3058 |
2905 |
0.00 |
|
|
|
| 30 |
E" |
1539 |
1462 |
0.00 |
|
|
|
| 30 |
E" |
1539 |
1462 |
0.00 |
|
|
|
| 31 |
E" |
1370 |
1302 |
0.00 |
|
|
|
| 31 |
E" |
1370 |
1302 |
0.00 |
|
|
|
| 32 |
E" |
1346 |
1278 |
0.00 |
|
|
|
| 32 |
E" |
1346 |
1278 |
0.00 |
|
|
|
| 33 |
E" |
1218 |
1157 |
0.00 |
|
|
|
| 33 |
E" |
1218 |
1157 |
0.00 |
|
|
|
| 34 |
E" |
1009 |
959 |
0.00 |
|
|
|
| 34 |
E" |
1009 |
959 |
0.00 |
|
|
|
| 35 |
E" |
572 |
543 |
0.00 |
|
|
|
| 35 |
E" |
572 |
543 |
0.00 |
|
|
|
| 36 |
E" |
308 |
293 |
0.00 |
|
|
|
| 36 |
E" |
308 |
293 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40548.5 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 38525.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.299 |
| N2 |
0.000 |
0.000 |
-1.299 |
| C3 |
0.000 |
1.415 |
0.788 |
| C4 |
1.225 |
-0.707 |
0.788 |
| C5 |
-1.225 |
-0.707 |
0.788 |
| C6 |
0.000 |
1.415 |
-0.788 |
| C7 |
-1.225 |
-0.707 |
-0.788 |
| C8 |
1.225 |
-0.707 |
-0.788 |
| H9 |
0.889 |
1.915 |
1.197 |
| H10 |
-0.889 |
1.915 |
1.197 |
| H11 |
1.213 |
-1.728 |
1.197 |
| H12 |
2.103 |
-0.187 |
1.197 |
| H13 |
-2.103 |
-0.187 |
1.197 |
| H14 |
-1.213 |
-1.728 |
1.197 |
| H15 |
-0.889 |
1.915 |
-1.197 |
| H16 |
0.889 |
1.915 |
-1.197 |
| H17 |
-1.213 |
-1.728 |
-1.197 |
| H18 |
-2.103 |
-0.187 |
-1.197 |
| H19 |
2.103 |
-0.187 |
-1.197 |
| H20 |
1.213 |
-1.728 |
-1.197 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5974 | 1.5042 | 1.5042 | 1.5042 | 2.5211 | 2.5211 | 2.5211 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | 3.2688 | 3.2688 | 3.2688 | 3.2688 | 3.2688 | 3.2688 |
N2 | 2.5974 | | 2.5211 | 2.5211 | 2.5211 | 1.5042 | 1.5042 | 1.5042 | 3.2688 | 3.2688 | 3.2688 | 3.2688 | 3.2688 | 3.2688 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | C3 | 1.5042 | 2.5211 | | 2.4505 | 2.4505 | 1.5759 | 2.9135 | 2.9135 | 1.0991 | 1.0991 | 3.3933 | 2.6750 | 2.6750 | 3.3933 | 2.2318 | 2.2318 | 3.9098 | 3.3058 | 3.3058 | 3.9098 | C4 | 1.5042 | 2.5211 | 2.4505 | | 2.4505 | 2.9135 | 2.9135 | 1.5759 | 2.6750 | 3.3933 | 1.0991 | 1.0991 | 3.3933 | 2.6750 | 3.9098 | 3.3058 | 3.3058 | 3.9098 | 2.2318 | 2.2318 | C5 | 1.5042 | 2.5211 | 2.4505 | 2.4505 | | 2.9135 | 1.5759 | 2.9135 | 3.3933 | 2.6750 | 2.6750 | 3.3933 | 1.0991 | 1.0991 | 3.3058 | 3.9098 | 2.2318 | 2.2318 | 3.9098 | 3.3058 | C6 | 2.5211 | 1.5042 | 1.5759 | 2.9135 | 2.9135 | | 2.4505 | 2.4505 | 2.2318 | 2.2318 | 3.9098 | 3.3058 | 3.3058 | 3.9098 | 1.0991 | 1.0991 | 3.3933 | 2.6750 | 2.6750 | 3.3933 | C7 | 2.5211 | 1.5042 | 2.9135 | 2.9135 | 1.5759 | 2.4505 | | 2.4505 | 3.9098 | 3.3058 | 3.3058 | 3.9098 | 2.2318 | 2.2318 | 2.6750 | 3.3933 | 1.0991 | 1.0991 | 3.3933 | 2.6750 | C8 | 2.5211 | 1.5042 | 2.9135 | 1.5759 | 2.9135 | 2.4505 | 2.4505 | | 3.3058 | 3.9098 | 2.2318 | 2.2318 | 3.9098 | 3.3058 | 3.3933 | 2.6750 | 2.6750 | 3.3933 | 1.0991 | 1.0991 | H9 | 2.1136 | 3.2688 | 1.0991 | 2.6750 | 3.3933 | 2.2318 | 3.9098 | 3.3058 | | 1.7787 | 3.6566 | 2.4270 | 3.6566 | 4.2057 | 2.9824 | 2.3940 | 4.8393 | 4.3706 | 3.4090 | 4.3706 | H10 | 2.1136 | 3.2688 | 1.0991 | 3.3933 | 2.6750 | 2.2318 | 3.3058 | 3.9098 | 1.7787 | | 4.2057 | 3.6566 | 2.4270 | 3.6566 | 2.3940 | 2.9824 | 4.3706 | 3.4090 | 4.3706 | 4.8393 | H11 | 2.1136 | 3.2688 | 3.3933 | 1.0991 | 2.6750 | 3.9098 | 3.3058 | 2.2318 | 3.6566 | 4.2057 | | 1.7787 | 3.6566 | 2.4270 | 4.8393 | 4.3706 | 3.4090 | 4.3706 | 2.9824 | 2.3940 | H12 | 2.1136 | 3.2688 | 2.6750 | 1.0991 | 3.3933 | 3.3058 | 3.9098 | 2.2318 | 2.4270 | 3.6566 | 1.7787 | | 4.2057 | 3.6566 | 4.3706 | 3.4090 | 4.3706 | 4.8393 | 2.3940 | 2.9824 | H13 | 2.1136 | 3.2688 | 2.6750 | 3.3933 | 1.0991 | 3.3058 | 2.2318 | 3.9098 | 3.6566 | 2.4270 | 3.6566 | 4.2057 | | 1.7787 | 3.4090 | 4.3706 | 2.9824 | 2.3940 | 4.8393 | 4.3706 | H14 | 2.1136 | 3.2688 | 3.3933 | 2.6750 | 1.0991 | 3.9098 | 2.2318 | 3.3058 | 4.2057 | 3.6566 | 2.4270 | 3.6566 | 1.7787 | | 4.3706 | 4.8393 | 2.3940 | 2.9824 | 4.3706 | 3.4090 | H15 | 3.2688 | 2.1136 | 2.2318 | 3.9098 | 3.3058 | 1.0991 | 2.6750 | 3.3933 | 2.9824 | 2.3940 | 4.8393 | 4.3706 | 3.4090 | 4.3706 | | 1.7787 | 3.6566 | 2.4270 | 3.6566 | 4.2057 | H16 | 3.2688 | 2.1136 | 2.2318 | 3.3058 | 3.9098 | 1.0991 | 3.3933 | 2.6750 | 2.3940 | 2.9824 | 4.3706 | 3.4090 | 4.3706 | 4.8393 | 1.7787 | | 4.2057 | 3.6566 | 2.4270 | 3.6566 | H17 | 3.2688 | 2.1136 | 3.9098 | 3.3058 | 2.2318 | 3.3933 | 1.0991 | 2.6750 | 4.8393 | 4.3706 | 3.4090 | 4.3706 | 2.9824 | 2.3940 | 3.6566 | 4.2057 | | 1.7787 | 3.6566 | 2.4270 | H18 | 3.2688 | 2.1136 | 3.3058 | 3.9098 | 2.2318 | 2.6750 | 1.0991 | 3.3933 | 4.3706 | 3.4090 | 4.3706 | 4.8393 | 2.3940 | 2.9824 | 2.4270 | 3.6566 | 1.7787 | | 4.2057 | 3.6566 | H19 | 3.2688 | 2.1136 | 3.3058 | 2.2318 | 3.9098 | 2.6750 | 3.3933 | 1.0991 | 3.4090 | 4.3706 | 2.9824 | 2.3940 | 4.8393 | 4.3706 | 3.6566 | 2.4270 | 3.6566 | 4.2057 | | 1.7787 | H20 | 3.2688 | 2.1136 | 3.9098 | 2.2318 | 3.3058 | 3.3933 | 2.6750 | 1.0991 | 4.3706 | 4.8393 | 2.3940 | 2.9824 | 4.3706 | 3.4090 | 4.2057 | 3.6566 | 2.4270 | 3.6566 | 1.7787 | |
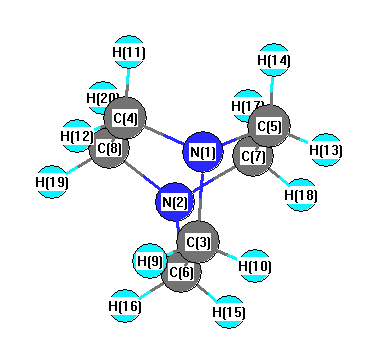 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
109.849 |
|
N1 |
C3 |
H9 |
107.543 |
| N1 |
C3 |
H10 |
107.543 |
|
N1 |
C4 |
C8 |
109.849 |
| N1 |
C4 |
H11 |
107.543 |
|
N1 |
C4 |
H12 |
107.543 |
| N1 |
C5 |
C7 |
109.849 |
|
N1 |
C5 |
H13 |
107.543 |
| N1 |
C5 |
H14 |
107.543 |
|
N2 |
C6 |
C3 |
109.849 |
| N2 |
C6 |
H15 |
107.543 |
|
N2 |
C6 |
H16 |
107.543 |
| N2 |
C7 |
C5 |
109.849 |
|
N2 |
C7 |
H17 |
107.543 |
| N2 |
C7 |
H18 |
107.543 |
|
N2 |
C8 |
C4 |
109.849 |
| N2 |
C8 |
H19 |
107.543 |
|
N2 |
C8 |
H20 |
107.543 |
| C3 |
N1 |
C4 |
109.091 |
|
C3 |
N1 |
C5 |
109.091 |
| C3 |
C6 |
H15 |
111.847 |
|
C3 |
C6 |
H16 |
111.847 |
| C4 |
N1 |
C5 |
109.091 |
|
C4 |
C8 |
H19 |
111.847 |
| C4 |
C8 |
H20 |
111.847 |
|
C5 |
C6 |
H15 |
101.086 |
| C5 |
C6 |
H16 |
150.789 |
|
C6 |
N2 |
C7 |
109.091 |
| C6 |
N2 |
C8 |
109.091 |
|
C6 |
C3 |
H9 |
111.847 |
| C6 |
C3 |
H10 |
111.847 |
|
C7 |
N2 |
C8 |
109.091 |
| C7 |
C5 |
H13 |
111.847 |
|
C7 |
C5 |
H14 |
111.847 |
| C8 |
C4 |
H11 |
111.847 |
|
C8 |
C4 |
H12 |
111.847 |
| H9 |
C3 |
H10 |
108.021 |
|
H11 |
C4 |
H12 |
108.021 |
| H13 |
C5 |
H14 |
108.021 |
|
H15 |
C6 |
H16 |
108.021 |
| H17 |
C7 |
H18 |
108.021 |
|
H19 |
C8 |
H20 |
108.021 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -343.656443 |
| Energy at 298.15K | -343.673337 |
| HF Energy | -342.881524 |
| Nuclear repulsion energy | 417.805087 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3109 |
2954 |
0.00 |
|
|
|
| 2 |
A1 |
3072 |
2919 |
0.00 |
|
|
|
| 3 |
A1 |
1560 |
1482 |
0.00 |
|
|
|
| 4 |
A1 |
1373 |
1304 |
0.00 |
|
|
|
| 5 |
A1 |
1294 |
1230 |
0.00 |
|
|
|
| 6 |
A1 |
1018 |
967 |
0.00 |
|
|
|
| 7 |
A1 |
954 |
906 |
0.00 |
|
|
|
| 8 |
A1 |
780 |
741 |
0.00 |
|
|
|
| 9 |
A1 |
607 |
577 |
0.00 |
|
|
|
| 10 |
A1 |
29 |
28 |
0.00 |
|
|
|
| 11 |
A2 |
3131 |
2975 |
0.00 |
|
|
|
| 12 |
A2 |
3059 |
2906 |
101.69 |
|
|
|
| 13 |
A2 |
1552 |
1474 |
4.55 |
|
|
|
| 14 |
A2 |
1404 |
1334 |
1.27 |
|
|
|
| 15 |
A2 |
1204 |
1143 |
0.00 |
|
|
|
| 16 |
A2 |
985 |
936 |
20.99 |
|
|
|
| 17 |
A2 |
826 |
785 |
0.00 |
|
|
|
| 18 |
A2 |
693 |
659 |
65.64 |
|
|
|
| 19 |
E |
3140 |
2983 |
76.46 |
|
|
|
| 19 |
E |
3140 |
2983 |
76.46 |
|
|
|
| 20 |
E |
3113 |
2958 |
0.00 |
|
|
|
| 20 |
E |
3113 |
2958 |
0.00 |
|
|
|
| 21 |
E |
3067 |
2914 |
88.17 |
|
|
|
| 21 |
E |
3067 |
2914 |
88.17 |
|
|
|
| 22 |
E |
3058 |
2905 |
0.00 |
|
|
|
| 22 |
E |
3058 |
2905 |
0.00 |
|
|
|
| 23 |
E |
1549 |
1472 |
11.23 |
|
|
|
| 23 |
E |
1549 |
1472 |
11.23 |
|
|
|
| 24 |
E |
1539 |
1462 |
0.00 |
|
|
|
| 24 |
E |
1539 |
1462 |
0.00 |
|
|
|
| 25 |
E |
1370 |
1302 |
0.00 |
|
|
|
| 25 |
E |
1370 |
1302 |
0.00 |
|
|
|
| 26 |
E |
1369 |
1301 |
6.12 |
|
|
|
| 26 |
E |
1369 |
1301 |
6.12 |
|
|
|
| 27 |
E |
1346 |
1278 |
0.00 |
|
|
|
| 27 |
E |
1346 |
1278 |
0.00 |
|
|
|
| 28 |
E |
1346 |
1278 |
0.00 |
|
|
|
| 28 |
E |
1346 |
1278 |
0.00 |
|
|
|
| 29 |
E |
1218 |
1157 |
0.00 |
|
|
|
| 29 |
E |
1218 |
1157 |
0.00 |
|
|
|
| 30 |
E |
1053 |
1001 |
27.33 |
|
|
|
| 30 |
E |
1053 |
1001 |
27.33 |
|
|
|
| 31 |
E |
1009 |
959 |
0.00 |
|
|
|
| 31 |
E |
1009 |
959 |
0.00 |
|
|
|
| 32 |
E |
899 |
854 |
4.20 |
|
|
|
| 32 |
E |
899 |
854 |
4.20 |
|
|
|
| 33 |
E |
845 |
803 |
6.92 |
|
|
|
| 33 |
E |
845 |
803 |
6.92 |
|
|
|
| 34 |
E |
572 |
543 |
0.00 |
|
|
|
| 34 |
E |
572 |
543 |
0.00 |
|
|
|
| 35 |
E |
424 |
403 |
0.06 |
|
|
|
| 35 |
E |
424 |
403 |
0.06 |
|
|
|
| 36 |
E |
308 |
293 |
0.00 |
|
|
|
| 36 |
E |
308 |
293 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40548.9 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 38525.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.299 |
| N2 |
0.000 |
0.000 |
-1.299 |
| C3 |
0.000 |
1.415 |
0.788 |
| C4 |
1.225 |
-0.708 |
0.788 |
| C5 |
-1.225 |
-0.707 |
0.788 |
| C6 |
-0.000 |
1.415 |
-0.788 |
| C7 |
-1.225 |
-0.708 |
-0.788 |
| C8 |
1.225 |
-0.707 |
-0.788 |
| H9 |
0.890 |
1.914 |
1.197 |
| H10 |
-0.889 |
1.915 |
1.197 |
| H11 |
1.213 |
-1.728 |
1.197 |
| H12 |
2.103 |
-0.188 |
1.197 |
| H13 |
-2.103 |
-0.187 |
1.197 |
| H14 |
-1.214 |
-1.727 |
1.197 |
| H15 |
-0.890 |
1.914 |
-1.197 |
| H16 |
0.889 |
1.915 |
-1.197 |
| H17 |
-1.213 |
-1.728 |
-1.197 |
| H18 |
-2.103 |
-0.188 |
-1.197 |
| H19 |
2.103 |
-0.187 |
-1.197 |
| H20 |
1.214 |
-1.727 |
-1.197 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5972 | 1.5041 | 1.5041 | 1.5041 | 2.5210 | 2.5210 | 2.5210 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | 3.2686 | 3.2690 | 3.2686 | 3.2690 | 3.2686 | 3.2690 |
N2 | 2.5972 | | 2.5210 | 2.5210 | 2.5210 | 1.5041 | 1.5041 | 1.5041 | 3.2686 | 3.2690 | 3.2686 | 3.2690 | 3.2686 | 3.2690 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | 2.1136 | C3 | 1.5041 | 2.5210 | | 2.4505 | 2.4505 | 1.5759 | 2.9136 | 2.9133 | 1.0992 | 1.0992 | 3.3932 | 2.6751 | 2.6748 | 3.3933 | 2.2317 | 2.2318 | 3.9098 | 3.3063 | 3.3053 | 3.9098 | C4 | 1.5041 | 2.5210 | 2.4505 | | 2.4505 | 2.9136 | 2.9133 | 1.5759 | 2.6748 | 3.3933 | 1.0992 | 1.0992 | 3.3932 | 2.6751 | 3.9098 | 3.3063 | 3.3053 | 3.9098 | 2.2317 | 2.2318 | C5 | 1.5041 | 2.5210 | 2.4505 | 2.4505 | | 2.9133 | 1.5759 | 2.9136 | 3.3932 | 2.6751 | 2.6748 | 3.3933 | 1.0992 | 1.0992 | 3.3053 | 3.9098 | 2.2317 | 2.2318 | 3.9098 | 3.3063 | C6 | 2.5210 | 1.5041 | 1.5759 | 2.9136 | 2.9133 | | 2.4505 | 2.4505 | 2.2317 | 2.2318 | 3.9098 | 3.3063 | 3.3053 | 3.9098 | 1.0992 | 1.0992 | 3.3932 | 2.6751 | 2.6748 | 3.3933 | C7 | 2.5210 | 1.5041 | 2.9136 | 2.9133 | 1.5759 | 2.4505 | | 2.4505 | 3.9098 | 3.3063 | 3.3053 | 3.9098 | 2.2317 | 2.2318 | 2.6748 | 3.3933 | 1.0992 | 1.0992 | 3.3932 | 2.6751 | C8 | 2.5210 | 1.5041 | 2.9133 | 1.5759 | 2.9136 | 2.4505 | 2.4505 | | 3.3053 | 3.9098 | 2.2317 | 2.2318 | 3.9098 | 3.3063 | 3.3932 | 2.6751 | 2.6748 | 3.3933 | 1.0992 | 1.0992 | H9 | 2.1136 | 3.2686 | 1.0992 | 2.6748 | 3.3932 | 2.2317 | 3.9098 | 3.3053 | | 1.7787 | 3.6565 | 2.4270 | 3.6565 | 4.2057 | 2.9826 | 2.3940 | 4.8391 | 4.3710 | 3.4081 | 4.3702 | H10 | 2.1136 | 3.2690 | 1.0992 | 3.3933 | 2.6751 | 2.2318 | 3.3063 | 3.9098 | 1.7787 | | 4.2057 | 3.6567 | 2.4270 | 3.6567 | 2.3940 | 2.9823 | 4.3710 | 3.4100 | 4.3702 | 4.8395 | H11 | 2.1136 | 3.2686 | 3.3932 | 1.0992 | 2.6748 | 3.9098 | 3.3053 | 2.2317 | 3.6565 | 4.2057 | | 1.7787 | 3.6565 | 2.4270 | 4.8391 | 4.3710 | 3.4081 | 4.3702 | 2.9826 | 2.3940 | H12 | 2.1136 | 3.2690 | 2.6751 | 1.0992 | 3.3933 | 3.3063 | 3.9098 | 2.2318 | 2.4270 | 3.6567 | 1.7787 | | 4.2057 | 3.6567 | 4.3710 | 3.4100 | 4.3702 | 4.8395 | 2.3940 | 2.9823 | H13 | 2.1136 | 3.2686 | 2.6748 | 3.3932 | 1.0992 | 3.3053 | 2.2317 | 3.9098 | 3.6565 | 2.4270 | 3.6565 | 4.2057 | | 1.7787 | 3.4081 | 4.3702 | 2.9826 | 2.3940 | 4.8391 | 4.3710 | H14 | 2.1136 | 3.2690 | 3.3933 | 2.6751 | 1.0992 | 3.9098 | 2.2318 | 3.3063 | 4.2057 | 3.6567 | 2.4270 | 3.6567 | 1.7787 | | 4.3702 | 4.8395 | 2.3940 | 2.9823 | 4.3710 | 3.4100 | H15 | 3.2686 | 2.1136 | 2.2317 | 3.9098 | 3.3053 | 1.0992 | 2.6748 | 3.3932 | 2.9826 | 2.3940 | 4.8391 | 4.3710 | 3.4081 | 4.3702 | | 1.7787 | 3.6565 | 2.4270 | 3.6565 | 4.2057 | H16 | 3.2690 | 2.1136 | 2.2318 | 3.3063 | 3.9098 | 1.0992 | 3.3933 | 2.6751 | 2.3940 | 2.9823 | 4.3710 | 3.4100 | 4.3702 | 4.8395 | 1.7787 | | 4.2057 | 3.6567 | 2.4270 | 3.6567 | H17 | 3.2686 | 2.1136 | 3.9098 | 3.3053 | 2.2317 | 3.3932 | 1.0992 | 2.6748 | 4.8391 | 4.3710 | 3.4081 | 4.3702 | 2.9826 | 2.3940 | 3.6565 | 4.2057 | | 1.7787 | 3.6565 | 2.4270 | H18 | 3.2690 | 2.1136 | 3.3063 | 3.9098 | 2.2318 | 2.6751 | 1.0992 | 3.3933 | 4.3710 | 3.4100 | 4.3702 | 4.8395 | 2.3940 | 2.9823 | 2.4270 | 3.6567 | 1.7787 | | 4.2057 | 3.6567 | H19 | 3.2686 | 2.1136 | 3.3053 | 2.2317 | 3.9098 | 2.6748 | 3.3932 | 1.0992 | 3.4081 | 4.3702 | 2.9826 | 2.3940 | 4.8391 | 4.3710 | 3.6565 | 2.4270 | 3.6565 | 4.2057 | | 1.7787 | H20 | 3.2690 | 2.1136 | 3.9098 | 2.2318 | 3.3063 | 3.3933 | 2.6751 | 1.0992 | 4.3702 | 4.8395 | 2.3940 | 2.9823 | 4.3710 | 3.4100 | 4.2057 | 3.6567 | 2.4270 | 3.6567 | 1.7787 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
109.847 |
|
N1 |
C3 |
H9 |
107.541 |
| N1 |
C3 |
H10 |
107.544 |
|
N1 |
C4 |
C8 |
109.847 |
| N1 |
C4 |
H11 |
107.541 |
|
N1 |
C4 |
H12 |
107.544 |
| N1 |
C5 |
C7 |
109.847 |
|
N1 |
C5 |
H13 |
107.541 |
| N1 |
C5 |
H14 |
107.544 |
|
N2 |
C6 |
C3 |
109.847 |
| N2 |
C6 |
H15 |
107.541 |
|
N2 |
C6 |
H16 |
107.544 |
| N2 |
C7 |
C5 |
109.847 |
|
N2 |
C7 |
H17 |
107.541 |
| N2 |
C7 |
H18 |
107.544 |
|
N2 |
C8 |
C4 |
109.847 |
| N2 |
C8 |
H19 |
107.541 |
|
N2 |
C8 |
H20 |
107.544 |
| C3 |
N1 |
C4 |
109.093 |
|
C3 |
N1 |
C5 |
109.093 |
| C3 |
C6 |
H15 |
111.848 |
|
C3 |
C6 |
H16 |
111.851 |
| C4 |
N1 |
C5 |
109.093 |
|
C4 |
C8 |
H19 |
111.848 |
| C4 |
C8 |
H20 |
111.851 |
|
C5 |
C6 |
H15 |
101.065 |
| C5 |
C6 |
H16 |
150.811 |
|
C6 |
N2 |
C7 |
109.093 |
| C6 |
N2 |
C8 |
109.093 |
|
C6 |
C3 |
H9 |
111.848 |
| C6 |
C3 |
H10 |
111.851 |
|
C7 |
N2 |
C8 |
109.093 |
| C7 |
C5 |
H13 |
111.848 |
|
C7 |
C5 |
H14 |
111.851 |
| C8 |
C4 |
H11 |
111.848 |
|
C8 |
C4 |
H12 |
111.851 |
| H9 |
C3 |
H10 |
108.019 |
|
H11 |
C4 |
H12 |
108.019 |
| H13 |
C5 |
H14 |
108.019 |
|
H15 |
C6 |
H16 |
108.019 |
| H17 |
C7 |
H18 |
108.019 |
|
H19 |
C8 |
H20 |
108.019 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
