return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (benzocyclobutadiene)

using model chemistry: MP2=FULL/6-31G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31G
 hartrees
Energy at 0K-306.920763
Energy at 298.15K 
HF Energy-306.209836
Nuclear repulsion energy308.036115
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3266 3103 18.17      
2 A1 3210 3050 21.49      
3 A1 3190 3031 11.54      
4 A1 1674 1590 0.94      
5 A1 1501 1426 2.62      
6 A1 1484 1410 2.55      
7 A1 1442 1370 9.90      
8 A1 1231 1169 0.00      
9 A1 1096 1041 0.01      
10 A1 1086 1032 0.31      
11 A1 982 933 1.94      
12 A1 801 761 0.14      
13 A1 545 518 0.01      
14 A2 848 805 0.00      
15 A2 828 787 0.00      
16 A2 705 670 0.00      
17 A2 484 460 0.00      
18 A2 278 264 0.00      
19 A2 400i 380i 0.00      
20 B1 803 763 4.24      
21 B1 722 686 82.47      
22 B1 608 578 60.79      
23 B1 321 305 0.22      
24 B1 140 133 7.25      
25 B2 3233 3072 4.12      
26 B2 3200 3040 39.27      
27 B2 3176 3018 0.00      
28 B2 1637 1556 1.08      
29 B2 1479 1405 4.25      
30 B2 1327 1261 0.53      
31 B2 1268 1205 15.20      
32 B2 1115 1060 1.01      
33 B2 1064 1011 1.84      
34 B2 855 813 0.06      
35 B2 665 632 1.33      
36 B2 398 378 3.30      

Unscaled Zero Point Vibrational Energy (zpe) 23131.0 cm-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 21976.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G
ABC
0.15674 0.07110 0.04891

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.464 -0.631
C2 0.000 0.697 -1.872
C3 0.000 -0.697 -1.872
C4 0.000 -1.464 -0.631
C5 0.000 -0.690 2.077
C6 0.000 0.690 2.077
C7 0.000 0.730 0.533
C8 0.000 -0.730 0.533
H9 0.000 2.555 -0.664
H10 0.000 1.238 -2.820
H11 0.000 -1.238 -2.820
H12 0.000 -2.555 -0.664
H13 0.000 -1.459 2.845
H14 0.000 1.459 2.845

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.45812.49162.92703.45942.81611.37542.48331.09172.20023.47674.01844.54133.4756
C21.45811.39452.49164.18473.94822.40442.79602.21581.09132.15483.46915.18594.7773
C32.49161.39451.45813.94824.18472.79602.40443.46912.15481.09132.21584.77735.1859
C42.92702.49161.45812.81613.45942.48331.37544.01843.47672.20021.09173.47564.5413
C53.45944.18473.94822.81611.37922.09761.54464.24665.26204.92683.31471.08732.2821
C62.81613.94824.18473.45941.37921.54462.09763.31474.92685.26204.24662.28211.0873
C71.37542.40442.79602.48332.09761.54461.46052.18163.39043.88723.49603.18432.4242
C82.48332.79602.40441.37541.54462.09761.46053.49603.88723.39042.18162.42423.1843
H91.09172.21583.46914.01844.24663.31472.18163.49602.52654.36265.10945.33103.6751
H102.20021.09132.15483.47675.26204.92683.39043.88722.52652.47574.36266.27365.6685
H113.47672.15481.09132.20024.92685.26203.88723.39044.36262.47572.52655.66856.2736
H124.01843.46912.21581.09173.31474.24663.49602.18165.10944.36262.52653.67515.3310
H134.54135.18594.77733.47561.08732.28213.18432.42425.33106.27365.66853.67512.9188
H143.47564.77735.18594.54132.28211.08732.42423.18433.67515.66856.27365.33102.9188

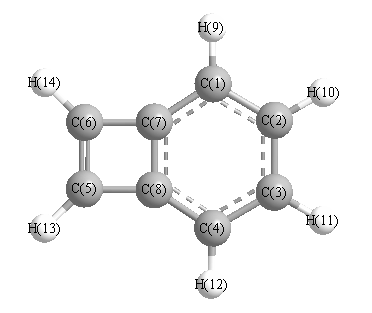
picture of benzocyclobutadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.702 C1 C2 H10 118.606
C1 C7 C6 149.292 C1 C7 N8 122.217
C2 C1 C7 116.081 C2 C1 H9 119.998
C2 C3 C4 121.702 C2 C3 H11 119.692
C3 C2 H10 119.692 C3 C4 N8 116.081
C3 C4 H12 119.998 C4 C3 H11 118.606
C4 N8 C5 149.292 C4 N8 C7 122.217
C5 C6 C7 91.509 C5 C6 H14 135.068
C5 N8 C7 88.491 C6 C5 N8 91.509
C6 C5 H13 135.068 C6 C7 N8 88.491
C7 C1 H9 123.921 C7 C6 H14 133.423
N8 C4 H12 123.921 N8 C5 H13 133.423
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability