Jump to
S1C2
Energy calculated at B2PLYP=FULLultrafine/6-31G
| | hartrees |
|---|
| Energy at 0K | -377.041474 |
| Energy at 298.15K | |
| HF Energy | -376.857275 |
| Nuclear repulsion energy | 186.357112 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3210 |
3210 |
24.87 |
22.63 |
0.36 |
0.53 |
| 2 |
A |
3192 |
3192 |
9.11 |
102.87 |
0.43 |
0.60 |
| 3 |
A |
3134 |
3134 |
8.80 |
97.14 |
0.09 |
0.17 |
| 4 |
A |
1564 |
1564 |
4.38 |
10.51 |
0.74 |
0.85 |
| 5 |
A |
1473 |
1473 |
6.76 |
3.93 |
0.74 |
0.85 |
| 6 |
A |
1432 |
1432 |
20.20 |
6.70 |
0.75 |
0.86 |
| 7 |
A |
1363 |
1363 |
14.95 |
3.69 |
0.74 |
0.85 |
| 8 |
A |
1264 |
1264 |
7.91 |
10.85 |
0.73 |
0.85 |
| 9 |
A |
1169 |
1169 |
21.06 |
4.52 |
0.71 |
0.83 |
| 10 |
A |
1135 |
1135 |
51.04 |
5.05 |
0.62 |
0.76 |
| 11 |
A |
1066 |
1066 |
152.28 |
2.35 |
0.42 |
0.59 |
| 12 |
A |
1032 |
1032 |
22.38 |
8.93 |
0.47 |
0.64 |
| 13 |
A |
917 |
917 |
43.11 |
6.71 |
0.31 |
0.47 |
| 14 |
A |
541 |
541 |
6.72 |
2.83 |
0.38 |
0.55 |
| 15 |
A |
464 |
464 |
24.77 |
1.28 |
0.72 |
0.84 |
| 16 |
A |
404 |
404 |
8.87 |
2.16 |
0.61 |
0.75 |
| 17 |
A |
236 |
236 |
13.07 |
0.11 |
0.61 |
0.76 |
| 18 |
A |
121 |
121 |
11.97 |
0.07 |
0.71 |
0.83 |
Unscaled Zero Point Vibrational Energy (zpe) 11858.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11858.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/6-31G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.752 |
-0.603 |
-0.315 |
| C2 |
0.453 |
0.021 |
0.340 |
| F3 |
-1.921 |
0.082 |
0.165 |
| F4 |
1.576 |
-0.778 |
0.004 |
| F5 |
0.671 |
1.312 |
-0.184 |
| H6 |
-0.695 |
-0.487 |
-1.397 |
| H7 |
-0.830 |
-1.655 |
-0.042 |
| H8 |
0.391 |
0.099 |
1.423 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5069 | 1.4369 | 2.3562 | 2.3889 | 1.0900 | 1.0900 | 2.1957 |
C2 | 1.5069 | | 2.3814 | 1.4180 | 1.4103 | 2.1434 | 2.1451 | 1.0876 | F3 | 1.4369 | 2.3814 | | 3.6043 | 2.8898 | 2.0661 | 2.0611 | 2.6320 | F4 | 2.3562 | 1.4180 | 3.6043 | | 2.2852 | 2.6836 | 2.5613 | 2.0465 | F5 | 2.3889 | 1.4103 | 2.8898 | 2.2852 | | 2.5638 | 3.3279 | 2.0322 | H6 | 1.0900 | 2.1434 | 2.0661 | 2.6836 | 2.5638 | | 1.7940 | 3.0785 | H7 | 1.0900 | 2.1451 | 2.0611 | 2.5613 | 3.3279 | 1.7940 | | 2.5918 | H8 | 2.1957 | 1.0876 | 2.6320 | 2.0465 | 2.0322 | 3.0785 | 2.5918 | |
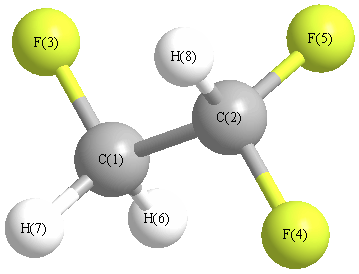 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.286 |
|
C1 |
C2 |
F5 |
109.903 |
| C1 |
C2 |
H8 |
114.651 |
|
C2 |
C1 |
F3 |
107.962 |
| C2 |
C1 |
H6 |
110.218 |
|
C2 |
C1 |
H7 |
110.357 |
| F3 |
C1 |
H6 |
108.932 |
|
F3 |
C1 |
H7 |
108.533 |
| F4 |
C2 |
F5 |
107.795 |
|
F4 |
C2 |
H8 |
108.807 |
| F5 |
C2 |
H8 |
108.194 |
|
H6 |
C1 |
H7 |
110.765 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/6-31G
| | hartrees |
|---|
| Energy at 0K | -377.037530 |
| Energy at 298.15K | |
| HF Energy | -376.853186 |
| Nuclear repulsion energy | 189.240588 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3155 |
3155 |
36.63 |
93.29 |
0.15 |
0.26 |
| 2 |
A' |
3120 |
3120 |
7.73 |
82.89 |
0.18 |
0.30 |
| 3 |
A' |
1560 |
1560 |
4.18 |
10.83 |
0.74 |
0.85 |
| 4 |
A' |
1470 |
1470 |
6.19 |
1.27 |
0.69 |
0.82 |
| 5 |
A' |
1427 |
1427 |
25.01 |
7.69 |
0.71 |
0.83 |
| 6 |
A' |
1163 |
1163 |
68.04 |
6.96 |
0.48 |
0.65 |
| 7 |
A' |
1046 |
1046 |
6.66 |
3.68 |
0.73 |
0.84 |
| 8 |
A' |
882 |
882 |
45.91 |
8.60 |
0.08 |
0.15 |
| 9 |
A' |
730 |
730 |
53.46 |
4.26 |
0.58 |
0.73 |
| 10 |
A' |
465 |
465 |
16.92 |
2.56 |
0.75 |
0.85 |
| 11 |
A' |
228 |
228 |
2.49 |
0.26 |
0.57 |
0.73 |
| 12 |
A" |
3185 |
3185 |
14.69 |
54.25 |
0.75 |
0.86 |
| 13 |
A" |
1422 |
1422 |
24.83 |
2.31 |
0.75 |
0.86 |
| 14 |
A" |
1281 |
1281 |
10.00 |
21.95 |
0.75 |
0.86 |
| 15 |
A" |
1136 |
1136 |
51.26 |
0.49 |
0.75 |
0.86 |
| 16 |
A" |
965 |
965 |
78.10 |
5.76 |
0.75 |
0.86 |
| 17 |
A" |
363 |
363 |
0.13 |
0.39 |
0.75 |
0.86 |
| 18 |
A" |
113 |
113 |
3.10 |
0.15 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11854.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11854.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.772 |
-0.818 |
0.000 |
| C2 |
0.376 |
0.634 |
0.000 |
| F3 |
-0.401 |
-1.639 |
0.000 |
| F4 |
-0.401 |
0.925 |
1.142 |
| F5 |
-0.401 |
0.925 |
-1.142 |
| H6 |
1.352 |
-1.047 |
-0.896 |
| H7 |
1.352 |
-1.047 |
0.896 |
| H8 |
1.236 |
1.305 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5058 | 1.4323 | 2.3916 | 2.3916 | 1.0912 | 1.0912 | 2.1737 |
C2 | 1.5058 | | 2.4030 | 1.4116 | 1.4116 | 2.1404 | 2.1404 | 1.0906 | F3 | 1.4323 | 2.4030 | | 2.8071 | 2.8071 | 2.0559 | 2.0559 | 3.3692 | F4 | 2.3916 | 1.4116 | 2.8071 | | 2.2842 | 3.3337 | 2.6496 | 2.0320 | F5 | 2.3916 | 1.4116 | 2.8071 | 2.2842 | | 2.6496 | 3.3337 | 2.0320 | H6 | 1.0912 | 2.1404 | 2.0559 | 3.3337 | 2.6496 | | 1.7920 | 2.5197 | H7 | 1.0912 | 2.1404 | 2.0559 | 2.6496 | 3.3337 | 1.7920 | | 2.5197 | H8 | 2.1737 | 1.0906 | 3.3692 | 2.0320 | 2.0320 | 2.5197 | 2.5197 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.081 |
|
C1 |
C2 |
F5 |
110.081 |
| C1 |
C2 |
H8 |
112.708 |
|
C2 |
C1 |
F3 |
109.722 |
| C2 |
C1 |
H6 |
109.985 |
|
C2 |
C1 |
H7 |
109.985 |
| F3 |
C1 |
H6 |
108.359 |
|
F3 |
C1 |
H7 |
108.359 |
| F4 |
C2 |
F5 |
108.008 |
|
F4 |
C2 |
H8 |
107.908 |
| F5 |
C2 |
H8 |
107.908 |
|
H6 |
C1 |
H7 |
110.395 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
