Jump to
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -218.022097 |
| Energy at 298.15K | -218.029949 |
| HF Energy | -217.889699 |
| Nuclear repulsion energy | 126.364313 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3151 |
3151 |
31.81 |
|
|
|
| 2 |
A' |
3089 |
3089 |
52.73 |
|
|
|
| 3 |
A' |
3078 |
3078 |
5.28 |
|
|
|
| 4 |
A' |
3062 |
3062 |
22.19 |
|
|
|
| 5 |
A' |
1585 |
1585 |
2.78 |
|
|
|
| 6 |
A' |
1570 |
1570 |
4.34 |
|
|
|
| 7 |
A' |
1561 |
1561 |
2.18 |
|
|
|
| 8 |
A' |
1483 |
1483 |
8.75 |
|
|
|
| 9 |
A' |
1455 |
1455 |
6.87 |
|
|
|
| 10 |
A' |
1382 |
1382 |
2.17 |
|
|
|
| 11 |
A' |
1174 |
1174 |
0.37 |
|
|
|
| 12 |
A' |
1068 |
1068 |
8.77 |
|
|
|
| 13 |
A' |
1007 |
1007 |
59.63 |
|
|
|
| 14 |
A' |
920 |
920 |
12.11 |
|
|
|
| 15 |
A' |
443 |
443 |
6.74 |
|
|
|
| 16 |
A' |
266 |
266 |
5.39 |
|
|
|
| 17 |
A" |
3156 |
3156 |
91.57 |
|
|
|
| 18 |
A" |
3134 |
3134 |
7.32 |
|
|
|
| 19 |
A" |
3113 |
3113 |
0.74 |
|
|
|
| 20 |
A" |
1566 |
1566 |
8.88 |
|
|
|
| 21 |
A" |
1357 |
1357 |
0.49 |
|
|
|
| 22 |
A" |
1299 |
1299 |
0.00 |
|
|
|
| 23 |
A" |
1204 |
1204 |
0.91 |
|
|
|
| 24 |
A" |
935 |
935 |
1.95 |
|
|
|
| 25 |
A" |
800 |
800 |
2.45 |
|
|
|
| 26 |
A" |
235 |
235 |
0.00 |
|
|
|
| 27 |
A" |
130 |
130 |
4.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21610.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21610.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.091 |
-0.793 |
0.000 |
| C2 |
0.000 |
0.725 |
0.000 |
| C3 |
-1.459 |
1.214 |
0.000 |
| F4 |
1.487 |
-1.184 |
0.000 |
| H5 |
-0.366 |
-1.229 |
0.893 |
| H6 |
-0.366 |
-1.229 |
-0.893 |
| H7 |
0.529 |
1.107 |
-0.881 |
| H8 |
0.529 |
1.107 |
0.881 |
| H9 |
-1.501 |
2.307 |
0.000 |
| H10 |
-1.998 |
0.858 |
-0.886 |
| H11 |
-1.998 |
0.858 |
0.886 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5206 | 2.5354 | 1.4501 | 1.0939 | 1.0939 | 2.1394 | 2.1394 | 3.4845 | 2.8060 | 2.8060 |
C2 | 1.5206 | | 1.5389 | 2.4202 | 2.1797 | 2.1797 | 1.0958 | 1.0958 | 2.1807 | 2.1896 | 2.1896 | C3 | 2.5354 | 1.5389 | | 3.7988 | 2.8215 | 2.8215 | 2.1769 | 2.1769 | 1.0939 | 1.0962 | 1.0962 | F4 | 1.4501 | 2.4202 | 3.7988 | | 2.0573 | 2.0573 | 2.6354 | 2.6354 | 4.5954 | 4.1354 | 4.1354 | H5 | 1.0939 | 2.1797 | 2.8215 | 2.0573 | | 1.7861 | 3.0668 | 2.5018 | 3.8197 | 3.1918 | 2.6498 | H6 | 1.0939 | 2.1797 | 2.8215 | 2.0573 | 1.7861 | | 2.5018 | 3.0668 | 3.8197 | 2.6498 | 3.1918 | H7 | 2.1394 | 1.0958 | 2.1769 | 2.6354 | 3.0668 | 2.5018 | | 1.7613 | 2.5170 | 2.5385 | 3.0929 | H8 | 2.1394 | 1.0958 | 2.1769 | 2.6354 | 2.5018 | 3.0668 | 1.7613 | | 2.5170 | 3.0929 | 2.5385 | H9 | 3.4845 | 2.1807 | 1.0939 | 4.5954 | 3.8197 | 3.8197 | 2.5170 | 2.5170 | | 1.7691 | 1.7691 | H10 | 2.8060 | 2.1896 | 1.0962 | 4.1354 | 3.1918 | 2.6498 | 2.5385 | 3.0929 | 1.7691 | | 1.7727 | H11 | 2.8060 | 2.1896 | 1.0962 | 4.1354 | 2.6498 | 3.1918 | 3.0929 | 2.5385 | 1.7691 | 1.7727 | |
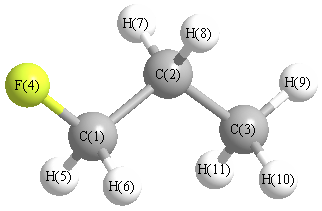 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.928 |
|
C1 |
C2 |
H7 |
108.616 |
| C1 |
C2 |
H8 |
108.616 |
|
C2 |
C1 |
F4 |
109.087 |
| C2 |
C1 |
H5 |
111.928 |
|
C2 |
C1 |
H6 |
111.928 |
| C2 |
C3 |
H9 |
110.708 |
|
C2 |
C3 |
H10 |
111.276 |
| C2 |
C3 |
H11 |
111.276 |
|
C3 |
C2 |
H7 |
110.288 |
| C3 |
C2 |
H8 |
110.288 |
|
F4 |
C1 |
H5 |
107.108 |
| F4 |
C1 |
H6 |
107.108 |
|
H5 |
C1 |
H6 |
109.450 |
| H7 |
C2 |
H8 |
106.954 |
|
H9 |
C3 |
H10 |
107.756 |
| H9 |
C3 |
H11 |
107.756 |
|
H10 |
C3 |
H11 |
107.911 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.079 |
|
|
|
| 2 |
C |
-0.289 |
|
|
|
| 3 |
C |
-0.428 |
|
|
|
| 4 |
F |
-0.377 |
|
|
|
| 5 |
H |
0.138 |
|
|
|
| 6 |
H |
0.138 |
|
|
|
| 7 |
H |
0.154 |
|
|
|
| 8 |
H |
0.154 |
|
|
|
| 9 |
H |
0.151 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
| 11 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.183 |
0.939 |
0.000 |
2.376 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.981 |
2.750 |
0.000 |
| y |
2.750 |
-26.126 |
0.000 |
| z |
0.000 |
0.000 |
-24.770 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.533 |
2.750 |
0.000 |
| y |
2.750 |
0.750 |
0.000 |
| z |
0.000 |
0.000 |
2.783 |
|
| Polar |
|---|
| 3z2-r2 | 5.566 |
|---|
| x2-y2 | -2.855 |
|---|
| xy | 2.750 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.021 |
-0.276 |
0.000 |
| y |
-0.276 |
4.862 |
0.000 |
| z |
0.000 |
0.000 |
4.554 |
<r2> (average value of r
2) Å
2
| <r2> |
105.792 |
| (<r2>)1/2 |
10.286 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -218.022639 |
| Energy at 298.15K | |
| HF Energy | -217.890123 |
| Nuclear repulsion energy | 128.968775 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3165 |
3165 |
24.08 |
|
|
|
| 2 |
A |
3154 |
3154 |
60.39 |
|
|
|
| 3 |
A |
3142 |
3142 |
35.65 |
|
|
|
| 4 |
A |
3108 |
3108 |
21.72 |
|
|
|
| 5 |
A |
3090 |
3090 |
26.00 |
|
|
|
| 6 |
A |
3067 |
3067 |
21.81 |
|
|
|
| 7 |
A |
3061 |
3061 |
24.37 |
|
|
|
| 8 |
A |
1576 |
1576 |
0.73 |
|
|
|
| 9 |
A |
1574 |
1574 |
9.84 |
|
|
|
| 10 |
A |
1559 |
1559 |
7.01 |
|
|
|
| 11 |
A |
1547 |
1547 |
2.33 |
|
|
|
| 12 |
A |
1479 |
1479 |
6.71 |
|
|
|
| 13 |
A |
1445 |
1445 |
9.67 |
|
|
|
| 14 |
A |
1436 |
1436 |
1.54 |
|
|
|
| 15 |
A |
1341 |
1341 |
0.40 |
|
|
|
| 16 |
A |
1285 |
1285 |
0.32 |
|
|
|
| 17 |
A |
1199 |
1199 |
1.65 |
|
|
|
| 18 |
A |
1149 |
1149 |
5.51 |
|
|
|
| 19 |
A |
1101 |
1101 |
12.08 |
|
|
|
| 20 |
A |
976 |
976 |
30.60 |
|
|
|
| 21 |
A |
948 |
948 |
24.94 |
|
|
|
| 22 |
A |
889 |
889 |
6.41 |
|
|
|
| 23 |
A |
796 |
796 |
1.33 |
|
|
|
| 24 |
A |
481 |
481 |
4.27 |
|
|
|
| 25 |
A |
318 |
318 |
2.15 |
|
|
|
| 26 |
A |
228 |
228 |
2.74 |
|
|
|
| 27 |
A |
147 |
147 |
3.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21629.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21629.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.790 |
0.556 |
0.297 |
| C2 |
-0.609 |
0.665 |
-0.289 |
| C3 |
-1.518 |
-0.503 |
0.124 |
| F4 |
1.400 |
-0.680 |
-0.167 |
| H5 |
1.441 |
1.372 |
-0.026 |
| H6 |
0.772 |
0.507 |
1.390 |
| H7 |
-0.527 |
0.709 |
-1.382 |
| H8 |
-1.040 |
1.622 |
0.037 |
| H9 |
-2.504 |
-0.418 |
-0.343 |
| H10 |
-1.070 |
-1.452 |
-0.181 |
| H11 |
-1.659 |
-0.527 |
1.211 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5203 | 2.5448 | 1.4540 | 1.0930 | 1.0940 | 2.1395 | 2.1337 | 3.4938 | 2.7784 | 2.8288 |
C2 | 1.5203 | | 1.5370 | 2.4208 | 2.1837 | 2.1797 | 1.0964 | 1.0985 | 2.1838 | 2.1700 | 2.1852 | C3 | 2.5448 | 1.5370 | | 2.9381 | 3.5059 | 2.8049 | 2.1717 | 2.1793 | 1.0944 | 1.0933 | 1.0965 | F4 | 1.4540 | 2.4208 | 2.9381 | | 2.0567 | 2.0559 | 2.6683 | 3.3605 | 3.9169 | 2.5884 | 3.3586 | H5 | 1.0930 | 2.1837 | 3.5059 | 2.0567 | | 1.7889 | 2.4802 | 2.4943 | 4.3434 | 3.7820 | 3.8397 | H6 | 1.0940 | 2.1797 | 2.8049 | 2.0559 | 1.7889 | | 3.0679 | 2.5215 | 3.8201 | 3.1147 | 2.6477 | H7 | 2.1395 | 1.0964 | 2.1717 | 2.6683 | 2.4802 | 3.0679 | | 1.7629 | 2.5008 | 2.5310 | 3.0869 | H8 | 2.1337 | 1.0985 | 2.1793 | 3.3605 | 2.4943 | 2.5215 | 1.7629 | | 2.5392 | 3.0818 | 2.5255 | H9 | 3.4938 | 2.1838 | 1.0944 | 3.9169 | 4.3434 | 3.8201 | 2.5008 | 2.5392 | | 1.7754 | 1.7727 | H10 | 2.7784 | 2.1700 | 1.0933 | 2.5884 | 3.7820 | 3.1147 | 2.5310 | 3.0818 | 1.7754 | | 1.7724 | H11 | 2.8288 | 2.1852 | 1.0965 | 3.3586 | 3.8397 | 2.6477 | 3.0869 | 2.5255 | 1.7727 | 1.7724 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.684 |
|
C1 |
C2 |
H7 |
108.608 |
| C1 |
C2 |
H8 |
108.041 |
|
C2 |
C1 |
F4 |
108.933 |
| C2 |
C1 |
H5 |
112.328 |
|
C2 |
C1 |
H6 |
111.942 |
| C2 |
C3 |
H9 |
111.059 |
|
C2 |
C3 |
H10 |
110.037 |
| C2 |
C3 |
H11 |
111.044 |
|
C3 |
C2 |
H7 |
109.981 |
| C3 |
C2 |
H8 |
110.460 |
|
F4 |
C1 |
H5 |
106.846 |
| F4 |
C1 |
H6 |
106.731 |
|
H5 |
C1 |
H6 |
109.773 |
| H7 |
C2 |
H8 |
106.870 |
|
H9 |
C3 |
H10 |
108.495 |
| H9 |
C3 |
H11 |
108.026 |
|
H10 |
C3 |
H11 |
108.082 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.072 |
|
|
|
| 2 |
C |
-0.291 |
|
|
|
| 3 |
C |
-0.410 |
|
|
|
| 4 |
F |
-0.376 |
|
|
|
| 5 |
H |
0.142 |
|
|
|
| 6 |
H |
0.139 |
|
|
|
| 7 |
H |
0.154 |
|
|
|
| 8 |
H |
0.135 |
|
|
|
| 9 |
H |
0.138 |
|
|
|
| 10 |
H |
0.163 |
|
|
|
| 11 |
H |
0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.141 |
1.810 |
0.567 |
2.214 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.354 |
2.293 |
0.813 |
| y |
2.293 |
-25.526 |
0.027 |
| z |
0.813 |
0.027 |
-24.844 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.169 |
2.293 |
0.813 |
| y |
2.293 |
0.573 |
0.027 |
| z |
0.813 |
0.027 |
1.596 |
|
| Polar |
|---|
| 3z2-r2 | 3.191 |
|---|
| x2-y2 | -1.828 |
|---|
| xy | 2.293 |
|---|
| xz | 0.813 |
|---|
| yz | 0.027 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.001 |
0.116 |
-0.006 |
| y |
0.116 |
4.799 |
0.029 |
| z |
-0.006 |
0.029 |
4.606 |
<r2> (average value of r
2) Å
2
| <r2> |
92.670 |
| (<r2>)1/2 |
9.627 |
