Jump to
S1C2
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -357.040785 |
| Energy at 298.15K | -357.045951 |
| HF Energy | -356.358883 |
| Nuclear repulsion energy | 227.481363 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3754 |
3567 |
68.40 |
|
|
|
| 2 |
A' |
3608 |
3428 |
81.76 |
|
|
|
| 3 |
A' |
3590 |
3411 |
63.47 |
|
|
|
| 4 |
A' |
1726 |
1640 |
181.16 |
|
|
|
| 5 |
A' |
1668 |
1585 |
116.70 |
|
|
|
| 6 |
A' |
1648 |
1566 |
147.76 |
|
|
|
| 7 |
A' |
1442 |
1370 |
1.75 |
|
|
|
| 8 |
A' |
1341 |
1274 |
79.74 |
|
|
|
| 9 |
A' |
1134 |
1078 |
220.76 |
|
|
|
| 10 |
A' |
1106 |
1050 |
92.84 |
|
|
|
| 11 |
A' |
765 |
727 |
1.19 |
|
|
|
| 12 |
A' |
605 |
575 |
68.91 |
|
|
|
| 13 |
A' |
533 |
506 |
2.04 |
|
|
|
| 14 |
A' |
422 |
401 |
6.66 |
|
|
|
| 15 |
A' |
277 |
264 |
12.21 |
|
|
|
| 16 |
A" |
763 |
725 |
0.03 |
|
|
|
| 17 |
A" |
659 |
626 |
340.18 |
|
|
|
| 18 |
A" |
610 |
580 |
12.24 |
|
|
|
| 19 |
A" |
567 |
538 |
265.19 |
|
|
|
| 20 |
A" |
412 |
391 |
16.07 |
|
|
|
| 21 |
A" |
69 |
66 |
6.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13349.1 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 12683.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.747 |
0.000 |
| C2 |
-0.042 |
-0.784 |
0.000 |
| O3 |
-1.117 |
-1.444 |
0.000 |
| O4 |
1.065 |
1.414 |
0.000 |
| O5 |
-1.255 |
1.311 |
0.000 |
| N6 |
1.219 |
-1.324 |
0.000 |
| H7 |
1.321 |
-2.329 |
0.000 |
| H8 |
2.038 |
-0.731 |
0.000 |
| H9 |
-1.181 |
2.294 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5310 | 2.4587 | 1.2567 | 1.3760 | 2.4024 | 3.3473 | 2.5170 | 1.9467 |
C2 | 1.5310 | | 1.2616 | 2.4609 | 2.4208 | 1.3714 | 2.0603 | 2.0804 | 3.2821 | O3 | 2.4587 | 1.2616 | | 3.5956 | 2.7582 | 2.3389 | 2.5937 | 3.2345 | 3.7385 | O4 | 1.2567 | 2.4609 | 3.5956 | | 2.3219 | 2.7423 | 3.7521 | 2.3554 | 2.4118 | O5 | 1.3760 | 2.4208 | 2.7582 | 2.3219 | | 3.6140 | 4.4593 | 3.8744 | 0.9860 | N6 | 2.4024 | 1.3714 | 2.3389 | 2.7423 | 3.6140 | | 1.0105 | 1.0113 | 4.3414 | H7 | 3.3473 | 2.0603 | 2.5937 | 3.7521 | 4.4593 | 1.0105 | | 1.7518 | 5.2568 | H8 | 2.5170 | 2.0804 | 3.2345 | 2.3554 | 3.8744 | 1.0113 | 1.7518 | | 4.4170 | H9 | 1.9467 | 3.2821 | 3.7385 | 2.4118 | 0.9860 | 4.3414 | 5.2568 | 4.4170 | |
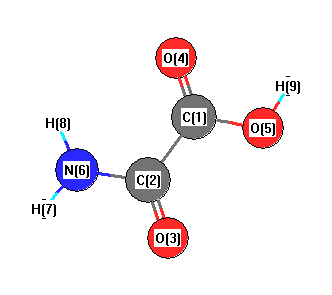 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
123.103 |
|
C1 |
C2 |
N6 |
111.621 |
| C1 |
O5 |
H9 |
109.909 |
|
C2 |
C1 |
O4 |
123.656 |
| C2 |
C1 |
O5 |
112.654 |
|
C2 |
N6 |
H7 |
118.988 |
| C2 |
N6 |
H8 |
120.907 |
|
O3 |
C2 |
N6 |
125.276 |
| O4 |
C1 |
O5 |
123.689 |
|
H7 |
N6 |
H8 |
120.105 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -357.044387 |
| Energy at 298.15K | -357.049695 |
| Nuclear repulsion energy | 228.572871 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3742 |
3555 |
75.78 |
|
|
|
| 2 |
A' |
3597 |
3418 |
96.51 |
|
|
|
| 3 |
A' |
3512 |
3336 |
97.64 |
|
|
|
| 4 |
A' |
1734 |
1648 |
212.66 |
|
|
|
| 5 |
A' |
1711 |
1625 |
160.73 |
|
|
|
| 6 |
A' |
1655 |
1572 |
9.78 |
|
|
|
| 7 |
A' |
1430 |
1359 |
17.33 |
|
|
|
| 8 |
A' |
1298 |
1233 |
517.41 |
|
|
|
| 9 |
A' |
1180 |
1121 |
23.53 |
|
|
|
| 10 |
A' |
1117 |
1061 |
1.27 |
|
|
|
| 11 |
A' |
777 |
738 |
18.69 |
|
|
|
| 12 |
A' |
616 |
585 |
10.35 |
|
|
|
| 13 |
A' |
545 |
517 |
1.69 |
|
|
|
| 14 |
A' |
415 |
394 |
5.18 |
|
|
|
| 15 |
A' |
278 |
265 |
38.98 |
|
|
|
| 16 |
A" |
733 |
696 |
32.59 |
|
|
|
| 17 |
A" |
686 |
652 |
207.40 |
|
|
|
| 18 |
A" |
630 |
599 |
172.94 |
|
|
|
| 19 |
A" |
618 |
587 |
203.13 |
|
|
|
| 20 |
A" |
414 |
393 |
2.33 |
|
|
|
| 21 |
A" |
113 |
107 |
8.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13399.3 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 12730.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.024 |
-0.787 |
0.000 |
| C2 |
0.000 |
0.751 |
0.000 |
| O3 |
-1.116 |
1.360 |
0.000 |
| O4 |
1.074 |
-1.464 |
0.000 |
| O5 |
-1.235 |
-1.338 |
0.000 |
| N6 |
1.230 |
1.326 |
0.000 |
| H7 |
1.316 |
2.334 |
0.000 |
| H8 |
2.060 |
0.746 |
0.000 |
| H9 |
-1.912 |
-0.616 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5376 | 2.4304 | 1.2491 | 1.3743 | 2.4327 | 3.3770 | 2.5485 | 1.9437 |
C2 | 1.5376 | | 1.2713 | 2.4610 | 2.4266 | 1.3579 | 2.0582 | 2.0600 | 2.3504 | O3 | 2.4304 | 1.2713 | | 3.5731 | 2.7007 | 2.3461 | 2.6191 | 3.2346 | 2.1303 | O4 | 1.2491 | 2.4610 | 3.5731 | | 2.3119 | 2.7942 | 3.8050 | 2.4200 | 3.1039 | O5 | 1.3743 | 2.4266 | 2.7007 | 2.3119 | | 3.6296 | 4.4706 | 3.8987 | 0.9902 | N6 | 2.4327 | 1.3579 | 2.3461 | 2.7942 | 3.6296 | | 1.0111 | 1.0126 | 3.6939 | H7 | 3.3770 | 2.0582 | 2.6191 | 3.8050 | 4.4706 | 1.0111 | | 1.7533 | 4.3725 | H8 | 2.5485 | 2.0600 | 3.2346 | 2.4200 | 3.8987 | 1.0126 | 1.7533 | | 4.1992 | H9 | 1.9437 | 2.3504 | 2.1303 | 3.1039 | 0.9902 | 3.6939 | 4.3725 | 4.1992 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.527 |
|
C1 |
C2 |
N6 |
114.175 |
| C1 |
O5 |
H9 |
109.505 |
|
C2 |
C1 |
O4 |
123.717 |
| C2 |
C1 |
O5 |
112.764 |
|
C2 |
N6 |
H7 |
119.926 |
| C2 |
N6 |
H8 |
119.985 |
|
O3 |
C2 |
N6 |
126.298 |
| O4 |
C1 |
O5 |
123.519 |
|
H7 |
N6 |
H8 |
120.089 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
