Jump to
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -192.758352 |
| Energy at 298.15K | -192.765520 |
| HF Energy | -192.625307 |
| Nuclear repulsion energy | 125.697101 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3117 |
3117 |
18.22 |
|
|
|
| 2 |
A1 |
3100 |
3100 |
10.46 |
|
|
|
| 3 |
A1 |
1595 |
1595 |
0.10 |
|
|
|
| 4 |
A1 |
1554 |
1554 |
2.73 |
|
|
|
| 5 |
A1 |
1380 |
1380 |
0.18 |
|
|
|
| 6 |
A1 |
1022 |
1022 |
0.00 |
|
|
|
| 7 |
A1 |
853 |
853 |
12.71 |
|
|
|
| 8 |
A1 |
790 |
790 |
10.41 |
|
|
|
| 9 |
A2 |
3153 |
3153 |
0.00 |
|
|
|
| 10 |
A2 |
1274 |
1274 |
0.00 |
|
|
|
| 11 |
A2 |
1135 |
1135 |
0.00 |
|
|
|
| 12 |
A2 |
881 |
881 |
0.00 |
|
|
|
| 13 |
B1 |
3180 |
3180 |
75.21 |
|
|
|
| 14 |
B1 |
3147 |
3147 |
31.16 |
|
|
|
| 15 |
B1 |
1229 |
1229 |
2.24 |
|
|
|
| 16 |
B1 |
1120 |
1120 |
0.65 |
|
|
|
| 17 |
B1 |
797 |
797 |
0.73 |
|
|
|
| 18 |
B1 |
50 |
50 |
5.23 |
|
|
|
| 19 |
B2 |
3093 |
3093 |
110.18 |
|
|
|
| 20 |
B2 |
1572 |
1572 |
0.95 |
|
|
|
| 21 |
B2 |
1328 |
1328 |
13.42 |
|
|
|
| 22 |
B2 |
1310 |
1310 |
0.84 |
|
|
|
| 23 |
B2 |
980 |
980 |
12.57 |
|
|
|
| 24 |
B2 |
918 |
918 |
57.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19290.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 19290.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.107 |
| C2 |
0.000 |
0.000 |
-1.077 |
| C3 |
0.000 |
1.069 |
0.047 |
| C4 |
0.000 |
-1.069 |
0.047 |
| H5 |
0.888 |
0.000 |
-1.711 |
| H6 |
-0.888 |
0.000 |
-1.711 |
| H7 |
0.894 |
1.692 |
0.115 |
| H8 |
-0.894 |
1.692 |
0.115 |
| H9 |
-0.894 |
-1.692 |
0.115 |
| H10 |
0.894 |
-1.692 |
0.115 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1842 | 1.5059 | 1.5059 | 2.9556 | 2.9556 | 2.1560 | 2.1560 | 2.1560 | 2.1560 |
C2 | 2.1842 | | 1.5508 | 1.5508 | 1.0918 | 1.0918 | 2.2549 | 2.2549 | 2.2549 | 2.2549 | C3 | 1.5059 | 1.5508 | | 2.1380 | 2.2413 | 2.2413 | 1.0923 | 1.0923 | 2.9033 | 2.9033 | C4 | 1.5059 | 1.5508 | 2.1380 | | 2.2413 | 2.2413 | 2.9033 | 2.9033 | 1.0923 | 1.0923 | H5 | 2.9556 | 1.0918 | 2.2413 | 2.2413 | | 1.7768 | 2.4902 | 3.0625 | 3.0625 | 2.4902 | H6 | 2.9556 | 1.0918 | 2.2413 | 2.2413 | 1.7768 | | 3.0625 | 2.4902 | 2.4902 | 3.0625 | H7 | 2.1560 | 2.2549 | 1.0923 | 2.9033 | 2.4902 | 3.0625 | | 1.7888 | 3.8282 | 3.3846 | H8 | 2.1560 | 2.2549 | 1.0923 | 2.9033 | 3.0625 | 2.4902 | 1.7888 | | 3.3846 | 3.8282 | H9 | 2.1560 | 2.2549 | 2.9033 | 1.0923 | 3.0625 | 2.4902 | 3.8282 | 3.3846 | | 1.7888 | H10 | 2.1560 | 2.2549 | 2.9033 | 1.0923 | 2.4902 | 3.0625 | 3.3846 | 3.8282 | 1.7888 | |
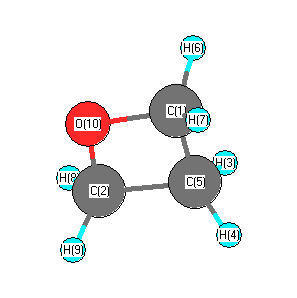 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.201 |
|
O1 |
C3 |
H7 |
111.158 |
| O1 |
C3 |
H8 |
111.158 |
|
O1 |
C4 |
C2 |
91.201 |
| O1 |
C4 |
H9 |
111.158 |
|
O1 |
C4 |
H10 |
111.158 |
| C2 |
C3 |
H7 |
116.025 |
|
C2 |
C3 |
H8 |
116.025 |
| C2 |
C4 |
H9 |
116.025 |
|
C2 |
C4 |
H10 |
116.025 |
| C3 |
O1 |
C4 |
90.448 |
|
C3 |
C2 |
C4 |
87.151 |
| C3 |
C2 |
H5 |
114.909 |
|
C3 |
C2 |
H6 |
114.909 |
| C4 |
C2 |
H5 |
114.909 |
|
C4 |
C2 |
H6 |
114.909 |
| H5 |
C2 |
H6 |
108.908 |
|
H7 |
C3 |
H8 |
109.930 |
| H9 |
C4 |
H10 |
109.930 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.542 |
|
|
|
| 2 |
C |
-0.359 |
|
|
|
| 3 |
C |
0.002 |
|
|
|
| 4 |
C |
0.002 |
|
|
|
| 5 |
H |
0.155 |
|
|
|
| 6 |
H |
0.155 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.585 |
2.585 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.450 |
0.000 |
0.000 |
| y |
0.000 |
-22.451 |
0.000 |
| z |
0.000 |
0.000 |
-28.786 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.168 |
0.000 |
0.000 |
| y |
0.000 |
4.168 |
0.000 |
| z |
0.000 |
0.000 |
-5.336 |
|
| Polar |
|---|
| 3z2-r2 | -10.671 |
|---|
| x2-y2 | -2.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.460 |
0.000 |
0.000 |
| y |
0.000 |
5.629 |
0.000 |
| z |
0.000 |
0.000 |
4.403 |
<r2> (average value of r
2) Å
2
| <r2> |
68.410 |
| (<r2>)1/2 |
8.271 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -192.758352 |
| Energy at 298.15K | -192.765521 |
| HF Energy | -192.625299 |
| Nuclear repulsion energy | 125.694673 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3179 |
3179 |
75.99 |
|
|
|
| 2 |
A' |
3147 |
3147 |
30.36 |
|
|
|
| 3 |
A' |
3117 |
3117 |
18.03 |
|
|
|
| 4 |
A' |
3100 |
3100 |
10.68 |
|
|
|
| 5 |
A' |
1596 |
1596 |
0.10 |
|
|
|
| 6 |
A' |
1554 |
1554 |
2.73 |
|
|
|
| 7 |
A' |
1380 |
1380 |
0.18 |
|
|
|
| 8 |
A' |
1229 |
1229 |
2.24 |
|
|
|
| 9 |
A' |
1120 |
1120 |
0.64 |
|
|
|
| 10 |
A' |
1023 |
1023 |
0.00 |
|
|
|
| 11 |
A' |
853 |
853 |
12.66 |
|
|
|
| 12 |
A' |
797 |
797 |
0.85 |
|
|
|
| 13 |
A' |
790 |
790 |
10.33 |
|
|
|
| 14 |
A' |
51 |
51 |
5.23 |
|
|
|
| 15 |
A" |
3153 |
3153 |
0.00 |
|
|
|
| 16 |
A" |
3093 |
3093 |
110.14 |
|
|
|
| 17 |
A" |
1572 |
1572 |
0.95 |
|
|
|
| 18 |
A" |
1329 |
1329 |
13.48 |
|
|
|
| 19 |
A" |
1310 |
1310 |
0.78 |
|
|
|
| 20 |
A" |
1275 |
1275 |
0.00 |
|
|
|
| 21 |
A" |
1135 |
1135 |
0.00 |
|
|
|
| 22 |
A" |
981 |
981 |
12.26 |
|
|
|
| 23 |
A" |
918 |
918 |
57.46 |
|
|
|
| 24 |
A" |
881 |
881 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19290.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 19290.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.003 |
-1.108 |
0.000 |
| C2 |
-0.001 |
1.077 |
0.000 |
| C3 |
-0.001 |
-0.047 |
1.069 |
| C4 |
-0.001 |
-0.047 |
-1.069 |
| H5 |
0.888 |
1.711 |
0.000 |
| H6 |
-0.889 |
1.712 |
0.000 |
| H7 |
0.892 |
-0.114 |
1.694 |
| H8 |
-0.896 |
-0.117 |
1.690 |
| H9 |
0.892 |
-0.114 |
-1.694 |
| H10 |
-0.896 |
-0.117 |
-1.690 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1843 | 1.5061 | 1.5061 | 2.9545 | 2.9573 | 2.1561 | 2.1561 | 2.1561 | 2.1561 |
C2 | 2.1843 | | 1.5506 | 1.5506 | 1.0919 | 1.0919 | 2.2551 | 2.2546 | 2.2551 | 2.2546 | C3 | 1.5061 | 1.5506 | | 2.1379 | 2.2411 | 2.2415 | 1.0923 | 1.0923 | 2.9048 | 2.9018 | C4 | 1.5061 | 1.5506 | 2.1379 | | 2.2411 | 2.2415 | 2.9048 | 2.9018 | 1.0923 | 1.0923 | H5 | 2.9545 | 1.0919 | 2.2411 | 2.2411 | | 1.7765 | 2.4904 | 3.0630 | 2.4904 | 3.0630 | H6 | 2.9573 | 1.0919 | 2.2415 | 2.2415 | 1.7765 | | 3.0622 | 2.4902 | 3.0622 | 2.4902 | H7 | 2.1561 | 2.2551 | 1.0923 | 2.9048 | 2.4904 | 3.0622 | | 1.7887 | 3.3887 | 3.8282 | H8 | 2.1561 | 2.2546 | 1.0923 | 2.9018 | 3.0630 | 2.4902 | 1.7887 | | 3.8282 | 3.3806 | H9 | 2.1561 | 2.2551 | 2.9048 | 1.0923 | 2.4904 | 3.0622 | 3.3887 | 3.8282 | | 1.7887 | H10 | 2.1561 | 2.2546 | 2.9018 | 1.0923 | 3.0630 | 2.4902 | 3.8282 | 3.3806 | 1.7887 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.205 |
|
O1 |
C3 |
H7 |
111.154 |
| O1 |
C3 |
H8 |
111.148 |
|
O1 |
C4 |
C2 |
91.205 |
| O1 |
C4 |
H9 |
111.154 |
|
O1 |
C4 |
H10 |
111.148 |
| C2 |
C3 |
H7 |
116.057 |
|
C2 |
C3 |
H8 |
116.009 |
| C2 |
C4 |
H9 |
116.057 |
|
C2 |
C4 |
H10 |
116.009 |
| C3 |
O1 |
C4 |
90.430 |
|
C3 |
C2 |
C4 |
87.159 |
| C3 |
C2 |
H5 |
114.904 |
|
C3 |
C2 |
H6 |
114.930 |
| C4 |
C2 |
H5 |
114.904 |
|
C4 |
C2 |
H6 |
114.930 |
| H5 |
C2 |
H6 |
108.877 |
|
H7 |
C3 |
H8 |
109.923 |
| H9 |
C4 |
H10 |
109.923 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.542 |
|
|
|
| 2 |
C |
-0.359 |
|
|
|
| 3 |
C |
0.002 |
|
|
|
| 4 |
C |
0.002 |
|
|
|
| 5 |
H |
0.155 |
|
|
|
| 6 |
H |
0.155 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.008 |
2.586 |
0.000 |
2.586 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.451 |
0.012 |
0.000 |
| y |
0.012 |
-28.785 |
0.000 |
| z |
0.000 |
0.000 |
-22.450 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.166 |
0.012 |
0.000 |
| y |
0.012 |
-5.335 |
0.000 |
| z |
0.000 |
0.000 |
4.169 |
|
| Polar |
|---|
| 3z2-r2 | 8.337 |
|---|
| x2-y2 | 4.334 |
|---|
| xy | 0.012 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.460 |
-0.001 |
0.000 |
| y |
-0.001 |
4.403 |
0.000 |
| z |
0.000 |
0.000 |
5.629 |
<r2> (average value of r
2) Å
2
| <r2> |
68.412 |
| (<r2>)1/2 |
8.271 |
